return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,2-dichlorobenzene)

using model chemistry: MP2=FULL/6-31+G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31+G**
 hartrees
Energy at 0K-1149.620567
Energy at 298.15K 
HF Energy-1148.509431
Nuclear repulsion energy469.214744
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31+G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3288 3088 6.66      
2 A1 3273 3074 3.15      
3 A1 1638 1539 4.52      
4 A1 1519 1426 51.66      
5 A1 1456 1368 7.13      
6 A1 1207 1134 0.30      
7 A1 1181 1109 40.93      
8 A1 1072 1007 9.21      
9 A1 677 636 13.34      
10 A1 491 461 5.01      
11 A1 206 193 0.02      
12 A2 889 835 0.00      
13 A2 824 774 0.00      
14 A2 431 405 0.00      
15 A2 121 114 0.00      
16 A2 835i 784i 0.00      
17 B1 892 838 0.59      
18 B1 749 703 81.28      
19 B1 400 375 4.10      
20 B1 218 205 1.84      
21 B2 3281 3082 3.21      
22 B2 3263 3064 0.68      
23 B2 1651 1550 9.38      
24 B2 1482 1392 17.12      
25 B2 1299 1220 3.56      
26 B2 1172 1101 2.00      
27 B2 1058 994 37.26      
28 B2 765 719 13.60      
29 B2 435 409 0.21      
30 B2 342 321 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 17222.4 cm-1
Scaled (by 0.9392) Zero Point Vibrational Energy (zpe) 16175.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31+G**
ABC
0.06358 0.04769 0.02725

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31+G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.700 -0.035
C2 0.000 -0.700 -0.035
C3 0.000 1.393 1.179
C4 0.000 -1.393 1.179
C5 0.000 0.698 2.387
C6 0.000 -0.698 2.387
Cl7 0.000 1.603 -1.510
Cl8 0.000 -1.603 -1.510
H9 0.000 2.474 1.163
H10 0.000 -2.474 1.163
H11 0.000 1.247 3.319
H12 0.000 -1.247 3.319

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.40101.39732.41972.42142.79641.72912.73502.13973.39263.39753.8778
C21.40102.41971.39732.79642.42142.73501.72913.39262.13973.87783.3975
C31.39732.41972.78541.39312.41452.69714.02521.08133.86662.14443.3979
C42.41971.39732.78542.41451.39314.02522.69713.86661.08133.39792.1444
C52.42142.79641.39312.41451.39644.00014.52512.15673.40001.08152.1569
C62.79642.42142.41451.39311.39644.52514.00013.40002.15672.15691.0815
Cl71.72912.73502.69714.02524.00014.52513.20522.81084.87434.84145.6066
Cl82.73501.72914.02522.69714.52514.00013.20524.87432.81085.60664.8414
H92.13973.39261.08133.86662.15673.40002.81084.87434.94772.48064.3004
H103.39262.13973.86661.08133.40002.15674.87432.81084.94774.30042.4806
H113.39753.87782.14443.39791.08152.15694.84145.60662.48064.30042.4942
H123.87783.39753.39792.14442.15691.08155.60664.84144.30042.48062.4942

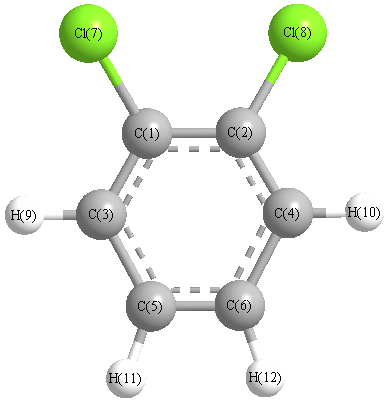
picture of 1,2-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 119.694 C1 C2 Cl8 121.448
C1 C3 C5 120.401 C1 C3 H9 118.816
C2 C1 C3 119.694 C2 C1 Cl7 121.448
C2 C4 C6 120.401 C2 C4 H10 118.816
C3 C1 Cl7 118.858 C3 C5 C6 119.904
C3 C5 H11 119.595 C4 C2 Cl8 118.858
C4 C6 C5 119.904 C4 C6 H12 119.595
C5 C3 H9 120.783 C5 C6 H12 120.501
C6 C4 H10 120.783 C6 C5 H11 120.501
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability