Jump to
S1C2
Energy calculated at QCISD/cc-pVTZ-PP
| | hartrees |
|---|
| Energy at 0K | -909.947842 |
| Energy at 298.15K | |
| HF Energy | -909.113154 |
| Nuclear repulsion energy | 279.912760 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVTZ-PP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3133 |
3133 |
0.00 |
|
|
|
| 2 |
Ag |
1504 |
1504 |
0.00 |
|
|
|
| 3 |
Ag |
1314 |
1314 |
0.00 |
|
|
|
| 4 |
Ag |
1092 |
1092 |
0.00 |
|
|
|
| 5 |
Ag |
705 |
705 |
0.00 |
|
|
|
| 6 |
Ag |
195 |
195 |
0.00 |
|
|
|
| 7 |
Au |
3212 |
3212 |
0.59 |
|
|
|
| 8 |
Au |
1123 |
1123 |
3.26 |
|
|
|
| 9 |
Au |
764 |
764 |
3.00 |
|
|
|
| 10 |
Au |
105 |
105 |
3.98 |
|
|
|
| 11 |
Bg |
3190 |
3190 |
0.00 |
|
|
|
| 12 |
Bg |
1307 |
1307 |
0.00 |
|
|
|
| 13 |
Bg |
964 |
964 |
0.00 |
|
|
|
| 14 |
Bu |
3139 |
3139 |
7.76 |
|
|
|
| 15 |
Bu |
1501 |
1501 |
6.48 |
|
|
|
| 16 |
Bu |
1230 |
1230 |
57.49 |
|
|
|
| 17 |
Bu |
632 |
632 |
53.37 |
|
|
|
| 18 |
Bu |
182 |
182 |
5.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12645.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12645.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVTZ-PP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.491 |
0.573 |
0.000 |
| C2 |
-0.491 |
-0.573 |
0.000 |
| Br3 |
-0.491 |
2.252 |
0.000 |
| Br4 |
0.491 |
-2.252 |
0.000 |
| H5 |
1.113 |
0.573 |
0.888 |
| H6 |
1.113 |
0.573 |
-0.888 |
| H7 |
-1.113 |
-0.573 |
0.888 |
| H8 |
-1.113 |
-0.573 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5085 | 1.9452 | 2.8247 | 1.0844 | 1.0844 | 2.1618 | 2.1618 |
C2 | 1.5085 | | 2.8247 | 1.9452 | 2.1618 | 2.1618 | 1.0844 | 1.0844 | Br3 | 1.9452 | 2.8247 | | 4.6098 | 2.4862 | 2.4862 | 3.0258 | 3.0258 | Br4 | 2.8247 | 1.9452 | 4.6098 | | 3.0258 | 3.0258 | 2.4862 | 2.4862 | H5 | 1.0844 | 2.1618 | 2.4862 | 3.0258 | | 1.7764 | 2.5036 | 3.0698 | H6 | 1.0844 | 2.1618 | 2.4862 | 3.0258 | 1.7764 | | 3.0698 | 2.5036 | H7 | 2.1618 | 1.0844 | 3.0258 | 2.4862 | 2.5036 | 3.0698 | | 1.7764 | H8 | 2.1618 | 1.0844 | 3.0258 | 2.4862 | 3.0698 | 2.5036 | 1.7764 | |
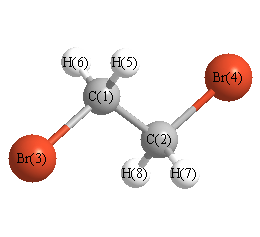 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.090 |
|
C1 |
C2 |
H7 |
111.930 |
| C1 |
C2 |
H8 |
111.930 |
|
C2 |
C1 |
Br3 |
109.090 |
| C2 |
C1 |
H5 |
111.930 |
|
C2 |
C1 |
H6 |
111.930 |
| Br3 |
C1 |
H5 |
106.826 |
|
Br3 |
C1 |
H6 |
106.826 |
| Br4 |
C2 |
H7 |
106.826 |
|
Br4 |
C2 |
H8 |
106.826 |
| H5 |
C1 |
H6 |
109.975 |
|
H7 |
C2 |
H8 |
109.975 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVTZ-PP
| | hartrees |
|---|
| Energy at 0K | -909.944791 |
| Energy at 298.15K | |
| HF Energy | -909.108936 |
| Nuclear repulsion energy | 297.096273 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVTZ-PP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3174 |
3174 |
0.22 |
|
|
|
| 2 |
A |
3118 |
3118 |
17.04 |
|
|
|
| 3 |
A |
1488 |
1488 |
0.21 |
|
|
|
| 4 |
A |
1333 |
1333 |
20.13 |
|
|
|
| 5 |
A |
1214 |
1214 |
1.62 |
|
|
|
| 6 |
A |
1059 |
1059 |
0.46 |
|
|
|
| 7 |
A |
929 |
929 |
5.96 |
|
|
|
| 8 |
A |
584 |
584 |
6.47 |
|
|
|
| 9 |
A |
231 |
231 |
0.90 |
|
|
|
| 10 |
A |
82 |
82 |
0.20 |
|
|
|
| 11 |
B |
3186 |
3186 |
1.65 |
|
|
|
| 12 |
B |
3109 |
3109 |
2.41 |
|
|
|
| 13 |
B |
1483 |
1483 |
9.80 |
|
|
|
| 14 |
B |
1305 |
1305 |
59.88 |
|
|
|
| 15 |
B |
1141 |
1141 |
2.22 |
|
|
|
| 16 |
B |
867 |
867 |
13.74 |
|
|
|
| 17 |
B |
619 |
619 |
11.18 |
|
|
|
| 18 |
B |
362 |
362 |
4.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12641.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12641.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVTZ-PP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.311 |
0.688 |
1.177 |
| C2 |
-0.311 |
-0.688 |
1.177 |
| Br3 |
-0.311 |
1.786 |
-0.293 |
| Br4 |
0.311 |
-1.786 |
-0.293 |
| H5 |
0.032 |
1.216 |
2.085 |
| H6 |
1.393 |
0.641 |
1.099 |
| H7 |
-0.032 |
-1.216 |
2.085 |
| H8 |
-1.393 |
-0.641 |
1.099 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5104 | 1.9367 | 2.8772 | 1.0874 | 1.0852 | 2.1375 | 2.1620 |
C2 | 1.5104 | | 2.8772 | 1.9367 | 2.1375 | 2.1620 | 1.0874 | 1.0852 | Br3 | 1.9367 | 2.8772 | | 3.6250 | 2.4694 | 2.4800 | 3.8396 | 2.9987 | Br4 | 2.8772 | 1.9367 | 3.6250 | | 3.8396 | 2.9987 | 2.4694 | 2.4800 | H5 | 1.0874 | 2.1375 | 2.4694 | 3.8396 | | 1.7763 | 2.4326 | 2.5396 | H6 | 1.0852 | 2.1620 | 2.4800 | 2.9987 | 1.7763 | | 2.5396 | 3.0657 | H7 | 2.1375 | 1.0874 | 3.8396 | 2.4694 | 2.4326 | 2.5396 | | 1.7763 | H8 | 2.1620 | 1.0852 | 2.9987 | 2.4800 | 2.5396 | 3.0657 | 1.7763 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.573 |
|
C1 |
C2 |
H7 |
109.661 |
| C1 |
C2 |
H8 |
111.761 |
|
C2 |
C1 |
Br3 |
112.573 |
| C2 |
C1 |
H5 |
109.661 |
|
C2 |
C1 |
H6 |
111.761 |
| Br3 |
C1 |
H5 |
106.054 |
|
Br3 |
C1 |
H6 |
106.897 |
| Br4 |
C2 |
H7 |
106.054 |
|
Br4 |
C2 |
H8 |
106.897 |
| H5 |
C1 |
H6 |
109.694 |
|
H7 |
C2 |
H8 |
109.694 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
