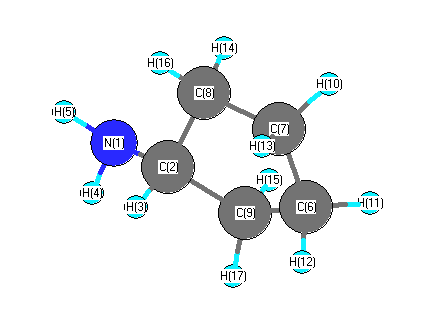
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3475 |
3354 |
2.24 |
|
|
|
| 2 |
A |
3119 |
3010 |
71.71 |
|
|
|
| 3 |
A |
3097 |
2989 |
10.60 |
|
|
|
| 4 |
A |
3067 |
2960 |
37.92 |
|
|
|
| 5 |
A |
3046 |
2940 |
34.46 |
|
|
|
| 6 |
A |
2913 |
2811 |
75.01 |
|
|
|
| 7 |
A |
1639 |
1582 |
21.76 |
|
|
|
| 8 |
A |
1494 |
1442 |
2.98 |
|
|
|
| 9 |
A |
1468 |
1416 |
3.77 |
|
|
|
| 10 |
A |
1398 |
1349 |
19.50 |
|
|
|
| 11 |
A |
1322 |
1276 |
1.20 |
|
|
|
| 12 |
A |
1296 |
1251 |
0.44 |
|
|
|
| 13 |
A |
1234 |
1191 |
1.11 |
|
|
|
| 14 |
A |
1193 |
1151 |
3.05 |
|
|
|
| 15 |
A |
1132 |
1092 |
9.50 |
|
|
|
| 16 |
A |
1066 |
1029 |
0.10 |
|
|
|
| 17 |
A |
986 |
952 |
16.63 |
|
|
|
| 18 |
A |
916 |
884 |
26.66 |
|
|
|
| 19 |
A |
899 |
868 |
23.13 |
|
|
|
| 20 |
A |
877 |
847 |
54.57 |
|
|
|
| 21 |
A |
755 |
729 |
1.54 |
|
|
|
| 22 |
A |
541 |
522 |
2.42 |
|
|
|
| 23 |
A |
462 |
446 |
8.98 |
|
|
|
| 24 |
A |
184 |
178 |
0.95 |
|
|
|
| 25 |
A |
3552 |
3428 |
0.00 |
|
|
|
| 26 |
A |
3102 |
2994 |
9.09 |
|
|
|
| 27 |
A |
3089 |
2980 |
35.83 |
|
|
|
| 28 |
A |
3053 |
2946 |
32.45 |
|
|
|
| 29 |
A |
3043 |
2936 |
25.45 |
|
|
|
| 30 |
A |
1465 |
1414 |
3.28 |
|
|
|
| 31 |
A |
1452 |
1401 |
0.03 |
|
|
|
| 32 |
A |
1387 |
1339 |
1.86 |
|
|
|
| 33 |
A |
1321 |
1275 |
0.05 |
|
|
|
| 34 |
A |
1285 |
1240 |
0.37 |
|
|
|
| 35 |
A |
1272 |
1227 |
1.41 |
|
|
|
| 36 |
A |
1219 |
1176 |
0.07 |
|
|
|
| 37 |
A |
1189 |
1147 |
0.14 |
|
|
|
| 38 |
A |
1046 |
1009 |
0.05 |
|
|
|
| 39 |
A |
976 |
942 |
0.16 |
|
|
|
| 40 |
A |
957 |
924 |
3.07 |
|
|
|
| 41 |
A |
818 |
789 |
0.00 |
|
|
|
| 42 |
A |
633 |
611 |
0.02 |
|
|
|
| 43 |
A |
356 |
343 |
11.60 |
|
|
|
| 44 |
A |
249 |
240 |
28.03 |
|
|
|
| 45 |
A |
35 |
34 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34538.7 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 33329.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.172 |
|
|
|
| 2 |
C |
-0.093 |
|
|
|
| 3 |
H |
0.002 |
|
|
|
| 4 |
H |
0.084 |
|
|
|
| 5 |
H |
0.084 |
|
|
|
| 6 |
C |
-0.059 |
|
|
|
| 7 |
C |
-0.059 |
|
|
|
| 8 |
C |
-0.011 |
|
|
|
| 9 |
C |
-0.011 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
| 11 |
H |
0.031 |
|
|
|
| 12 |
H |
0.032 |
|
|
|
| 13 |
H |
0.032 |
|
|
|
| 14 |
H |
0.035 |
|
|
|
| 15 |
H |
0.035 |
|
|
|
| 16 |
H |
0.019 |
|
|
|
| 17 |
H |
0.019 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.964 |
0.753 |
0.000 |
1.223 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.554 |
1.291 |
0.000 |
| y |
1.291 |
-35.031 |
0.000 |
| z |
0.000 |
0.000 |
-38.023 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.027 |
1.291 |
0.000 |
| y |
1.291 |
4.758 |
0.000 |
| z |
0.000 |
0.000 |
0.269 |
|
| Polar |
|---|
| 3z2-r2 | 0.539 |
|---|
| x2-y2 | -6.523 |
|---|
| xy | 1.291 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
171.355 |
| (<r2>)1/2 |
13.090 |