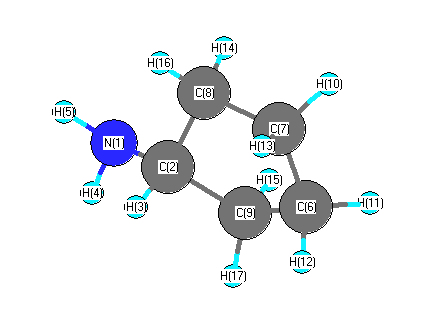
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3357 |
3337 |
5.64 |
|
|
|
| 2 |
A |
3043 |
3025 |
67.50 |
|
|
|
| 3 |
A |
3021 |
3004 |
10.66 |
|
|
|
| 4 |
A |
2991 |
2974 |
39.58 |
|
|
|
| 5 |
A |
2970 |
2953 |
32.11 |
|
|
|
| 6 |
A |
2825 |
2809 |
77.78 |
|
|
|
| 7 |
A |
1592 |
1583 |
17.90 |
|
|
|
| 8 |
A |
1445 |
1436 |
2.64 |
|
|
|
| 9 |
A |
1420 |
1412 |
3.29 |
|
|
|
| 10 |
A |
1348 |
1340 |
16.98 |
|
|
|
| 11 |
A |
1277 |
1269 |
1.77 |
|
|
|
| 12 |
A |
1251 |
1244 |
0.96 |
|
|
|
| 13 |
A |
1193 |
1186 |
0.86 |
|
|
|
| 14 |
A |
1151 |
1144 |
2.46 |
|
|
|
| 15 |
A |
1095 |
1089 |
6.55 |
|
|
|
| 16 |
A |
1037 |
1031 |
0.05 |
|
|
|
| 17 |
A |
957 |
952 |
17.27 |
|
|
|
| 18 |
A |
893 |
888 |
30.15 |
|
|
|
| 19 |
A |
876 |
871 |
27.33 |
|
|
|
| 20 |
A |
859 |
854 |
39.48 |
|
|
|
| 21 |
A |
733 |
729 |
1.76 |
|
|
|
| 22 |
A |
527 |
524 |
2.19 |
|
|
|
| 23 |
A |
455 |
453 |
8.55 |
|
|
|
| 24 |
A |
181 |
180 |
0.83 |
|
|
|
| 25 |
A |
3432 |
3413 |
0.56 |
|
|
|
| 26 |
A |
3027 |
3009 |
8.35 |
|
|
|
| 27 |
A |
3013 |
2996 |
39.26 |
|
|
|
| 28 |
A |
2978 |
2961 |
24.01 |
|
|
|
| 29 |
A |
2966 |
2949 |
27.53 |
|
|
|
| 30 |
A |
1415 |
1407 |
2.52 |
|
|
|
| 31 |
A |
1404 |
1396 |
0.01 |
|
|
|
| 32 |
A |
1341 |
1333 |
2.03 |
|
|
|
| 33 |
A |
1275 |
1268 |
0.01 |
|
|
|
| 34 |
A |
1241 |
1234 |
0.36 |
|
|
|
| 35 |
A |
1234 |
1227 |
2.12 |
|
|
|
| 36 |
A |
1181 |
1174 |
0.04 |
|
|
|
| 37 |
A |
1149 |
1142 |
0.11 |
|
|
|
| 38 |
A |
1019 |
1013 |
0.05 |
|
|
|
| 39 |
A |
946 |
941 |
0.26 |
|
|
|
| 40 |
A |
933 |
927 |
2.81 |
|
|
|
| 41 |
A |
794 |
790 |
0.00 |
|
|
|
| 42 |
A |
616 |
612 |
0.03 |
|
|
|
| 43 |
A |
349 |
347 |
10.77 |
|
|
|
| 44 |
A |
247 |
246 |
25.90 |
|
|
|
| 45 |
A |
3 |
3 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33528.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 33334.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.132 |
|
|
|
| 2 |
C |
-0.089 |
|
|
|
| 3 |
H |
-0.014 |
|
|
|
| 4 |
H |
0.073 |
|
|
|
| 5 |
H |
0.073 |
|
|
|
| 6 |
C |
-0.033 |
|
|
|
| 7 |
C |
-0.033 |
|
|
|
| 8 |
C |
0.016 |
|
|
|
| 9 |
C |
0.016 |
|
|
|
| 10 |
H |
0.016 |
|
|
|
| 11 |
H |
0.016 |
|
|
|
| 12 |
H |
0.020 |
|
|
|
| 13 |
H |
0.020 |
|
|
|
| 14 |
H |
0.021 |
|
|
|
| 15 |
H |
0.021 |
|
|
|
| 16 |
H |
0.004 |
|
|
|
| 17 |
H |
0.004 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.197 |
0.170 |
0.000 |
1.209 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.996 |
3.459 |
0.000 |
| y |
3.459 |
-38.262 |
0.000 |
| z |
0.000 |
0.000 |
-38.464 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.634 |
3.459 |
0.000 |
| y |
3.459 |
0.468 |
0.000 |
| z |
0.000 |
0.000 |
0.165 |
|
| Polar |
|---|
| 3z2-r2 | 0.331 |
|---|
| x2-y2 | -0.735 |
|---|
| xy | 3.459 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.318 |
0.355 |
0.000 |
| y |
0.355 |
10.285 |
0.000 |
| z |
0.000 |
0.000 |
9.531 |
<r2> (average value of r
2) Å
2
| <r2> |
172.984 |
| (<r2>)1/2 |
13.152 |