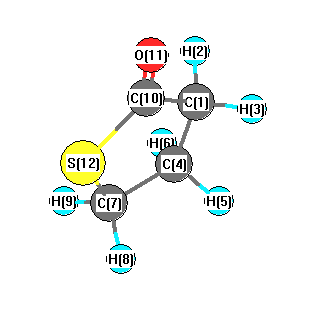
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3060 |
3043 |
10.06 |
|
|
|
| 2 |
A |
3055 |
3037 |
6.49 |
|
|
|
| 3 |
A |
3038 |
3020 |
14.71 |
|
|
|
| 4 |
A |
2989 |
2972 |
20.18 |
|
|
|
| 5 |
A |
2986 |
2968 |
22.08 |
|
|
|
| 6 |
A |
2966 |
2949 |
6.99 |
|
|
|
| 7 |
A |
1776 |
1766 |
325.94 |
|
|
|
| 8 |
A |
1417 |
1409 |
0.80 |
|
|
|
| 9 |
A |
1401 |
1393 |
5.82 |
|
|
|
| 10 |
A |
1375 |
1367 |
6.86 |
|
|
|
| 11 |
A |
1303 |
1296 |
0.72 |
|
|
|
| 12 |
A |
1253 |
1246 |
5.42 |
|
|
|
| 13 |
A |
1239 |
1232 |
5.74 |
|
|
|
| 14 |
A |
1196 |
1189 |
1.99 |
|
|
|
| 15 |
A |
1138 |
1131 |
0.73 |
|
|
|
| 16 |
A |
1066 |
1060 |
10.52 |
|
|
|
| 17 |
A |
1036 |
1030 |
26.24 |
|
|
|
| 18 |
A |
996 |
990 |
63.93 |
|
|
|
| 19 |
A |
982 |
976 |
4.26 |
|
|
|
| 20 |
A |
907 |
901 |
8.07 |
|
|
|
| 21 |
A |
833 |
828 |
8.38 |
|
|
|
| 22 |
A |
808 |
803 |
13.52 |
|
|
|
| 23 |
A |
673 |
669 |
2.18 |
|
|
|
| 24 |
A |
604 |
600 |
31.13 |
|
|
|
| 25 |
A |
576 |
573 |
14.78 |
|
|
|
| 26 |
A |
463 |
461 |
1.58 |
|
|
|
| 27 |
A |
457 |
455 |
1.17 |
|
|
|
| 28 |
A |
381 |
379 |
3.43 |
|
|
|
| 29 |
A |
226 |
225 |
1.16 |
|
|
|
| 30 |
A |
130 |
129 |
0.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20164.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20047.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.005 |
|
|
|
| 2 |
H |
0.050 |
|
|
|
| 3 |
H |
0.065 |
|
|
|
| 4 |
C |
-0.039 |
|
|
|
| 5 |
H |
0.034 |
|
|
|
| 6 |
H |
0.049 |
|
|
|
| 7 |
C |
-0.132 |
|
|
|
| 8 |
H |
0.071 |
|
|
|
| 9 |
H |
0.066 |
|
|
|
| 10 |
C |
-0.004 |
|
|
|
| 11 |
O |
-0.163 |
|
|
|
| 12 |
S |
0.007 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.311 |
1.680 |
0.349 |
3.729 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.136 |
-0.278 |
-0.049 |
| y |
-0.278 |
-42.324 |
0.186 |
| z |
-0.049 |
0.186 |
-42.151 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.899 |
-0.278 |
-0.049 |
| y |
-0.278 |
1.820 |
0.186 |
| z |
-0.049 |
0.186 |
2.079 |
|
| Polar |
|---|
| 3z2-r2 | 4.158 |
|---|
| x2-y2 | -3.812 |
|---|
| xy | -0.278 |
|---|
| xz | -0.049 |
|---|
| yz | 0.186 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.742 |
-0.500 |
0.148 |
| y |
-0.500 |
10.075 |
0.164 |
| z |
0.148 |
0.164 |
6.387 |
<r2> (average value of r
2) Å
2
| <r2> |
180.954 |
| (<r2>)1/2 |
13.452 |