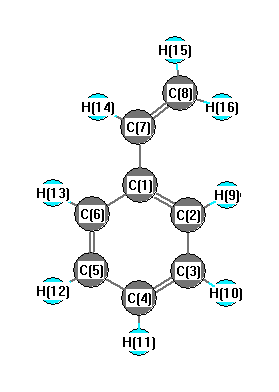
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3178 |
3160 |
9.99 |
|
|
|
| 2 |
A' |
3137 |
3119 |
13.72 |
|
|
|
| 3 |
A' |
3127 |
3109 |
27.68 |
|
|
|
| 4 |
A' |
3119 |
3101 |
7.94 |
|
|
|
| 5 |
A' |
3108 |
3090 |
0.39 |
|
|
|
| 6 |
A' |
3100 |
3082 |
8.64 |
|
|
|
| 7 |
A' |
3085 |
3067 |
1.96 |
|
|
|
| 8 |
A' |
3073 |
3055 |
15.46 |
|
|
|
| 9 |
A' |
1650 |
1640 |
9.87 |
|
|
|
| 10 |
A' |
1612 |
1602 |
0.82 |
|
|
|
| 11 |
A' |
1584 |
1575 |
2.06 |
|
|
|
| 12 |
A' |
1479 |
1470 |
8.24 |
|
|
|
| 13 |
A' |
1433 |
1425 |
5.40 |
|
|
|
| 14 |
A' |
1390 |
1382 |
3.34 |
|
|
|
| 15 |
A' |
1367 |
1359 |
2.05 |
|
|
|
| 16 |
A' |
1297 |
1289 |
2.60 |
|
|
|
| 17 |
A' |
1272 |
1265 |
1.50 |
|
|
|
| 18 |
A' |
1200 |
1193 |
2.32 |
|
|
|
| 19 |
A' |
1157 |
1151 |
0.11 |
|
|
|
| 20 |
A' |
1136 |
1130 |
0.09 |
|
|
|
| 21 |
A' |
1078 |
1072 |
5.41 |
|
|
|
| 22 |
A' |
1029 |
1023 |
1.20 |
|
|
|
| 23 |
A' |
1003 |
998 |
5.72 |
|
|
|
| 24 |
A' |
982 |
977 |
0.05 |
|
|
|
| 25 |
A' |
769 |
765 |
0.24 |
|
|
|
| 26 |
A' |
610 |
606 |
0.03 |
|
|
|
| 27 |
A' |
541 |
537 |
4.35 |
|
|
|
| 28 |
A' |
435 |
433 |
0.10 |
|
|
|
| 29 |
A' |
225 |
223 |
0.49 |
|
|
|
| 30 |
A" |
995 |
989 |
11.66 |
|
|
|
| 31 |
A" |
969 |
963 |
0.45 |
|
|
|
| 32 |
A" |
947 |
942 |
0.00 |
|
|
|
| 33 |
A" |
901 |
896 |
1.41 |
|
|
|
| 34 |
A" |
879 |
874 |
29.94 |
|
|
|
| 35 |
A" |
825 |
820 |
0.27 |
|
|
|
| 36 |
A" |
779 |
775 |
20.72 |
|
|
|
| 37 |
A" |
689 |
685 |
31.12 |
|
|
|
| 38 |
A" |
637 |
633 |
0.08 |
|
|
|
| 39 |
A" |
433 |
430 |
6.25 |
|
|
|
| 40 |
A" |
397 |
395 |
0.18 |
|
|
|
| 41 |
A" |
202 |
200 |
1.38 |
|
|
|
| 42 |
A" |
55 |
54 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28442.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28277.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.029 |
|
|
|
| 2 |
C |
-0.005 |
|
|
|
| 3 |
C |
0.034 |
|
|
|
| 4 |
C |
0.024 |
|
|
|
| 5 |
C |
0.029 |
|
|
|
| 6 |
C |
0.005 |
|
|
|
| 7 |
C |
-0.036 |
|
|
|
| 8 |
C |
0.010 |
|
|
|
| 9 |
H |
-0.031 |
|
|
|
| 10 |
H |
-0.019 |
|
|
|
| 11 |
H |
-0.020 |
|
|
|
| 12 |
H |
-0.019 |
|
|
|
| 13 |
H |
-0.031 |
|
|
|
| 14 |
H |
-0.021 |
|
|
|
| 15 |
H |
0.029 |
|
|
|
| 16 |
H |
0.023 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.068 |
-0.170 |
0.000 |
0.184 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.603 |
0.115 |
0.000 |
| y |
0.115 |
-42.745 |
0.000 |
| z |
0.000 |
0.000 |
-52.117 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.828 |
0.115 |
0.000 |
| y |
0.115 |
4.615 |
0.000 |
| z |
0.000 |
0.000 |
-9.443 |
|
| Polar |
|---|
| 3z2-r2 | -18.885 |
|---|
| x2-y2 | 0.142 |
|---|
| xy | 0.115 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.467 |
-2.917 |
0.000 |
| y |
-2.917 |
19.213 |
0.000 |
| z |
0.000 |
0.000 |
4.836 |
<r2> (average value of r
2) Å
2
| <r2> |
280.602 |
| (<r2>)1/2 |
16.751 |