return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-344.637780
Energy at 298.15K 
HF Energy-343.455198
Nuclear repulsion energy317.797985
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3215 3065        
2 A' 3208 3057        
3 A' 3197 3047        
4 A' 3184 3035        
5 A' 3173 3024        
6 A' 2914 2777        
7 A' 1769 1686        
8 A' 1648 1571        
9 A' 1632 1555        
10 A' 1513 1442        
11 A' 1473 1404        
12 A' 1410 1344        
13 A' 1349 1286        
14 A' 1315 1254        
15 A' 1222 1165        
16 A' 1171 1116        
17 A' 1162 1108        
18 A' 1085 1034        
19 A' 1031 983        
20 A' 997 950        
21 A' 829 790        
22 A' 647 617        
23 A' 612 584        
24 A' 433 412        
25 A' 217 207        
26 A" 1009 962        
27 A" 958 914        
28 A" 949 905        
29 A" 908 866        
30 A" 850 810        
31 A" 734 699        
32 A" 625 596        
33 A" 449 428        
34 A" 401 382        
35 A" 236 225        
36 A" 121 115        

Unscaled Zero Point Vibrational Energy (zpe) 23823.6 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 22706.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.17106 0.05126 0.03944

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.581 0.000
C2 -1.044 -0.373 0.000
C3 -0.735 -1.745 0.000
C4 0.615 -2.165 0.000
C5 1.656 -1.214 0.000
C6 1.347 0.161 0.000
C7 -0.328 2.042 0.000
O8 -1.465 2.491 0.000
H9 0.567 2.722 0.000
H10 -2.082 -0.018 0.000
H11 -1.541 -2.490 0.000
H12 0.854 -3.237 0.000
H13 2.703 -1.544 0.000
H14 2.151 0.911 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.41472.43952.81462.44321.41131.49712.40672.21392.16633.43663.91253.43892.1760
C21.41471.40572.44192.82852.45032.51912.89513.48881.09692.17473.43533.92593.4435
C32.43951.40571.41422.45002.82313.80854.29814.65212.18961.09762.17993.44453.9224
C42.81462.44191.41421.41042.43904.31175.09954.88703.44702.17991.09792.17883.4387
C52.44322.82852.45001.41041.41003.81374.84494.08413.92513.44252.17581.09742.1825
C61.41132.45032.82312.43901.41002.51903.65182.67683.43383.92073.43362.17881.0993
C71.49712.51913.80854.31173.81372.51901.22181.12362.70524.69195.40964.69602.7249
O82.40672.89514.29815.09954.84493.65181.22182.04422.58384.98226.17935.80133.9456
H92.21393.48884.65214.88704.08412.67681.12362.04423.81035.62195.96524.77112.4056
H102.16631.09692.18963.44703.92513.43382.70522.58383.81032.53104.35645.02254.3335
H113.43662.17471.09762.17993.44253.92074.69194.98225.62192.53102.50804.34815.0200
H123.91253.43532.17991.09792.17583.43365.40966.17935.96524.35642.50802.50704.3462
H133.43893.92593.44452.17881.09742.17884.69605.80134.77115.02254.34812.50702.5170
H142.17603.44353.92243.43872.18251.09932.72493.94562.40564.33355.02004.34622.5170

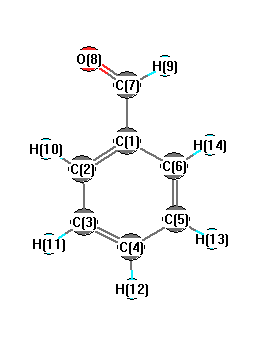
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.759 C1 C2 H10 118.660
C1 C6 C5 119.989 C1 C6 H14 119.647
C1 C7 O8 124.229 C1 C7 H9 114.544
C2 C1 C6 120.238 C2 C1 C7 119.774
C2 C3 C4 119.982 C2 C3 H11 120.123
C3 C2 H10 121.581 C3 C4 C5 120.314
C3 C4 H12 119.872 C4 C3 H11 119.895
C4 C5 C6 119.717 C4 C5 H13 120.123
C5 C4 H12 119.814 C5 C6 H14 120.363
C6 C1 C7 119.988 C6 C5 H13 120.160
O8 C7 H9 121.227
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability