return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-344.590012
Energy at 298.15K 
HF Energy-343.457078
Nuclear repulsion energy318.642873
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3238 3106 6.45      
2 A' 3230 3099 13.57      
3 A' 3218 3087 14.40      
4 A' 3205 3075 1.67      
5 A' 3195 3065 4.45      
6 A' 2945 2825 125.06      
7 A' 1805 1731 185.27      
8 A' 1680 1612 12.70      
9 A' 1663 1595 7.98      
10 A' 1537 1474 0.33      
11 A' 1494 1434 9.83      
12 A' 1426 1368 12.29      
13 A' 1338 1284 8.20      
14 A' 1321 1267 11.84      
15 A' 1237 1187 45.20      
16 A' 1186 1138 9.00      
17 A' 1171 1123 0.88      
18 A' 1100 1055 3.82      
19 A' 1048 1006 1.21      
20 A' 1012 971 0.10      
21 A' 840 805 31.45      
22 A' 656 629 19.75      
23 A' 621 596 0.76      
24 A' 439 421 0.15      
25 A' 221 212 7.31      
26 A" 1029 988 0.01      
27 A" 988 948 0.04      
28 A" 979 939 0.06      
29 A" 933 895 1.06      
30 A" 870 835 0.04      
31 A" 752 721 57.98      
32 A" 654 627 6.55      
33 A" 461 442 6.25      
34 A" 411 394 0.04      
35 A" 237 228 7.09      
36 A" 120 115 3.65      

Unscaled Zero Point Vibrational Energy (zpe) 24127.5 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 23148.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.17224 0.05146 0.03962

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.579 0.000
C2 -1.041 -0.373 0.000
C3 -0.734 -1.740 0.000
C4 0.611 -2.160 0.000
C5 1.650 -1.214 0.000
C6 1.342 0.158 0.000
C7 -0.326 2.040 0.000
O8 -1.457 2.489 0.000
H9 0.568 2.716 0.000
H10 -2.077 -0.019 0.000
H11 -1.539 -2.483 0.000
H12 0.849 -3.230 0.000
H13 2.695 -1.543 0.000
H14 2.146 0.905 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.41002.43222.80672.43621.40651.49662.40252.21172.16153.42723.90273.42982.1705
C21.41001.40132.43422.81882.44102.51582.89203.48271.09492.16873.42593.91443.4331
C32.43221.40131.40962.44142.81303.80154.29064.64272.18261.09572.17413.43443.9103
C42.80672.43421.40961.40542.43104.30335.08904.87713.43672.17451.09602.17293.4282
C52.43622.81882.44141.40541.40583.80614.83364.07623.91353.43242.17011.09552.1760
C61.40652.44102.81302.43101.40582.51443.64262.67293.42373.90873.42432.17341.0974
C71.49662.51583.80154.30333.80612.51441.21711.12122.70294.68275.39934.68612.7197
O82.40252.89204.29065.08904.83363.64261.21712.03742.58424.97326.16705.78753.9357
H92.21173.48274.64274.87714.07622.67291.12122.03743.80535.61035.95354.76102.4023
H102.16151.09492.18263.43673.91353.42372.70292.58423.80532.52194.34415.00904.3229
H113.42722.16871.09572.17453.43243.90874.68274.97325.61032.52192.50204.33695.0060
H123.90273.42592.17411.09602.17013.42435.39936.16705.95354.34412.50202.50094.3342
H133.42983.91443.43442.17291.09552.17344.68615.78754.76105.00904.33692.50092.5090
H142.17053.43313.91033.42822.17601.09742.71973.93572.40234.32295.00604.33422.5090

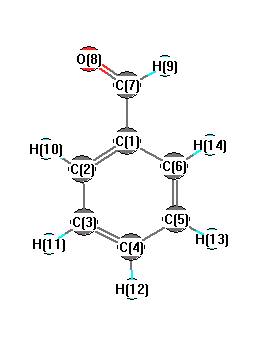
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.803 C1 C2 H10 118.745
C1 C6 C5 120.055 C1 C6 H14 119.691
C1 C7 O8 124.266 C1 C7 H9 114.558
C2 C1 C6 120.155 C2 C1 C7 119.859
C2 C3 C4 119.991 C2 C3 H11 120.081
C3 C2 H10 121.452 C3 C4 C5 120.288
C3 C4 H12 119.868 C4 C3 H11 119.928
C4 C5 C6 119.707 C4 C5 H13 120.141
C5 C4 H12 119.843 C5 C6 H14 120.254
C6 C1 C7 119.986 C6 C5 H13 120.152
O8 C7 H9 121.176
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability