return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-344.577586
Energy at 298.15K 
HF Energy-343.458019
Nuclear repulsion energy319.044781
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3246 3106 5.62      
2 A' 3238 3098 12.66      
3 A' 3226 3087 14.24      
4 A' 3214 3075 1.67      
5 A' 3204 3065 4.06      
6 A' 2965 2837 115.06      
7 A' 1857 1776 167.58      
8 A' 1692 1619 8.24      
9 A' 1674 1601 5.37      
10 A' 1545 1479 0.37      
11 A' 1502 1437 8.74      
12 A' 1438 1375 15.21      
13 A' 1347 1288 4.61      
14 A' 1333 1275 14.34      
15 A' 1247 1193 42.73      
16 A' 1192 1140 5.41      
17 A' 1176 1125 0.67      
18 A' 1105 1058 3.62      
19 A' 1054 1009 1.59      
20 A' 1018 974 0.19      
21 A' 844 807 32.42      
22 A' 660 631 20.52      
23 A' 624 597 0.66      
24 A' 441 422 0.12      
25 A' 223 213 7.27      
26 A" 1035 990 0.09      
27 A" 990 947 0.05      
28 A" 981 938 0.03      
29 A" 932 891 1.14      
30 A" 872 835 0.03      
31 A" 753 720 60.37      
32 A" 654 626 4.54      
33 A" 463 443 6.43      
34 A" 412 394 0.04      
35 A" 236 226 7.06      
36 A" 119 114 3.88      

Unscaled Zero Point Vibrational Energy (zpe) 24254.1 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23203.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.17275 0.05157 0.03971

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.578 0.000
C2 -1.042 -0.370 0.000
C3 -0.737 -1.737 0.000
C4 0.605 -2.161 0.000
C5 1.643 -1.216 0.000
C6 1.340 0.155 0.000
C7 -0.321 2.040 0.000
O8 -1.445 2.491 0.000
H9 0.573 2.715 0.000
H10 -2.077 -0.017 0.000
H11 -1.544 -2.478 0.000
H12 0.840 -3.231 0.000
H13 2.687 -1.548 0.000
H14 2.145 0.899 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.40832.42902.80462.43321.40501.49682.39722.21232.16063.42343.90023.42642.1692
C21.40831.39992.43262.81532.43842.51552.88923.48171.09442.16663.42403.91053.4302
C32.42901.39991.40792.43702.80913.79944.28624.64012.18021.09542.17283.43003.9059
C42.80462.43261.40791.40362.42924.30155.08324.87543.43402.17241.09562.17073.4256
C52.43322.81532.43701.40361.40443.80284.82504.07413.90953.42802.16881.09522.1742
C61.40502.43842.80912.42921.40442.51213.63462.67223.42133.90453.42222.17131.0968
C71.49682.51553.79944.30153.80282.51211.21111.11982.70474.68035.39704.68182.7170
O82.39722.88924.28625.08324.82503.63461.21112.03022.58614.96956.16105.77793.9270
H92.21233.48174.64014.87544.07412.67221.11982.03023.80595.60735.95144.75792.4016
H102.16061.09442.18023.43403.90953.42132.70472.58613.80592.51814.34075.00474.3207
H113.42342.16661.09542.17243.42803.90454.68034.96955.60732.51812.50024.33245.0013
H123.90023.42402.17281.09562.16883.42225.39706.16105.95144.34072.50022.49914.3313
H133.42643.91053.43002.17071.09522.17134.68185.77794.75795.00474.33242.49912.5062
H142.16923.43023.90593.42562.17421.09682.71703.92702.40164.32075.00134.33132.5062

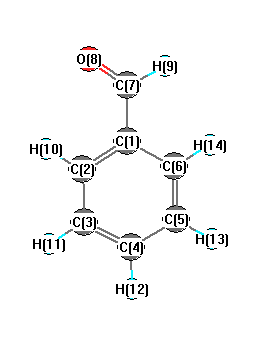
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.760 C1 C2 H10 118.852
C1 C6 C5 120.021 C1 C6 H14 119.732
C1 C7 O8 124.227 C1 C7 H9 114.679
C2 C1 C6 120.165 C2 C1 C7 119.939
C2 C3 C4 120.083 C2 C3 H11 120.024
C3 C2 H10 121.388 C3 C4 C5 120.182
C3 C4 H12 119.917 C4 C3 H11 119.892
C4 C5 C6 119.789 C4 C5 H13 120.108
C5 C4 H12 119.901 C5 C6 H14 120.247
C6 C1 C7 119.895 C6 C5 H13 120.103
O8 C7 H9 121.094
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability