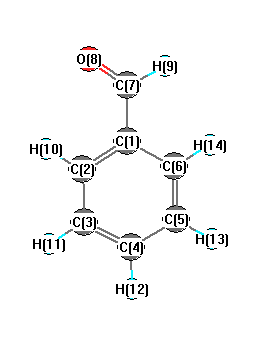
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3138 |
3120 |
12.25 |
320.64 |
0.11 |
0.19 |
| 2 |
A' |
3133 |
3114 |
11.00 |
37.85 |
0.31 |
0.47 |
| 3 |
A' |
3123 |
3105 |
9.60 |
103.90 |
0.74 |
0.85 |
| 4 |
A' |
3112 |
3094 |
0.65 |
87.82 |
0.69 |
0.82 |
| 5 |
A' |
3098 |
3080 |
4.62 |
53.44 |
0.39 |
0.56 |
| 6 |
A' |
2775 |
2759 |
151.95 |
160.65 |
0.30 |
0.46 |
| 7 |
A' |
1734 |
1724 |
209.26 |
68.01 |
0.37 |
0.54 |
| 8 |
A' |
1608 |
1599 |
27.79 |
76.19 |
0.52 |
0.68 |
| 9 |
A' |
1592 |
1583 |
12.37 |
9.65 |
0.50 |
0.67 |
| 10 |
A' |
1469 |
1461 |
1.99 |
0.79 |
0.30 |
0.46 |
| 11 |
A' |
1437 |
1428 |
13.73 |
1.55 |
0.31 |
0.48 |
| 12 |
A' |
1386 |
1378 |
3.57 |
2.03 |
0.65 |
0.79 |
| 13 |
A' |
1351 |
1344 |
7.81 |
2.16 |
0.68 |
0.81 |
| 14 |
A' |
1275 |
1268 |
14.41 |
1.59 |
0.48 |
0.64 |
| 15 |
A' |
1197 |
1190 |
57.07 |
21.40 |
0.25 |
0.40 |
| 16 |
A' |
1140 |
1133 |
6.97 |
5.20 |
0.41 |
0.58 |
| 17 |
A' |
1137 |
1130 |
9.85 |
9.32 |
0.52 |
0.68 |
| 18 |
A' |
1064 |
1058 |
7.89 |
0.99 |
0.21 |
0.34 |
| 19 |
A' |
1017 |
1011 |
3.10 |
15.35 |
0.10 |
0.18 |
| 20 |
A' |
985 |
980 |
0.30 |
19.98 |
0.09 |
0.16 |
| 21 |
A' |
820 |
815 |
27.07 |
10.60 |
0.13 |
0.23 |
| 22 |
A' |
640 |
636 |
18.98 |
3.20 |
0.37 |
0.54 |
| 23 |
A' |
605 |
601 |
0.44 |
5.41 |
0.75 |
0.86 |
| 24 |
A' |
429 |
426 |
0.15 |
6.88 |
0.36 |
0.53 |
| 25 |
A' |
210 |
209 |
6.67 |
0.47 |
0.52 |
0.69 |
| 26 |
A" |
999 |
994 |
0.26 |
5.59 |
0.75 |
0.86 |
| 27 |
A" |
981 |
976 |
0.01 |
0.68 |
0.75 |
0.86 |
| 28 |
A" |
963 |
957 |
0.03 |
0.15 |
0.75 |
0.86 |
| 29 |
A" |
913 |
907 |
0.88 |
0.87 |
0.75 |
0.86 |
| 30 |
A" |
840 |
835 |
0.03 |
5.46 |
0.75 |
0.86 |
| 31 |
A" |
742 |
738 |
28.25 |
1.68 |
0.75 |
0.86 |
| 32 |
A" |
687 |
683 |
24.41 |
0.29 |
0.75 |
0.86 |
| 33 |
A" |
455 |
453 |
4.92 |
0.54 |
0.75 |
0.86 |
| 34 |
A" |
403 |
401 |
0.04 |
0.00 |
0.75 |
0.86 |
| 35 |
A" |
237 |
236 |
5.15 |
1.13 |
0.75 |
0.86 |
| 36 |
A" |
123 |
122 |
2.29 |
2.99 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 23409.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23273.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.083 |
|
|
|
| 2 |
C |
0.055 |
|
|
|
| 3 |
C |
0.033 |
|
|
|
| 4 |
C |
0.032 |
|
|
|
| 5 |
C |
0.030 |
|
|
|
| 6 |
C |
0.021 |
|
|
|
| 7 |
C |
0.174 |
|
|
|
| 8 |
O |
-0.172 |
|
|
|
| 9 |
H |
-0.022 |
|
|
|
| 10 |
H |
-0.017 |
|
|
|
| 11 |
H |
-0.008 |
|
|
|
| 12 |
H |
-0.009 |
|
|
|
| 13 |
H |
-0.011 |
|
|
|
| 14 |
H |
-0.023 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.999 |
-2.295 |
0.000 |
3.044 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.283 |
5.209 |
0.000 |
| y |
5.209 |
-46.250 |
0.000 |
| z |
0.000 |
0.000 |
-48.436 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.060 |
5.209 |
0.000 |
| y |
5.209 |
0.110 |
0.000 |
| z |
0.000 |
0.000 |
-3.170 |
|
| Polar |
|---|
| 3z2-r2 | -6.340 |
|---|
| x2-y2 | 1.967 |
|---|
| xy | 5.209 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.444 |
-1.081 |
0.000 |
| y |
-1.081 |
16.116 |
0.000 |
| z |
0.000 |
0.000 |
4.418 |
<r2> (average value of r
2) Å
2
| <r2> |
264.663 |
| (<r2>)1/2 |
16.268 |