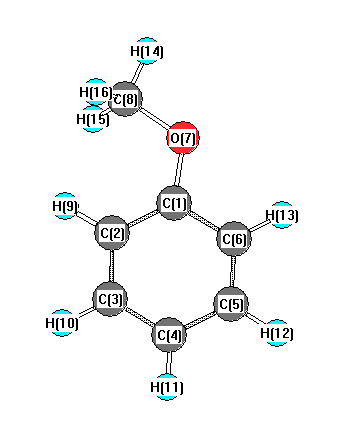
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3147 |
3129 |
4.29 |
|
|
|
| 2 |
A' |
3140 |
3122 |
13.77 |
|
|
|
| 3 |
A' |
3133 |
3115 |
17.50 |
|
|
|
| 4 |
A' |
3116 |
3098 |
10.44 |
|
|
|
| 5 |
A' |
3109 |
3091 |
2.82 |
|
|
|
| 6 |
A' |
3077 |
3059 |
17.41 |
|
|
|
| 7 |
A' |
2926 |
2909 |
59.71 |
|
|
|
| 8 |
A' |
1617 |
1607 |
69.02 |
|
|
|
| 9 |
A' |
1595 |
1585 |
17.63 |
|
|
|
| 10 |
A' |
1482 |
1474 |
91.63 |
|
|
|
| 11 |
A' |
1444 |
1435 |
2.52 |
|
|
|
| 12 |
A' |
1423 |
1415 |
36.75 |
|
|
|
| 13 |
A' |
1404 |
1396 |
6.38 |
|
|
|
| 14 |
A' |
1378 |
1370 |
12.40 |
|
|
|
| 15 |
A' |
1280 |
1272 |
0.17 |
|
|
|
| 16 |
A' |
1253 |
1246 |
205.71 |
|
|
|
| 17 |
A' |
1155 |
1148 |
2.20 |
|
|
|
| 18 |
A' |
1147 |
1140 |
13.19 |
|
|
|
| 19 |
A' |
1131 |
1124 |
1.91 |
|
|
|
| 20 |
A' |
1069 |
1063 |
11.39 |
|
|
|
| 21 |
A' |
1049 |
1043 |
48.23 |
|
|
|
| 22 |
A' |
1020 |
1014 |
1.42 |
|
|
|
| 23 |
A' |
977 |
971 |
0.24 |
|
|
|
| 24 |
A' |
781 |
776 |
13.04 |
|
|
|
| 25 |
A' |
603 |
600 |
0.42 |
|
|
|
| 26 |
A' |
543 |
540 |
4.78 |
|
|
|
| 27 |
A' |
435 |
432 |
1.00 |
|
|
|
| 28 |
A' |
251 |
249 |
1.90 |
|
|
|
| 29 |
A" |
2994 |
2977 |
39.12 |
|
|
|
| 30 |
A" |
1408 |
1400 |
6.12 |
|
|
|
| 31 |
A" |
1117 |
1111 |
0.34 |
|
|
|
| 32 |
A" |
957 |
951 |
0.19 |
|
|
|
| 33 |
A" |
934 |
928 |
0.01 |
|
|
|
| 34 |
A" |
864 |
859 |
5.89 |
|
|
|
| 35 |
A" |
798 |
793 |
0.02 |
|
|
|
| 36 |
A" |
744 |
739 |
41.53 |
|
|
|
| 37 |
A" |
684 |
681 |
18.48 |
|
|
|
| 38 |
A" |
504 |
501 |
6.78 |
|
|
|
| 39 |
A" |
410 |
408 |
0.01 |
|
|
|
| 40 |
A" |
271 |
270 |
0.41 |
|
|
|
| 41 |
A" |
205 |
203 |
0.18 |
|
|
|
| 42 |
A" |
100 |
99 |
3.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28335.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28171.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.108 |
|
|
|
| 2 |
C |
-0.037 |
|
|
|
| 3 |
C |
0.021 |
|
|
|
| 4 |
C |
0.015 |
|
|
|
| 5 |
C |
0.030 |
|
|
|
| 6 |
C |
0.015 |
|
|
|
| 7 |
O |
-0.239 |
|
|
|
| 8 |
C |
0.064 |
|
|
|
| 9 |
H |
-0.023 |
|
|
|
| 10 |
H |
-0.021 |
|
|
|
| 11 |
H |
-0.025 |
|
|
|
| 12 |
H |
-0.020 |
|
|
|
| 13 |
H |
-0.020 |
|
|
|
| 14 |
H |
0.048 |
|
|
|
| 15 |
H |
0.042 |
|
|
|
| 16 |
H |
0.042 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.294 |
0.392 |
0.000 |
1.352 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.173 |
3.064 |
0.000 |
| y |
3.064 |
-43.657 |
0.000 |
| z |
0.000 |
0.000 |
-50.627 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.969 |
3.064 |
0.000 |
| y |
3.064 |
2.243 |
0.000 |
| z |
0.000 |
0.000 |
-8.211 |
|
| Polar |
|---|
| 3z2-r2 | -16.423 |
|---|
| x2-y2 | 2.484 |
|---|
| xy | 3.064 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.007 |
1.742 |
0.000 |
| y |
1.742 |
15.367 |
0.000 |
| z |
0.000 |
0.000 |
5.544 |
<r2> (average value of r
2) Å
2
| <r2> |
277.242 |
| (<r2>)1/2 |
16.651 |