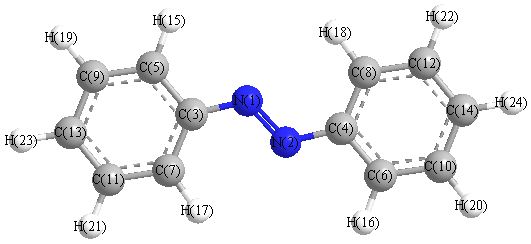
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3147 |
3129 |
0.00 |
|
|
|
| 2 |
Ag |
3139 |
3121 |
0.00 |
|
|
|
| 3 |
Ag |
3129 |
3111 |
0.00 |
|
|
|
| 4 |
Ag |
3119 |
3101 |
0.00 |
|
|
|
| 5 |
Ag |
3108 |
3090 |
0.00 |
|
|
|
| 6 |
Ag |
1609 |
1599 |
0.00 |
|
|
|
| 7 |
Ag |
1592 |
1582 |
0.00 |
|
|
|
| 8 |
Ag |
1486 |
1478 |
0.00 |
|
|
|
| 9 |
Ag |
1450 |
1442 |
0.00 |
|
|
|
| 10 |
Ag |
1425 |
1417 |
0.00 |
|
|
|
| 11 |
Ag |
1382 |
1374 |
0.00 |
|
|
|
| 12 |
Ag |
1277 |
1270 |
0.00 |
|
|
|
| 13 |
Ag |
1177 |
1170 |
0.00 |
|
|
|
| 14 |
Ag |
1134 |
1128 |
0.00 |
|
|
|
| 15 |
Ag |
1122 |
1115 |
0.00 |
|
|
|
| 16 |
Ag |
1057 |
1051 |
0.00 |
|
|
|
| 17 |
Ag |
1015 |
1010 |
0.00 |
|
|
|
| 18 |
Ag |
985 |
979 |
0.00 |
|
|
|
| 19 |
Ag |
914 |
908 |
0.00 |
|
|
|
| 20 |
Ag |
659 |
655 |
0.00 |
|
|
|
| 21 |
Ag |
601 |
597 |
0.00 |
|
|
|
| 22 |
Ag |
299 |
297 |
0.00 |
|
|
|
| 23 |
Ag |
218 |
217 |
0.00 |
|
|
|
| 24 |
Au |
984 |
978 |
0.38 |
|
|
|
| 25 |
Au |
965 |
960 |
0.00 |
|
|
|
| 26 |
Au |
930 |
925 |
5.30 |
|
|
|
| 27 |
Au |
835 |
831 |
0.01 |
|
|
|
| 28 |
Au |
781 |
776 |
38.33 |
|
|
|
| 29 |
Au |
685 |
681 |
59.68 |
|
|
|
| 30 |
Au |
541 |
538 |
9.33 |
|
|
|
| 31 |
Au |
402 |
400 |
0.00 |
|
|
|
| 32 |
Au |
295 |
294 |
0.56 |
|
|
|
| 33 |
Au |
60 |
60 |
1.20 |
|
|
|
| 34 |
Au |
25 |
25 |
0.07 |
|
|
|
| 35 |
Bg |
983 |
978 |
0.00 |
|
|
|
| 36 |
Bg |
966 |
960 |
0.00 |
|
|
|
| 37 |
Bg |
925 |
920 |
0.00 |
|
|
|
| 38 |
Bg |
836 |
832 |
0.00 |
|
|
|
| 39 |
Bg |
761 |
757 |
0.00 |
|
|
|
| 40 |
Bg |
681 |
677 |
0.00 |
|
|
|
| 41 |
Bg |
475 |
472 |
0.00 |
|
|
|
| 42 |
Bg |
406 |
403 |
0.00 |
|
|
|
| 43 |
Bg |
243 |
242 |
0.00 |
|
|
|
| 44 |
Bg |
104 |
104 |
0.00 |
|
|
|
| 45 |
Bu |
3147 |
3129 |
13.73 |
|
|
|
| 46 |
Bu |
3139 |
3120 |
39.13 |
|
|
|
| 47 |
Bu |
3129 |
3111 |
29.32 |
|
|
|
| 48 |
Bu |
3119 |
3101 |
15.79 |
|
|
|
| 49 |
Bu |
3108 |
3090 |
4.98 |
|
|
|
| 50 |
Bu |
1606 |
1597 |
5.49 |
|
|
|
| 51 |
Bu |
1587 |
1578 |
3.53 |
|
|
|
| 52 |
Bu |
1464 |
1456 |
7.27 |
|
|
|
| 53 |
Bu |
1437 |
1428 |
14.07 |
|
|
|
| 54 |
Bu |
1378 |
1370 |
5.59 |
|
|
|
| 55 |
Bu |
1272 |
1265 |
2.09 |
|
|
|
| 56 |
Bu |
1232 |
1225 |
25.69 |
|
|
|
| 57 |
Bu |
1134 |
1128 |
0.33 |
|
|
|
| 58 |
Bu |
1124 |
1118 |
28.85 |
|
|
|
| 59 |
Bu |
1062 |
1056 |
19.56 |
|
|
|
| 60 |
Bu |
1013 |
1008 |
14.77 |
|
|
|
| 61 |
Bu |
983 |
978 |
4.95 |
|
|
|
| 62 |
Bu |
819 |
814 |
0.79 |
|
|
|
| 63 |
Bu |
608 |
604 |
0.74 |
|
|
|
| 64 |
Bu |
527 |
524 |
5.31 |
|
|
|
| 65 |
Bu |
514 |
511 |
23.37 |
|
|
|
| 66 |
Bu |
86 |
85 |
1.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40706.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 40470.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.155 |
|
|
|
| 2 |
N |
-0.155 |
|
|
|
| 3 |
C |
0.033 |
|
|
|
| 4 |
C |
0.033 |
|
|
|
| 5 |
C |
0.047 |
|
|
|
| 6 |
C |
0.047 |
|
|
|
| 7 |
C |
0.064 |
|
|
|
| 8 |
C |
0.064 |
|
|
|
| 9 |
C |
0.035 |
|
|
|
| 10 |
C |
0.035 |
|
|
|
| 11 |
C |
0.031 |
|
|
|
| 12 |
C |
0.031 |
|
|
|
| 13 |
C |
0.030 |
|
|
|
| 14 |
C |
0.030 |
|
|
|
| 15 |
H |
-0.023 |
|
|
|
| 16 |
H |
-0.023 |
|
|
|
| 17 |
H |
-0.019 |
|
|
|
| 18 |
H |
-0.019 |
|
|
|
| 19 |
H |
-0.015 |
|
|
|
| 20 |
H |
-0.015 |
|
|
|
| 21 |
H |
-0.013 |
|
|
|
| 22 |
H |
-0.013 |
|
|
|
| 23 |
H |
-0.015 |
|
|
|
| 24 |
H |
-0.015 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-66.936 |
5.142 |
0.000 |
| y |
5.142 |
-69.525 |
0.000 |
| z |
0.000 |
0.000 |
-85.671 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.661 |
5.142 |
0.000 |
| y |
5.142 |
6.778 |
0.000 |
| z |
0.000 |
0.000 |
-17.440 |
|
| Polar |
|---|
| 3z2-r2 | -34.880 |
|---|
| x2-y2 | 2.589 |
|---|
| xy | 5.142 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
38.226 |
12.817 |
0.000 |
| y |
12.817 |
32.676 |
0.000 |
| z |
0.000 |
0.000 |
7.703 |
<r2> (average value of r
2) Å
2
| <r2> |
1106.846 |
| (<r2>)1/2 |
33.269 |