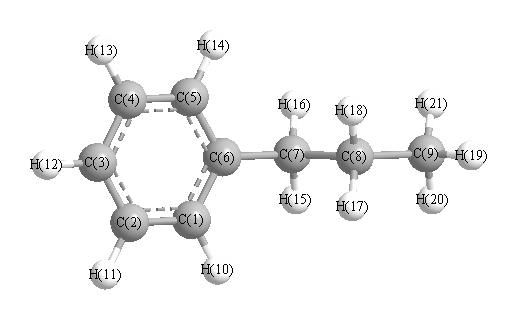
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3133 |
3115 |
16.90 |
|
|
|
| 2 |
A' |
3111 |
3093 |
4.16 |
|
|
|
| 3 |
A' |
3094 |
3076 |
10.70 |
|
|
|
| 4 |
A' |
3051 |
3033 |
28.31 |
|
|
|
| 5 |
A' |
2964 |
2947 |
71.27 |
|
|
|
| 6 |
A' |
2962 |
2945 |
15.03 |
|
|
|
| 7 |
A' |
2953 |
2936 |
5.03 |
|
|
|
| 8 |
A' |
1615 |
1606 |
6.73 |
|
|
|
| 9 |
A' |
1480 |
1471 |
11.72 |
|
|
|
| 10 |
A' |
1432 |
1424 |
5.01 |
|
|
|
| 11 |
A' |
1417 |
1409 |
0.03 |
|
|
|
| 12 |
A' |
1407 |
1399 |
0.38 |
|
|
|
| 13 |
A' |
1346 |
1338 |
0.58 |
|
|
|
| 14 |
A' |
1321 |
1314 |
2.90 |
|
|
|
| 15 |
A' |
1254 |
1247 |
2.26 |
|
|
|
| 16 |
A' |
1195 |
1188 |
2.08 |
|
|
|
| 17 |
A' |
1156 |
1150 |
0.13 |
|
|
|
| 18 |
A' |
1076 |
1070 |
6.54 |
|
|
|
| 19 |
A' |
1040 |
1034 |
0.40 |
|
|
|
| 20 |
A' |
1027 |
1022 |
3.06 |
|
|
|
| 21 |
A' |
985 |
979 |
0.38 |
|
|
|
| 22 |
A' |
968 |
963 |
0.01 |
|
|
|
| 23 |
A' |
899 |
893 |
0.03 |
|
|
|
| 24 |
A' |
880 |
875 |
2.15 |
|
|
|
| 25 |
A' |
805 |
800 |
0.27 |
|
|
|
| 26 |
A' |
742 |
737 |
17.93 |
|
|
|
| 27 |
A' |
695 |
691 |
28.40 |
|
|
|
| 28 |
A' |
578 |
574 |
3.67 |
|
|
|
| 29 |
A' |
493 |
490 |
6.32 |
|
|
|
| 30 |
A' |
304 |
302 |
0.04 |
|
|
|
| 31 |
A' |
269 |
267 |
0.45 |
|
|
|
| 32 |
A' |
99 |
99 |
0.26 |
|
|
|
| 33 |
A" |
3120 |
3101 |
32.22 |
|
|
|
| 34 |
A" |
3096 |
3078 |
5.84 |
|
|
|
| 35 |
A" |
3041 |
3023 |
42.28 |
|
|
|
| 36 |
A" |
3011 |
2994 |
19.10 |
|
|
|
| 37 |
A" |
2988 |
2971 |
0.13 |
|
|
|
| 38 |
A" |
1593 |
1584 |
0.65 |
|
|
|
| 39 |
A" |
1436 |
1427 |
3.23 |
|
|
|
| 40 |
A" |
1419 |
1411 |
7.88 |
|
|
|
| 41 |
A" |
1370 |
1362 |
0.34 |
|
|
|
| 42 |
A" |
1295 |
1288 |
0.61 |
|
|
|
| 43 |
A" |
1266 |
1259 |
0.17 |
|
|
|
| 44 |
A" |
1196 |
1189 |
0.17 |
|
|
|
| 45 |
A" |
1135 |
1128 |
0.02 |
|
|
|
| 46 |
A" |
1084 |
1078 |
5.72 |
|
|
|
| 47 |
A" |
1033 |
1027 |
0.79 |
|
|
|
| 48 |
A" |
945 |
940 |
0.01 |
|
|
|
| 49 |
A" |
845 |
840 |
0.88 |
|
|
|
| 50 |
A" |
829 |
824 |
0.06 |
|
|
|
| 51 |
A" |
730 |
726 |
1.76 |
|
|
|
| 52 |
A" |
611 |
608 |
0.04 |
|
|
|
| 53 |
A" |
400 |
397 |
0.00 |
|
|
|
| 54 |
A" |
338 |
336 |
0.05 |
|
|
|
| 55 |
A" |
245 |
244 |
0.01 |
|
|
|
| 56 |
A" |
89 |
89 |
0.01 |
|
|
|
| 57 |
A" |
40 |
40 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39450.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 39222.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.012 |
|
|
|
| 2 |
C |
0.028 |
|
|
|
| 3 |
C |
0.019 |
|
|
|
| 4 |
C |
0.028 |
|
|
|
| 5 |
C |
0.012 |
|
|
|
| 6 |
C |
-0.043 |
|
|
|
| 7 |
C |
0.025 |
|
|
|
| 8 |
C |
-0.062 |
|
|
|
| 9 |
C |
-0.049 |
|
|
|
| 10 |
H |
-0.040 |
|
|
|
| 11 |
H |
-0.022 |
|
|
|
| 12 |
H |
-0.023 |
|
|
|
| 13 |
H |
-0.022 |
|
|
|
| 14 |
H |
-0.040 |
|
|
|
| 15 |
H |
0.017 |
|
|
|
| 16 |
H |
0.017 |
|
|
|
| 17 |
H |
0.024 |
|
|
|
| 18 |
H |
0.024 |
|
|
|
| 19 |
H |
0.029 |
|
|
|
| 20 |
H |
0.031 |
|
|
|
| 21 |
H |
0.031 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.150 |
0.380 |
0.000 |
0.409 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-58.753 |
-0.164 |
0.000 |
| y |
-0.164 |
-52.424 |
0.000 |
| z |
0.000 |
0.000 |
-51.987 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.547 |
-0.164 |
0.000 |
| y |
-0.164 |
2.946 |
0.000 |
| z |
0.000 |
0.000 |
3.601 |
|
| Polar |
|---|
| 3z2-r2 | 7.203 |
|---|
| x2-y2 | -6.329 |
|---|
| xy | -0.164 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.784 |
-1.201 |
0.000 |
| y |
-1.201 |
19.853 |
0.000 |
| z |
0.000 |
0.000 |
15.182 |
<r2> (average value of r
2) Å
2
| <r2> |
444.132 |
| (<r2>)1/2 |
21.074 |