return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H7 (cyclopentenyl radical)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A2
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-194.115032
Energy at 298.15K-194.122106
HF Energy-193.384263
Nuclear repulsion energy161.801989
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3236 3084        
2 A1 3188 3039        
3 A1 3033 2891        
4 A1 1489 1419        
5 A1 1478 1409        
6 A1 1289 1228        
7 A1 1092 1040        
8 A1 1045 996        
9 A1 921 878        
10 A1 804 766        
11 A2 3050 2907        
12 A2 1223 1166        
13 A2 983 937        
14 A2 580 553        
15 A2 73 69        
16 B1 3069 2925        
17 B1 1134 1080        
18 B1 925 882        
19 B1 798 761        
20 B1 588 560        
21 B1 394 375        
22 B2 3224 3073        
23 B2 3018 2876        
24 B2 1457 1388        
25 B2 1417 1351        
26 B2 1314 1253        
27 B2 1288 1227        
28 B2 1066 1016        
29 B2 917 874        
30 B2 716 682        

Unscaled Zero Point Vibrational Energy (zpe) 22402.5 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 21351.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.25662 0.25025 0.13305

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.282
C2 0.000 1.159 0.509
C3 0.000 -1.159 0.509
C4 0.000 0.777 -0.971
C5 0.000 -0.777 -0.971
H6 0.000 0.000 2.381
H7 0.000 2.233 0.719
H8 0.000 -2.233 0.719
H9 0.887 1.194 -1.490
H10 -0.887 1.194 -1.490
H11 0.887 -1.194 -1.490
H12 -0.887 -1.194 -1.490

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.39301.39302.38292.38291.09872.30332.30333.14623.14623.14623.1462
C21.39302.31751.52802.43642.20151.09513.39872.18742.18743.21253.2125
C31.39302.31752.43641.52802.20153.39871.09513.21253.21252.18742.1874
C42.38291.52802.43641.55393.44032.23103.45231.10981.10982.22332.2233
C52.38292.43641.52801.55393.44033.45232.23102.22332.22331.10981.1098
H61.09872.20152.20153.44033.44032.78372.78374.14704.14704.14704.1470
H72.30331.09513.39872.23103.45232.78374.46692.59782.59784.17354.1735
H82.30333.39871.09513.45232.23102.78374.46694.17354.17352.59782.5978
H93.14622.18743.21251.10982.22334.14702.59784.17351.77492.38862.9759
H103.14622.18743.21251.10982.22334.14702.59784.17351.77492.97592.3886
H113.14623.21252.18742.22331.10984.14704.17352.59782.38862.97591.7749
H123.14623.21252.18742.22331.10984.14704.17352.59782.97592.38861.7749

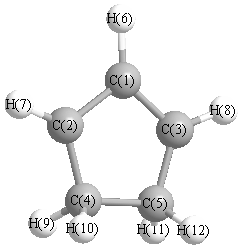
picture of cyclopentenyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 109.245 C1 C2 H7 135.211
C1 C3 C5 109.245 C1 C3 H8 135.211
C2 C1 C3 112.572 C2 C1 H6 123.714
C2 C4 C5 104.469 C2 C4 H9 111.053
C2 C4 H10 111.053 C3 C1 H6 123.714
C3 C5 C4 104.469 C3 C5 H11 111.053
C3 C5 H12 111.053 C4 C2 H7 115.544
C4 C5 H11 112.092 C4 C5 H12 112.092
C5 C3 H8 115.544 C5 C4 H9 112.092
C5 C4 H10 112.092 H9 C4 H10 106.199
H11 C5 H12 106.199
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability