return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H7 (cyclopentenyl radical)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A2
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-194.078482
Energy at 298.15K-194.085712
HF Energy-193.384926
Nuclear repulsion energy162.411275
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3271 3130 8.42      
2 A1 3222 3082 8.64      
3 A1 3064 2931 38.97      
4 A1 1532 1466 0.09      
5 A1 1496 1431 2.96      
6 A1 1317 1260 0.09      
7 A1 1116 1068 3.05      
8 A1 1061 1015 0.01      
9 A1 936 896 2.16      
10 A1 820 784 0.16      
11 A2 3083 2950 0.00      
12 A2 1240 1186 0.00      
13 A2 1005 962 0.00      
14 A2 634 607 0.00      
15 A2 89 85 0.00      
16 B1 3102 2968 46.77      
17 B1 1153 1103 3.31      
18 B1 985 942 4.83      
19 B1 816 781 3.42      
20 B1 622 595 49.88      
21 B1 419 401 7.11      
22 B2 3258 3117 6.81      
23 B2 3050 2918 35.16      
24 B2 1469 1406 0.42      
25 B2 1407 1347 5.16      
26 B2 1326 1269 1.97      
27 B2 1311 1254 1.80      
28 B2 1065 1019 0.51      
29 B2 920 880 12.85      
30 B2 726 695 0.18      

Unscaled Zero Point Vibrational Energy (zpe) 22758.1 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21772.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.25903 0.25169 0.13408

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.275
C2 0.000 1.151 0.509
C3 0.000 -1.151 0.509
C4 0.000 0.775 -0.969
C5 0.000 -0.775 -0.969
H6 0.000 0.000 2.371
H7 0.000 2.222 0.726
H8 0.000 -2.222 0.726
H9 0.886 1.193 -1.487
H10 -0.886 1.193 -1.487
H11 0.886 -1.193 -1.487
H12 -0.886 -1.193 -1.487

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.38261.38262.37392.37391.09642.28842.28843.13553.13553.13553.1355
C21.38262.30201.52472.42802.18931.09273.37982.18332.18333.20303.2030
C31.38262.30202.42801.52472.18933.37981.09273.20303.20302.18332.1833
C42.37391.52472.42801.55103.42892.22853.44361.10751.10752.21962.2196
C52.37392.42801.52471.55103.42893.44362.22852.21962.21961.10751.1075
H61.09642.18932.18933.42893.42892.76432.76434.13374.13374.13374.1337
H72.28841.09273.37982.22853.44362.76434.44362.59622.59624.16434.1643
H82.28843.37981.09273.44362.22852.76434.44364.16434.16432.59622.5962
H93.13552.18333.20301.10752.21964.13372.59624.16431.77152.38572.9715
H103.13552.18333.20301.10752.21964.13372.59624.16431.77152.97152.3857
H113.13553.20302.18332.21961.10754.13374.16432.59622.38572.97151.7715
H123.13553.20302.18332.21961.10754.13374.16432.59622.97152.38571.7715

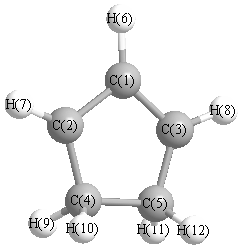
picture of cyclopentenyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 109.383 C1 C2 H7 134.864
C1 C3 C5 109.383 C1 C3 H8 134.864
C2 C1 C3 112.716 C2 C1 H6 123.642
C2 C4 C5 104.259 C2 C4 H9 111.100
C2 C4 H10 111.100 C3 C1 H6 123.642
C3 C5 C4 104.259 C3 C5 H11 111.100
C3 C5 H12 111.100 C4 C2 H7 115.752
C4 C5 H11 112.140 C4 C5 H12 112.140
C5 C3 H8 115.752 C5 C4 H9 112.140
C5 C4 H10 112.140 H9 C4 H10 106.222
H11 C5 H12 106.222
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability