return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-1149.658606
Energy at 298.15K-1149.662760
HF Energy-1148.558643
Nuclear repulsion energy445.722616
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3248 3116 0.00      
2 Ag 1664 1596 0.00      
3 Ag 1194 1145 0.00      
4 Ag 1138 1092 0.00      
5 Ag 761 730 0.00      
6 Ag 332 319 0.00      
7 Au 966 927 0.00      
8 Au 413 396 0.00      
9 B1g 839 805 0.00      
10 B1u 3231 3100 1.69      
11 B1u 1526 1464 95.70      
12 B1u 1130 1084 84.70      
13 B1u 1024 982 40.81      
14 B1u 558 536 23.85      
15 B2g 932 894 0.00      
16 B2g 636 610 0.00      
17 B2g 299 286 0.00      
18 B2u 3246 3115 2.55      
19 B2u 1433 1375 5.88      
20 B2u 1272 1220 0.01      
21 B2u 1120 1075 5.29      
22 B2u 220 211 0.94      
23 B3g 3232 3101 0.00      
24 B3g 1647 1580 0.00      
25 B3g 1310 1257 0.00      
26 B3g 637 611 0.00      
27 B3g 352 338 0.00      
28 B3u 830 796 47.37      
29 B3u 486 467 15.28      
30 B3u 103 99 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 17888.6 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 17162.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.18657 0.02192 0.01961

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.395
C2 0.000 0.000 -1.395
C3 0.000 1.220 0.702
C4 0.000 -1.220 0.702
C5 0.000 -1.220 -0.702
C6 0.000 1.220 -0.702
Cl7 0.000 0.000 3.147
Cl8 0.000 0.000 -3.147
H9 0.000 2.164 1.255
H10 0.000 -2.164 1.255
H11 0.000 -2.164 -1.255
H12 0.000 2.164 -1.255

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.79091.40351.40352.42662.42661.75114.54202.16902.16903.42173.4217
C22.79092.42662.42661.40351.40354.54201.75113.42173.42172.16902.1690
C31.40352.42662.44052.81561.40422.73214.03741.09403.42953.90962.1726
C41.40352.42662.44051.40422.81562.73214.03743.42951.09402.17263.9096
C52.42661.40352.81561.40422.44054.03742.73213.90962.17261.09403.4295
C62.42661.40351.40422.81562.44054.03742.73212.17263.90963.42951.0940
Cl71.75114.54202.73212.73214.03744.03746.29312.87482.87484.90464.9046
Cl84.54201.75114.03744.03742.73212.73216.29314.90464.90462.87482.8748
H92.16903.42171.09403.42953.90962.17262.87484.90464.32895.00362.5093
H102.16903.42173.42951.09402.17263.90962.87484.90464.32892.50935.0036
H113.42172.16903.90962.17261.09403.42954.90462.87485.00362.50934.3289
H123.42172.16902.17263.90963.42951.09404.90462.87482.50935.00364.3289

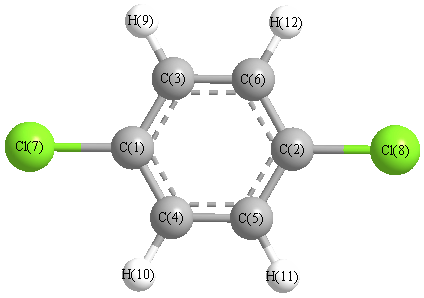
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.606 C1 C3 H9 120.057
C1 C4 C5 119.606 C1 C4 H10 120.057
C2 C5 C4 119.606 C2 C5 H11 120.057
C2 C6 C3 119.606 C2 C6 H12 120.057
C3 C1 C4 120.787 C3 C1 Cl7 119.606
C3 C6 H12 120.337 C4 C1 Cl7 119.606
C4 C5 H11 120.337 C5 C2 C6 120.787
C5 C2 Cl8 119.606 C5 C4 H10 120.337
C6 C2 Cl8 119.606 C6 C3 H9 120.337
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability