return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-1149.650147
Energy at 298.15K-1149.654330
HF Energy-1148.558998
Nuclear repulsion energy446.175523
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3255 3114 0.00      
2 Ag 1674 1601 0.00      
3 Ag 1198 1146 0.00      
4 Ag 1147 1097 0.00      
5 Ag 767 734 0.00      
6 Ag 334 319 0.00      
7 Au 970 928 0.00      
8 Au 414 396 0.00      
9 B1g 843 806 0.00      
10 B1u 3238 3098 1.45      
11 B1u 1534 1467 93.89      
12 B1u 1138 1089 81.68      
13 B1u 1031 986 39.52      
14 B1u 564 539 22.12      
15 B2g 936 895 0.00      
16 B2g 637 610 0.00      
17 B2g 300 287 0.00      
18 B2u 3254 3113 2.12      
19 B2u 1441 1379 5.40      
20 B2u 1289 1234 0.01      
21 B2u 1125 1076 5.53      
22 B2u 221 211 0.95      
23 B3g 3239 3099 0.00      
24 B3g 1656 1584 0.00      
25 B3g 1315 1258 0.00      
26 B3g 639 612 0.00      
27 B3g 354 339 0.00      
28 B3u 833 797 47.92      
29 B3u 489 468 15.14      
30 B3u 103 99 0.43      

Unscaled Zero Point Vibrational Energy (zpe) 17968.1 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 17190.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.18698 0.02196 0.01965

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.395
C2 0.000 0.000 -1.395
C3 0.000 1.219 0.701
C4 0.000 -1.219 0.701
C5 0.000 -1.219 -0.701
C6 0.000 1.219 -0.701
Cl7 0.000 0.000 3.143
Cl8 0.000 0.000 -3.143
H9 0.000 2.163 1.254
H10 0.000 -2.163 1.254
H11 0.000 -2.163 -1.254
H12 0.000 2.163 -1.254

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.78911.40211.40212.42452.42451.74894.53792.16732.16733.41913.4191
C22.78912.42452.42451.40211.40214.53791.74893.41913.41912.16732.1673
C31.40212.42452.43762.81231.40262.72934.03321.09373.42633.90602.1709
C41.40212.42452.43761.40262.81232.72934.03323.42631.09372.17093.9060
C52.42451.40212.81231.40262.43764.03322.72933.90602.17091.09373.4263
C62.42451.40211.40262.81232.43764.03322.72932.17093.90603.42631.0937
Cl71.74894.53792.72932.72934.03324.03326.28682.87212.87214.90014.9001
Cl84.53791.74894.03324.03322.72932.72936.28684.90014.90012.87212.8721
H92.16733.41911.09373.42633.90602.17092.87214.90014.32554.99962.5072
H102.16733.41913.42631.09372.17093.90602.87214.90014.32552.50724.9996
H113.41912.16733.90602.17091.09373.42634.90012.87214.99962.50724.3255
H123.41912.16732.17093.90603.42631.09374.90012.87212.50724.99964.3255

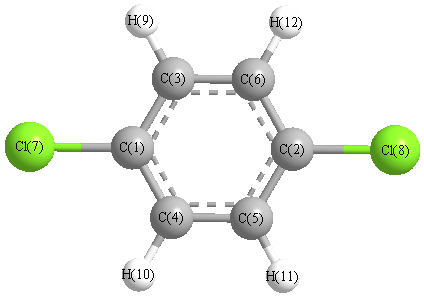
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.630 C1 C3 H9 120.039
C1 C4 C5 119.630 C1 C4 H10 120.039
C2 C5 C4 119.630 C2 C5 H11 120.039
C2 C6 C3 119.630 C2 C6 H12 120.039
C3 C1 C4 120.739 C3 C1 Cl7 119.630
C3 C6 H12 120.331 C4 C1 Cl7 119.630
C4 C5 H11 120.331 C5 C2 C6 120.739
C5 C2 Cl8 119.630 C5 C4 H10 120.331
C6 C2 Cl8 119.630 C6 C3 H9 120.331
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability