return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4O2 (parabenzoquinone)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-380.405919
Energy at 298.15K-380.410371
Nuclear repulsion energy320.525468
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3244 3104 0.00      
2 Ag 1835 1755 0.00      
3 Ag 1717 1642 0.00      
4 Ag 1161 1110 0.00      
5 Ag 784 750 0.00      
6 Ag 450 431 0.00      
7 Au 1014 970 0.00      
8 Au 338 324 0.00      
9 B1g 766 733 0.00      
10 B1u 3223 3083 1.23      
11 B1u 1830 1750 240.76      
12 B1u 1386 1326 5.22      
13 B1u 948 906 9.09      
14 B1u 763 730 0.10      
15 B2g 1010 966 0.00      
16 B2g 784 750 0.00      
17 B2g 225 215 0.00      
18 B2u 3242 3102 5.52      
19 B2u 1700 1626 13.83      
20 B2u 1342 1284 83.86      
21 B2u 1086 1039 38.72      
22 B2u 415 397 25.23      
23 B3g 3223 3084 0.00      
24 B3g 1431 1369 0.00      
25 B3g 1245 1192 0.00      
26 B3g 607 581 0.00      
27 B3g 451 432 0.00      
28 B3u 910 871 75.84      
29 B3u 515 493 3.08      
30 B3u 92 88 10.89      

Unscaled Zero Point Vibrational Energy (zpe) 18867.9 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 18051.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.17245 0.05459 0.04147

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.459
C2 0.000 0.000 -1.459
C3 0.000 1.277 0.675
C4 0.000 -1.277 0.675
C5 0.000 -1.277 -0.675
C6 0.000 1.277 -0.675
O7 0.000 0.000 2.678
O8 0.000 0.000 -2.678
H9 0.000 2.201 1.264
H10 0.000 -2.201 1.264
H11 0.000 -2.201 -1.264
H12 0.000 2.201 -1.264

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 O8 H9 H10 H11 H12
C12.91871.49821.49822.48732.48731.21824.13692.20932.20933.50133.5013
C22.91872.48732.48731.49821.49824.13691.21823.50133.50132.20932.2093
C31.49822.48732.55332.88851.35062.37463.58771.09563.52673.98152.1481
C41.49822.48732.55331.35062.88852.37463.58773.52671.09562.14813.9815
C52.48731.49822.88851.35062.55333.58772.37463.98152.14811.09563.5267
C62.48731.49821.35062.88852.55333.58772.37462.14813.98153.52671.0956
O71.21824.13692.37462.37463.58773.58775.35512.61562.61564.51424.5142
O84.13691.21823.58773.58772.37462.37465.35514.51424.51422.61562.6156
H92.20933.50131.09563.52673.98152.14812.61564.51424.40135.07552.5279
H102.20933.50133.52671.09562.14813.98152.61564.51424.40132.52795.0755
H113.50132.20933.98152.14811.09563.52674.51422.61565.07552.52794.4013
H123.50132.20932.14813.98153.52671.09564.51422.61562.52795.07554.4013

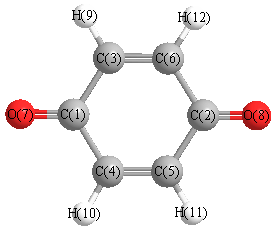
picture of parabenzoquinone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 121.555 C1 C3 H9 115.946
C1 C4 C5 121.555 C1 C4 H10 115.946
C2 C5 C4 121.555 C2 C5 H11 115.946
C2 C6 C3 121.555 C2 C6 H12 115.946
C3 C1 C4 116.889 C3 C1 O7 121.555
C3 C6 H12 122.498 C4 C1 O7 121.555
C4 C5 H11 122.498 C5 C2 C6 116.889
C5 C2 O8 121.555 C5 C4 H10 122.498
C6 C2 O8 121.555 C6 C3 H9 122.498
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability