Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -5223.430265 |
| Energy at 298.15K | -5223.440473 |
| HF Energy | -5222.843407 |
| Nuclear repulsion energy | 415.391220 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3134 |
3007 |
0.00 |
|
|
|
| 2 |
Ag |
1490 |
1430 |
0.00 |
|
|
|
| 3 |
Ag |
1306 |
1253 |
0.00 |
|
|
|
| 4 |
Ag |
1090 |
1046 |
0.00 |
|
|
|
| 5 |
Ag |
698 |
670 |
0.00 |
|
|
|
| 6 |
Ag |
195 |
187 |
0.00 |
|
|
|
| 7 |
Au |
3218 |
3087 |
2.06 |
|
|
|
| 8 |
Au |
1112 |
1067 |
3.68 |
|
|
|
| 9 |
Au |
759 |
728 |
3.21 |
|
|
|
| 10 |
Au |
108 |
104 |
3.91 |
|
|
|
| 11 |
Bg |
3198 |
3068 |
0.00 |
|
|
|
| 12 |
Bg |
1291 |
1238 |
0.00 |
|
|
|
| 13 |
Bg |
957 |
918 |
0.00 |
|
|
|
| 14 |
Bu |
3137 |
3010 |
10.27 |
|
|
|
| 15 |
Bu |
1482 |
1422 |
5.09 |
|
|
|
| 16 |
Bu |
1216 |
1167 |
60.74 |
|
|
|
| 17 |
Bu |
621 |
596 |
53.04 |
|
|
|
| 18 |
Bu |
185 |
177 |
5.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12597.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12086.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.494 |
0.578 |
0.000 |
| C2 |
-0.494 |
-0.578 |
0.000 |
| Br3 |
-0.494 |
2.274 |
0.000 |
| Br4 |
0.494 |
-2.274 |
0.000 |
| H5 |
1.126 |
0.575 |
0.900 |
| H6 |
1.126 |
0.575 |
-0.900 |
| H7 |
-1.126 |
-0.575 |
0.900 |
| H8 |
-1.126 |
-0.575 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5205 | 1.9625 | 2.8528 | 1.0999 | 1.0999 | 2.1823 | 2.1823 |
C2 | 1.5205 | | 2.8528 | 1.9625 | 2.1823 | 2.1823 | 1.0999 | 1.0999 | Br3 | 1.9625 | 2.8528 | | 4.6548 | 2.5140 | 2.5140 | 3.0544 | 3.0544 | Br4 | 2.8528 | 1.9625 | 4.6548 | | 3.0544 | 3.0544 | 2.5140 | 2.5140 | H5 | 1.0999 | 2.1823 | 2.5140 | 3.0544 | | 1.8000 | 2.5284 | 3.1037 | H6 | 1.0999 | 2.1823 | 2.5140 | 3.0544 | 1.8000 | | 3.1037 | 2.5284 | H7 | 2.1823 | 1.0999 | 3.0544 | 2.5140 | 2.5284 | 3.1037 | | 1.8000 | H8 | 2.1823 | 1.0999 | 3.0544 | 2.5140 | 3.1037 | 2.5284 | 1.8000 | |
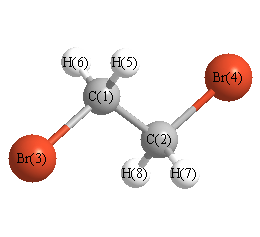 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.326 |
|
C1 |
C2 |
H7 |
111.772 |
| C1 |
C2 |
H8 |
111.772 |
|
C2 |
C1 |
Br3 |
109.326 |
| C2 |
C1 |
H5 |
111.772 |
|
C2 |
C1 |
H6 |
111.772 |
| Br3 |
C1 |
H5 |
106.959 |
|
Br3 |
C1 |
H6 |
106.959 |
| Br4 |
C2 |
H7 |
106.959 |
|
Br4 |
C2 |
H8 |
106.959 |
| H5 |
C1 |
H6 |
109.819 |
|
H7 |
C2 |
H8 |
109.819 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -5223.427040 |
| Energy at 298.15K | |
| HF Energy | -5222.839423 |
| Nuclear repulsion energy | 449.591180 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3181 |
3052 |
0.36 |
|
|
|
| 2 |
A |
3118 |
2991 |
20.12 |
|
|
|
| 3 |
A |
1471 |
1412 |
0.18 |
|
|
|
| 4 |
A |
1321 |
1268 |
24.46 |
|
|
|
| 5 |
A |
1209 |
1159 |
2.15 |
|
|
|
| 6 |
A |
1059 |
1016 |
0.33 |
|
|
|
| 7 |
A |
924 |
886 |
6.00 |
|
|
|
| 8 |
A |
573 |
550 |
7.43 |
|
|
|
| 9 |
A |
232 |
223 |
0.84 |
|
|
|
| 10 |
A |
83 |
80 |
0.22 |
|
|
|
| 11 |
B |
3192 |
3063 |
2.92 |
|
|
|
| 12 |
B |
3108 |
2982 |
3.14 |
|
|
|
| 13 |
B |
1464 |
1404 |
7.83 |
|
|
|
| 14 |
B |
1292 |
1240 |
63.96 |
|
|
|
| 15 |
B |
1130 |
1084 |
2.71 |
|
|
|
| 16 |
B |
857 |
823 |
13.90 |
|
|
|
| 17 |
B |
609 |
584 |
11.93 |
|
|
|
| 18 |
B |
361 |
347 |
4.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12591.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12080.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.311 |
0.694 |
1.185 |
| C2 |
-0.311 |
-0.694 |
1.185 |
| Br3 |
-0.311 |
1.807 |
-0.295 |
| Br4 |
0.311 |
-1.807 |
-0.295 |
| H5 |
0.025 |
1.224 |
2.109 |
| H6 |
1.409 |
0.647 |
1.113 |
| H7 |
-0.025 |
-1.224 |
2.109 |
| H8 |
-1.409 |
-0.647 |
1.113 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5221 | 1.9541 | 2.9069 | 1.1027 | 1.1006 | 2.1558 | 2.1825 |
C2 | 1.5221 | | 2.9069 | 1.9541 | 2.1558 | 2.1825 | 1.1027 | 1.1006 | Br3 | 1.9541 | 2.9069 | | 3.6676 | 2.4971 | 2.5076 | 3.8797 | 3.0349 | Br4 | 2.9069 | 1.9541 | 3.6676 | | 3.8797 | 3.0349 | 2.4971 | 2.5076 | H5 | 1.1027 | 2.1558 | 2.4971 | 3.8797 | | 1.7997 | 2.4486 | 2.5591 | H6 | 1.1006 | 2.1825 | 2.5076 | 3.0349 | 1.7997 | | 2.5591 | 3.1002 | H7 | 2.1558 | 1.1027 | 3.8797 | 2.4971 | 2.4486 | 2.5591 | | 1.7997 | H8 | 2.1825 | 1.1006 | 3.0349 | 2.5076 | 2.5591 | 3.1002 | 1.7997 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.898 |
|
C1 |
C2 |
H7 |
109.389 |
| C1 |
C2 |
H8 |
111.637 |
|
C2 |
C1 |
Br3 |
112.898 |
| C2 |
C1 |
H5 |
109.389 |
|
C2 |
C1 |
H6 |
111.637 |
| Br3 |
C1 |
H5 |
106.176 |
|
Br3 |
C1 |
H6 |
107.012 |
| Br4 |
C2 |
H7 |
106.176 |
|
Br4 |
C2 |
H8 |
107.012 |
| H5 |
C1 |
H6 |
109.540 |
|
H7 |
C2 |
H8 |
109.540 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
