Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -5223.426007 |
| Energy at 298.15K | -5223.436241 |
| HF Energy | -5222.843413 |
| Nuclear repulsion energy | 416.019595 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3140 |
3004 |
0.00 |
|
|
|
| 2 |
Ag |
1493 |
1428 |
0.00 |
|
|
|
| 3 |
Ag |
1316 |
1259 |
0.00 |
|
|
|
| 4 |
Ag |
1093 |
1045 |
0.00 |
|
|
|
| 5 |
Ag |
713 |
682 |
0.00 |
|
|
|
| 6 |
Ag |
196 |
188 |
0.00 |
|
|
|
| 7 |
Au |
3223 |
3084 |
2.04 |
|
|
|
| 8 |
Au |
1120 |
1072 |
3.70 |
|
|
|
| 9 |
Au |
760 |
727 |
3.29 |
|
|
|
| 10 |
Au |
108 |
104 |
3.85 |
|
|
|
| 11 |
Bg |
3203 |
3065 |
0.00 |
|
|
|
| 12 |
Bg |
1296 |
1240 |
0.00 |
|
|
|
| 13 |
Bg |
961 |
920 |
0.00 |
|
|
|
| 14 |
Bu |
3143 |
3007 |
10.03 |
|
|
|
| 15 |
Bu |
1486 |
1421 |
5.28 |
|
|
|
| 16 |
Bu |
1225 |
1172 |
57.33 |
|
|
|
| 17 |
Bu |
636 |
609 |
45.82 |
|
|
|
| 18 |
Bu |
186 |
178 |
5.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12648.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12101.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.492 |
0.580 |
0.000 |
| C2 |
-0.492 |
-0.580 |
0.000 |
| Br3 |
-0.492 |
2.272 |
0.000 |
| Br4 |
0.492 |
-2.272 |
0.000 |
| H5 |
1.125 |
0.575 |
0.899 |
| H6 |
1.125 |
0.575 |
-0.899 |
| H7 |
-1.125 |
-0.575 |
0.899 |
| H8 |
-1.125 |
-0.575 |
-0.899 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5209 | 1.9567 | 2.8516 | 1.0998 | 1.0998 | 2.1810 | 2.1810 |
C2 | 1.5209 | | 2.8516 | 1.9567 | 2.1810 | 2.1810 | 1.0998 | 1.0998 | Br3 | 1.9567 | 2.8516 | | 4.6483 | 2.5105 | 2.5105 | 3.0513 | 3.0513 | Br4 | 2.8516 | 1.9567 | 4.6483 | | 3.0513 | 3.0513 | 2.5105 | 2.5105 | H5 | 1.0998 | 2.1810 | 2.5105 | 3.0513 | | 1.7977 | 2.5273 | 3.1015 | H6 | 1.0998 | 2.1810 | 2.5105 | 3.0513 | 1.7977 | | 3.1015 | 2.5273 | H7 | 2.1810 | 1.0998 | 3.0513 | 2.5105 | 2.5273 | 3.1015 | | 1.7977 | H8 | 2.1810 | 1.0998 | 3.0513 | 2.5105 | 3.1015 | 2.5273 | 1.7977 | |
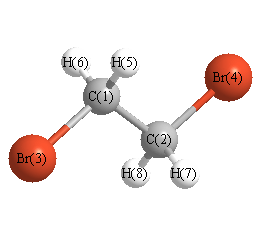 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.532 |
|
C1 |
C2 |
H7 |
111.646 |
| C1 |
C2 |
H8 |
111.646 |
|
C2 |
C1 |
Br3 |
109.532 |
| C2 |
C1 |
H5 |
111.646 |
|
C2 |
C1 |
H6 |
111.646 |
| Br3 |
C1 |
H5 |
107.089 |
|
Br3 |
C1 |
H6 |
107.089 |
| Br4 |
C2 |
H7 |
107.089 |
|
Br4 |
C2 |
H8 |
107.089 |
| H5 |
C1 |
H6 |
109.632 |
|
H7 |
C2 |
H8 |
109.632 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -5223.422839 |
| Energy at 298.15K | |
| HF Energy | -5222.839448 |
| Nuclear repulsion energy | 450.239398 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3049 |
0.43 |
|
|
|
| 2 |
A |
3123 |
2988 |
19.19 |
|
|
|
| 3 |
A |
1475 |
1411 |
0.20 |
|
|
|
| 4 |
A |
1330 |
1272 |
22.69 |
|
|
|
| 5 |
A |
1215 |
1162 |
1.81 |
|
|
|
| 6 |
A |
1063 |
1017 |
0.26 |
|
|
|
| 7 |
A |
929 |
889 |
5.79 |
|
|
|
| 8 |
A |
584 |
558 |
6.18 |
|
|
|
| 9 |
A |
234 |
223 |
0.82 |
|
|
|
| 10 |
A |
83 |
80 |
0.22 |
|
|
|
| 11 |
B |
3197 |
3059 |
2.74 |
|
|
|
| 12 |
B |
3114 |
2979 |
3.10 |
|
|
|
| 13 |
B |
1468 |
1404 |
8.02 |
|
|
|
| 14 |
B |
1302 |
1246 |
60.00 |
|
|
|
| 15 |
B |
1137 |
1088 |
2.68 |
|
|
|
| 16 |
B |
862 |
825 |
13.32 |
|
|
|
| 17 |
B |
621 |
594 |
9.90 |
|
|
|
| 18 |
B |
364 |
348 |
4.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12643.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12095.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.311 |
0.695 |
1.182 |
| C2 |
-0.311 |
-0.695 |
1.182 |
| Br3 |
-0.311 |
1.805 |
-0.295 |
| Br4 |
0.311 |
-1.805 |
-0.295 |
| H5 |
0.025 |
1.223 |
2.107 |
| H6 |
1.408 |
0.648 |
1.111 |
| H7 |
-0.025 |
-1.223 |
2.107 |
| H8 |
-1.408 |
-0.648 |
1.111 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5222 | 1.9491 | 2.9035 | 1.1026 | 1.1006 | 2.1551 | 2.1821 |
C2 | 1.5222 | | 2.9035 | 1.9491 | 2.1551 | 2.1821 | 1.1026 | 1.1006 | Br3 | 1.9491 | 2.9035 | | 3.6634 | 2.4936 | 2.5039 | 3.8750 | 3.0329 | Br4 | 2.9035 | 1.9491 | 3.6634 | | 3.8750 | 3.0329 | 2.4936 | 2.5039 | H5 | 1.1026 | 2.1551 | 2.4936 | 3.8750 | | 1.7984 | 2.4458 | 2.5578 | H6 | 1.1006 | 2.1821 | 2.5039 | 3.0329 | 1.7984 | | 2.5578 | 3.0998 | H7 | 2.1551 | 1.1026 | 3.8750 | 2.4936 | 2.4458 | 2.5578 | | 1.7984 | H8 | 2.1821 | 1.1006 | 3.0329 | 2.5039 | 2.5578 | 3.0998 | 1.7984 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.954 |
|
C1 |
C2 |
H7 |
109.344 |
| C1 |
C2 |
H8 |
111.594 |
|
C2 |
C1 |
Br3 |
112.954 |
| C2 |
C1 |
H5 |
109.344 |
|
C2 |
C1 |
H6 |
111.594 |
| Br3 |
C1 |
H5 |
106.264 |
|
Br3 |
C1 |
H6 |
107.073 |
| Br4 |
C2 |
H7 |
106.264 |
|
Br4 |
C2 |
H8 |
107.073 |
| H5 |
C1 |
H6 |
109.425 |
|
H7 |
C2 |
H8 |
109.425 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
