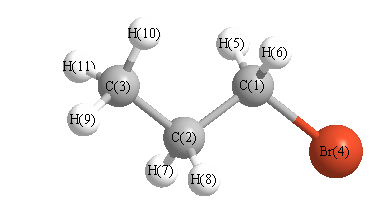
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3259 |
2805 |
39.05 |
|
|
|
| 2 |
A' |
3254 |
2801 |
24.85 |
|
|
|
| 3 |
A' |
3203 |
2757 |
11.96 |
|
|
|
| 4 |
A' |
3174 |
2732 |
35.72 |
|
|
|
| 5 |
A' |
1608 |
1384 |
3.56 |
|
|
|
| 6 |
A' |
1592 |
1370 |
0.37 |
|
|
|
| 7 |
A' |
1590 |
1369 |
4.28 |
|
|
|
| 8 |
A' |
1524 |
1312 |
0.15 |
|
|
|
| 9 |
A' |
1480 |
1274 |
6.75 |
|
|
|
| 10 |
A' |
1351 |
1162 |
63.15 |
|
|
|
| 11 |
A' |
1194 |
1027 |
1.27 |
|
|
|
| 12 |
A' |
1108 |
953 |
0.78 |
|
|
|
| 13 |
A' |
964 |
829 |
6.07 |
|
|
|
| 14 |
A' |
691 |
594 |
41.82 |
|
|
|
| 15 |
A' |
328 |
282 |
3.58 |
|
|
|
| 16 |
A' |
230 |
198 |
1.55 |
|
|
|
| 17 |
A" |
3329 |
2865 |
12.59 |
|
|
|
| 18 |
A" |
3256 |
2803 |
52.44 |
|
|
|
| 19 |
A" |
3233 |
2782 |
2.87 |
|
|
|
| 20 |
A" |
1594 |
1372 |
7.19 |
|
|
|
| 21 |
A" |
1418 |
1220 |
0.01 |
|
|
|
| 22 |
A" |
1335 |
1149 |
0.31 |
|
|
|
| 23 |
A" |
1139 |
980 |
1.84 |
|
|
|
| 24 |
A" |
916 |
788 |
0.00 |
|
|
|
| 25 |
A" |
797 |
686 |
3.02 |
|
|
|
| 26 |
A" |
258 |
222 |
0.06 |
|
|
|
| 27 |
A" |
122 |
105 |
0.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21971.3 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 18910.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.098 |
|
|
|
| 2 |
C |
-0.066 |
|
|
|
| 3 |
C |
-0.055 |
|
|
|
| 4 |
Br |
-0.157 |
|
|
|
| 5 |
H |
0.085 |
|
|
|
| 6 |
H |
0.085 |
|
|
|
| 7 |
H |
0.047 |
|
|
|
| 8 |
H |
0.047 |
|
|
|
| 9 |
H |
0.042 |
|
|
|
| 10 |
H |
0.034 |
|
|
|
| 11 |
H |
0.034 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.430 |
2.019 |
0.000 |
2.474 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.133 |
-0.154 |
0.000 |
| y |
-0.154 |
-38.311 |
0.000 |
| z |
0.000 |
0.000 |
-38.785 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.585 |
-0.154 |
0.000 |
| y |
-0.154 |
0.648 |
0.000 |
| z |
0.000 |
0.000 |
-0.063 |
|
| Polar |
|---|
| 3z2-r2 | -0.127 |
|---|
| x2-y2 | -0.822 |
|---|
| xy | -0.154 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
207.882 |
| (<r2>)1/2 |
14.418 |