Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -157.980625 |
| Energy at 298.15K | -157.991254 |
| HF Energy | -157.308489 |
| Nuclear repulsion energy | 129.599844 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
118 |
112 |
|
|
|
|
|
Au |
229 |
219 |
|
|
|
|
|
Bu |
255 |
243 |
|
|
|
|
|
Bg |
268 |
255 |
|
|
|
|
|
Ag |
424 |
404 |
|
|
|
|
|
Au |
734 |
699 |
|
|
|
|
|
Bg |
804 |
766 |
|
|
|
|
|
Ag |
852 |
812 |
|
|
|
|
|
Au |
951 |
907 |
|
|
|
|
|
Bu |
973 |
928 |
|
|
|
|
|
Bu |
1040 |
991 |
|
|
|
|
|
Ag |
1091 |
1040 |
|
|
|
|
|
Ag |
1170 |
1115 |
|
|
|
|
|
Bg |
1206 |
1149 |
|
|
|
|
|
Au |
1282 |
1222 |
|
|
|
|
|
Bu |
1309 |
1248 |
|
|
|
|
|
Bg |
1322 |
1260 |
|
|
|
|
|
Ag |
1394 |
1329 |
|
|
|
|
|
Bu |
1408 |
1342 |
|
|
|
|
|
Ag |
1414 |
1348 |
|
|
|
|
|
Ag |
1475 |
1406 |
|
|
|
|
|
Bu |
1479 |
1410 |
|
|
|
|
|
Bg |
1487 |
1417 |
|
|
|
|
|
Au |
1488 |
1419 |
|
|
|
|
|
Ag |
1494 |
1424 |
|
|
|
|
|
Bu |
1498 |
1428 |
|
|
|
|
|
Ag |
3021 |
2879 |
|
|
|
|
|
Bu |
3026 |
2884 |
|
|
|
|
|
Bu |
3034 |
2892 |
|
|
|
|
|
Ag |
3036 |
2894 |
|
|
|
|
|
Bg |
3054 |
2911 |
|
|
|
|
|
Au |
3075 |
2931 |
|
|
|
|
|
Bg |
3115 |
2969 |
|
|
|
|
|
Au |
3118 |
2972 |
|
|
|
|
|
Ag |
3121 |
2974 |
|
|
|
|
|
Bu |
3121 |
2975 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28943.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 27585.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.425 |
0.641 |
0.000 |
| C2 |
0.425 |
-0.641 |
0.000 |
| C3 |
0.425 |
1.921 |
0.000 |
| C4 |
-0.425 |
-1.921 |
0.000 |
| H5 |
-1.091 |
0.635 |
0.888 |
| H6 |
-1.091 |
0.635 |
-0.888 |
| H7 |
1.091 |
-0.635 |
0.888 |
| H8 |
1.091 |
-0.635 |
-0.888 |
| H9 |
-0.209 |
2.828 |
0.000 |
| H10 |
1.077 |
1.962 |
0.894 |
| H11 |
1.077 |
1.962 |
-0.894 |
| H12 |
0.209 |
-2.828 |
0.000 |
| H13 |
-1.077 |
-1.962 |
0.894 |
| H14 |
-1.077 |
-1.962 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5370 | 1.5362 | 2.5614 | 1.1104 | 1.1104 | 2.1709 | 2.1709 | 2.1978 | 2.1910 | 2.1910 | 3.5259 | 2.8283 | 2.8283 |
C2 | 1.5370 | | 2.5614 | 1.5362 | 2.1709 | 2.1709 | 1.1104 | 1.1104 | 3.5259 | 2.8283 | 2.8283 | 2.1978 | 2.1910 | 2.1910 | C3 | 1.5362 | 2.5614 | | 3.9343 | 2.1769 | 2.1769 | 2.7866 | 2.7866 | 1.1064 | 1.1077 | 1.1077 | 4.7535 | 4.2581 | 4.2581 | C4 | 2.5614 | 1.5362 | 3.9343 | | 2.7866 | 2.7866 | 2.1769 | 2.1769 | 4.7535 | 4.2581 | 4.2581 | 1.1064 | 1.1077 | 1.1077 | H5 | 1.1104 | 2.1709 | 2.1769 | 2.7866 | | 1.7763 | 2.5246 | 3.0869 | 2.5249 | 2.5420 | 3.1046 | 3.8039 | 2.5971 | 3.1499 | H6 | 1.1104 | 2.1709 | 2.1769 | 2.7866 | 1.7763 | | 3.0869 | 2.5246 | 2.5249 | 3.1046 | 2.5420 | 3.8039 | 3.1499 | 2.5971 | H7 | 2.1709 | 1.1104 | 2.7866 | 2.1769 | 2.5246 | 3.0869 | | 1.7763 | 3.8039 | 2.5971 | 3.1499 | 2.5249 | 2.5420 | 3.1046 | H8 | 2.1709 | 1.1104 | 2.7866 | 2.1769 | 3.0869 | 2.5246 | 1.7763 | | 3.8039 | 3.1499 | 2.5971 | 2.5249 | 3.1046 | 2.5420 | H9 | 2.1978 | 3.5259 | 1.1064 | 4.7535 | 2.5249 | 2.5249 | 3.8039 | 3.8039 | | 1.7898 | 1.7898 | 5.6711 | 4.9494 | 4.9494 | H10 | 2.1910 | 2.8283 | 1.1077 | 4.2581 | 2.5420 | 3.1046 | 2.5971 | 3.1499 | 1.7898 | | 1.7884 | 4.9494 | 4.4765 | 4.8205 | H11 | 2.1910 | 2.8283 | 1.1077 | 4.2581 | 3.1046 | 2.5420 | 3.1499 | 2.5971 | 1.7898 | 1.7884 | | 4.9494 | 4.8205 | 4.4765 | H12 | 3.5259 | 2.1978 | 4.7535 | 1.1064 | 3.8039 | 3.8039 | 2.5249 | 2.5249 | 5.6711 | 4.9494 | 4.9494 | | 1.7898 | 1.7898 | H13 | 2.8283 | 2.1910 | 4.2581 | 1.1077 | 2.5971 | 3.1499 | 2.5420 | 3.1046 | 4.9494 | 4.4765 | 4.8205 | 1.7898 | | 1.7884 | H14 | 2.8283 | 2.1910 | 4.2581 | 1.1077 | 3.1499 | 2.5971 | 3.1046 | 2.5420 | 4.9494 | 4.8205 | 4.4765 | 1.7898 | 1.7884 | |
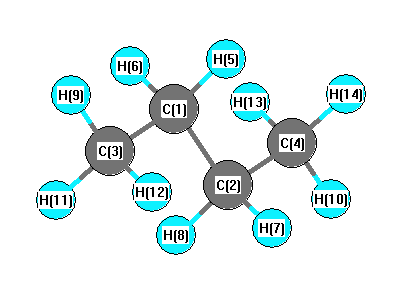 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.913 |
|
C1 |
C2 |
H7 |
109.104 |
| C1 |
C2 |
H8 |
109.104 |
|
C1 |
C3 |
H9 |
111.510 |
| C1 |
C3 |
H11 |
110.895 |
|
C1 |
C3 |
H12 |
30.956 |
| C2 |
C1 |
C3 |
112.913 |
|
C2 |
C1 |
H5 |
109.104 |
| C2 |
C1 |
H6 |
109.104 |
|
C2 |
C4 |
H10 |
17.282 |
| C2 |
C4 |
H13 |
110.895 |
|
C2 |
C4 |
H14 |
110.895 |
| C3 |
C1 |
H5 |
109.632 |
|
C3 |
C1 |
H6 |
109.632 |
| C4 |
C2 |
H7 |
109.632 |
|
C4 |
C2 |
H8 |
109.632 |
| H5 |
C1 |
H6 |
106.234 |
|
H7 |
C2 |
H8 |
106.234 |
| H9 |
C3 |
H11 |
107.865 |
|
H9 |
C3 |
H12 |
142.466 |
| H10 |
C4 |
H13 |
94.141 |
|
H10 |
C4 |
H14 |
114.283 |
| H11 |
C3 |
H12 |
93.666 |
|
H13 |
C4 |
H14 |
107.651 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -157.979546 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3130 |
2983 |
|
|
|
|
| 2 |
A |
3116 |
2970 |
|
|
|
|
| 3 |
A |
3068 |
2924 |
|
|
|
|
| 4 |
A |
3039 |
2896 |
|
|
|
|
| 5 |
A |
3028 |
2886 |
|
|
|
|
| 6 |
A |
1500 |
1430 |
|
|
|
|
| 7 |
A |
1493 |
1423 |
|
|
|
|
| 8 |
A |
1476 |
1407 |
|
|
|
|
| 9 |
A |
1408 |
1342 |
|
|
|
|
| 10 |
A |
1382 |
1317 |
|
|
|
|
| 11 |
A |
1306 |
1245 |
|
|
|
|
| 12 |
A |
1194 |
1138 |
|
|
|
|
| 13 |
A |
1109 |
1057 |
|
|
|
|
| 14 |
A |
990 |
943 |
|
|
|
|
| 15 |
A |
842 |
802 |
|
|
|
|
| 16 |
A |
797 |
760 |
|
|
|
|
| 17 |
A |
322 |
307 |
|
|
|
|
| 18 |
A |
271 |
259 |
|
|
|
|
| 19 |
A |
111 |
105 |
|
|
|
|
| 20 |
B |
3122 |
2976 |
|
|
|
|
| 21 |
B |
3117 |
2971 |
|
|
|
|
| 22 |
B |
3074 |
2929 |
|
|
|
|
| 23 |
B |
3036 |
2893 |
|
|
|
|
| 24 |
B |
3024 |
2882 |
|
|
|
|
| 25 |
B |
1494 |
1424 |
|
|
|
|
| 26 |
B |
1487 |
1417 |
|
|
|
|
| 27 |
B |
1471 |
1402 |
|
|
|
|
| 28 |
B |
1414 |
1347 |
|
|
|
|
| 29 |
B |
1360 |
1296 |
|
|
|
|
| 30 |
B |
1281 |
1221 |
|
|
|
|
| 31 |
B |
1153 |
1099 |
|
|
|
|
| 32 |
B |
983 |
937 |
|
|
|
|
| 33 |
B |
960 |
915 |
|
|
|
|
| 34 |
B |
745 |
710 |
|
|
|
|
| 35 |
B |
433 |
412 |
|
|
|
|
| 36 |
B |
221 |
211 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28977.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 27618.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
