Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -157.959370 |
| Energy at 298.15K | -157.970049 |
| HF Energy | -157.308907 |
| Nuclear repulsion energy | 129.781073 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
122 |
117 |
0.00 |
|
|
|
|
Au |
233 |
222 |
0.00 |
|
|
|
|
Bu |
259 |
247 |
0.02 |
|
|
|
|
Bg |
270 |
258 |
0.00 |
|
|
|
|
Ag |
429 |
410 |
0.00 |
|
|
|
|
Au |
741 |
709 |
2.05 |
|
|
|
|
Bg |
813 |
778 |
0.00 |
|
|
|
|
Ag |
861 |
824 |
0.00 |
|
|
|
|
Au |
962 |
920 |
0.73 |
|
|
|
|
Bu |
984 |
941 |
3.22 |
|
|
|
|
Bu |
1052 |
1006 |
0.10 |
|
|
|
|
Ag |
1102 |
1055 |
0.00 |
|
|
|
|
Ag |
1183 |
1132 |
0.00 |
|
|
|
|
Bg |
1218 |
1166 |
0.00 |
|
|
|
|
Au |
1295 |
1239 |
0.18 |
|
|
|
|
Bu |
1326 |
1268 |
5.52 |
|
|
|
|
Bg |
1336 |
1278 |
0.00 |
|
|
|
|
Ag |
1411 |
1350 |
0.00 |
|
|
|
|
Bu |
1425 |
1363 |
2.05 |
|
|
|
|
Ag |
1433 |
1371 |
0.00 |
|
|
|
|
Ag |
1489 |
1425 |
0.00 |
|
|
|
|
Bu |
1493 |
1428 |
1.39 |
|
|
|
|
Bg |
1499 |
1434 |
0.00 |
|
|
|
|
Au |
1500 |
1435 |
11.24 |
|
|
|
|
Ag |
1507 |
1442 |
0.00 |
|
|
|
|
Bu |
1511 |
1446 |
4.87 |
|
|
|
|
Ag |
3050 |
2918 |
0.00 |
|
|
|
|
Bu |
3055 |
2923 |
68.25 |
|
|
|
|
Bu |
3059 |
2927 |
51.79 |
|
|
|
|
Ag |
3062 |
2929 |
0.00 |
|
|
|
|
Bg |
3085 |
2951 |
0.00 |
|
|
|
|
Au |
3105 |
2970 |
27.24 |
|
|
|
|
Bg |
3142 |
3006 |
0.00 |
|
|
|
|
Au |
3145 |
3008 |
99.71 |
|
|
|
|
Ag |
3148 |
3011 |
0.00 |
|
|
|
|
Bu |
3148 |
3012 |
70.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29224.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 27959.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.424 |
0.640 |
0.000 |
| C2 |
0.424 |
-0.640 |
0.000 |
| C3 |
0.424 |
1.919 |
0.000 |
| C4 |
-0.424 |
-1.919 |
0.000 |
| H5 |
-1.089 |
0.635 |
0.886 |
| H6 |
-1.089 |
0.635 |
-0.886 |
| H7 |
1.089 |
-0.635 |
0.886 |
| H8 |
1.089 |
-0.635 |
-0.886 |
| H9 |
-0.209 |
2.824 |
0.000 |
| H10 |
1.075 |
1.961 |
0.893 |
| H11 |
1.075 |
1.961 |
-0.893 |
| H12 |
0.209 |
-2.824 |
0.000 |
| H13 |
-1.075 |
-1.961 |
0.893 |
| H14 |
-1.075 |
-1.961 |
-0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5351 | 1.5341 | 2.5586 | 1.1082 | 1.1082 | 2.1678 | 2.1678 | 2.1944 | 2.1882 | 2.1882 | 3.5212 | 2.8258 | 2.8258 |
C2 | 1.5351 | | 2.5586 | 1.5341 | 2.1678 | 2.1678 | 1.1082 | 1.1082 | 3.5212 | 2.8258 | 2.8258 | 2.1944 | 2.1882 | 2.1882 | C3 | 1.5341 | 2.5586 | | 3.9298 | 2.1731 | 2.1731 | 2.7837 | 2.7837 | 1.1045 | 1.1058 | 1.1058 | 4.7474 | 4.2537 | 4.2537 | C4 | 2.5586 | 1.5341 | 3.9298 | | 2.7837 | 2.7837 | 2.1731 | 2.1731 | 4.7474 | 4.2537 | 4.2537 | 1.1045 | 1.1058 | 1.1058 | H5 | 1.1082 | 2.1678 | 2.1731 | 2.7837 | | 1.7725 | 2.5213 | 3.0820 | 2.5202 | 2.5382 | 3.0995 | 3.7992 | 2.5958 | 3.1468 | H6 | 1.1082 | 2.1678 | 2.1731 | 2.7837 | 1.7725 | | 3.0820 | 2.5213 | 2.5202 | 3.0995 | 2.5382 | 3.7992 | 3.1468 | 2.5958 | H7 | 2.1678 | 1.1082 | 2.7837 | 2.1731 | 2.5213 | 3.0820 | | 1.7725 | 3.7992 | 2.5958 | 3.1468 | 2.5202 | 2.5382 | 3.0995 | H8 | 2.1678 | 1.1082 | 2.7837 | 2.1731 | 3.0820 | 2.5213 | 1.7725 | | 3.7992 | 3.1468 | 2.5958 | 2.5202 | 3.0995 | 2.5382 | H9 | 2.1944 | 3.5212 | 1.1045 | 4.7474 | 2.5202 | 2.5202 | 3.7992 | 3.7992 | | 1.7864 | 1.7864 | 5.6632 | 4.9437 | 4.9437 | H10 | 2.1882 | 2.8258 | 1.1058 | 4.2537 | 2.5382 | 3.0995 | 2.5958 | 3.1468 | 1.7864 | | 1.7851 | 4.9437 | 4.4726 | 4.8157 | H11 | 2.1882 | 2.8258 | 1.1058 | 4.2537 | 3.0995 | 2.5382 | 3.1468 | 2.5958 | 1.7864 | 1.7851 | | 4.9437 | 4.8157 | 4.4726 | H12 | 3.5212 | 2.1944 | 4.7474 | 1.1045 | 3.7992 | 3.7992 | 2.5202 | 2.5202 | 5.6632 | 4.9437 | 4.9437 | | 1.7864 | 1.7864 | H13 | 2.8258 | 2.1882 | 4.2537 | 1.1058 | 2.5958 | 3.1468 | 2.5382 | 3.0995 | 4.9437 | 4.4726 | 4.8157 | 1.7864 | | 1.7851 | H14 | 2.8258 | 2.1882 | 4.2537 | 1.1058 | 3.1468 | 2.5958 | 3.0995 | 2.5382 | 4.9437 | 4.8157 | 4.4726 | 1.7864 | 1.7851 | |
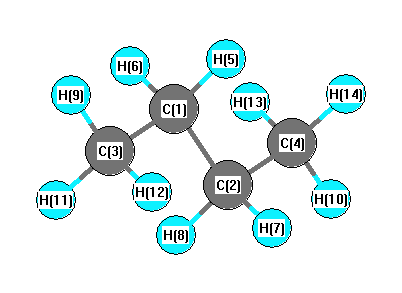 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.953 |
|
C1 |
C2 |
H7 |
109.126 |
| C1 |
C2 |
H8 |
109.126 |
|
C1 |
C3 |
H9 |
111.502 |
| C1 |
C3 |
H11 |
110.932 |
|
C1 |
C3 |
H12 |
30.945 |
| C2 |
C1 |
C3 |
112.953 |
|
C2 |
C1 |
H5 |
109.126 |
| C2 |
C1 |
H6 |
109.126 |
|
C2 |
C4 |
H10 |
17.274 |
| C2 |
C4 |
H13 |
110.932 |
|
C2 |
C4 |
H14 |
110.932 |
| C3 |
C1 |
H5 |
109.602 |
|
C3 |
C1 |
H6 |
109.602 |
| C4 |
C2 |
H7 |
109.602 |
|
C4 |
C2 |
H8 |
109.602 |
| H5 |
C1 |
H6 |
106.205 |
|
H7 |
C2 |
H8 |
106.205 |
| H9 |
C3 |
H11 |
107.840 |
|
H9 |
C3 |
H12 |
142.447 |
| H10 |
C4 |
H13 |
94.188 |
|
H10 |
C4 |
H14 |
114.314 |
| H11 |
C3 |
H12 |
93.710 |
|
H13 |
C4 |
H14 |
107.632 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -157.958221 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 131.592966 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3157 |
3020 |
32.43 |
|
|
|
| 2 |
A |
3143 |
3007 |
0.90 |
|
|
|
| 3 |
A |
3098 |
2964 |
25.39 |
|
|
|
| 4 |
A |
3065 |
2932 |
10.60 |
|
|
|
| 5 |
A |
3056 |
2924 |
36.46 |
|
|
|
| 6 |
A |
1512 |
1447 |
0.19 |
|
|
|
| 7 |
A |
1506 |
1441 |
3.95 |
|
|
|
| 8 |
A |
1489 |
1425 |
0.13 |
|
|
|
| 9 |
A |
1425 |
1363 |
1.75 |
|
|
|
| 10 |
A |
1400 |
1339 |
0.26 |
|
|
|
| 11 |
A |
1321 |
1264 |
0.20 |
|
|
|
| 12 |
A |
1206 |
1154 |
0.05 |
|
|
|
| 13 |
A |
1122 |
1073 |
0.05 |
|
|
|
| 14 |
A |
1002 |
958 |
0.20 |
|
|
|
| 15 |
A |
851 |
814 |
0.01 |
|
|
|
| 16 |
A |
805 |
770 |
1.06 |
|
|
|
| 17 |
A |
326 |
312 |
0.00 |
|
|
|
| 18 |
A |
274 |
262 |
0.03 |
|
|
|
| 19 |
A |
113 |
108 |
0.01 |
|
|
|
| 20 |
B |
3149 |
3013 |
50.00 |
|
|
|
| 21 |
B |
3144 |
3008 |
77.00 |
|
|
|
| 22 |
B |
3103 |
2969 |
28.90 |
|
|
|
| 23 |
B |
3061 |
2929 |
23.06 |
|
|
|
| 24 |
B |
3053 |
2921 |
30.06 |
|
|
|
| 25 |
B |
1506 |
1441 |
7.79 |
|
|
|
| 26 |
B |
1499 |
1434 |
6.45 |
|
|
|
| 27 |
B |
1485 |
1420 |
0.32 |
|
|
|
| 28 |
B |
1431 |
1369 |
5.08 |
|
|
|
| 29 |
B |
1377 |
1317 |
3.80 |
|
|
|
| 30 |
B |
1295 |
1239 |
0.55 |
|
|
|
| 31 |
B |
1166 |
1115 |
1.65 |
|
|
|
| 32 |
B |
994 |
951 |
1.00 |
|
|
|
| 33 |
B |
970 |
928 |
1.31 |
|
|
|
| 34 |
B |
753 |
720 |
2.30 |
|
|
|
| 35 |
B |
437 |
418 |
0.19 |
|
|
|
| 36 |
B |
222 |
212 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29256.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 27989.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
