Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -158.215059 |
| Energy at 298.15K | -158.225587 |
| HF Energy | -158.215059 |
| Nuclear repulsion energy | 129.607408 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
128 |
127 |
0.00 |
|
|
|
|
Au |
220 |
219 |
0.00 |
|
|
|
|
Bu |
249 |
247 |
0.07 |
|
|
|
|
Bg |
262 |
260 |
0.00 |
|
|
|
|
Ag |
415 |
413 |
0.00 |
|
|
|
|
Au |
723 |
719 |
4.52 |
|
|
|
|
Bg |
785 |
781 |
0.00 |
|
|
|
|
Ag |
829 |
824 |
0.00 |
|
|
|
|
Au |
926 |
920 |
1.55 |
|
|
|
|
Bu |
944 |
939 |
6.08 |
|
|
|
|
Bu |
1014 |
1008 |
0.17 |
|
|
|
|
Ag |
1062 |
1056 |
0.00 |
|
|
|
|
Ag |
1130 |
1123 |
0.00 |
|
|
|
|
Bg |
1155 |
1148 |
0.00 |
|
|
|
|
Au |
1235 |
1228 |
0.02 |
|
|
|
|
Bu |
1262 |
1254 |
6.51 |
|
|
|
|
Bg |
1280 |
1273 |
0.00 |
|
|
|
|
Ag |
1337 |
1330 |
0.00 |
|
|
|
|
Bu |
1347 |
1340 |
2.33 |
|
|
|
|
Ag |
1348 |
1340 |
0.00 |
|
|
|
|
Ag |
1411 |
1403 |
0.00 |
|
|
|
|
Bu |
1414 |
1406 |
1.88 |
|
|
|
|
Bg |
1419 |
1411 |
0.00 |
|
|
|
|
Au |
1421 |
1413 |
12.83 |
|
|
|
|
Ag |
1430 |
1422 |
0.00 |
|
|
|
|
Bu |
1436 |
1427 |
5.51 |
|
|
|
|
Ag |
2945 |
2928 |
0.00 |
|
|
|
|
Bu |
2952 |
2935 |
55.67 |
|
|
|
|
Bu |
2964 |
2947 |
73.16 |
|
|
|
|
Ag |
2965 |
2948 |
0.00 |
|
|
|
|
Bg |
2972 |
2955 |
0.00 |
|
|
|
|
Au |
2996 |
2978 |
30.04 |
|
|
|
|
Bg |
3039 |
3021 |
0.00 |
|
|
|
|
Au |
3041 |
3024 |
88.62 |
|
|
|
|
Ag |
3050 |
3032 |
0.00 |
|
|
|
|
Bu |
3050 |
3033 |
63.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28077.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27914.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.421 |
0.641 |
0.000 |
| C2 |
0.421 |
-0.641 |
0.000 |
| C3 |
0.421 |
1.920 |
0.000 |
| C4 |
-0.421 |
-1.920 |
0.000 |
| H5 |
-1.093 |
0.635 |
0.887 |
| H6 |
-1.093 |
0.635 |
-0.887 |
| H7 |
1.093 |
-0.635 |
0.887 |
| H8 |
1.093 |
-0.635 |
-0.887 |
| H9 |
-0.213 |
2.830 |
0.000 |
| H10 |
1.078 |
1.970 |
0.894 |
| H11 |
1.078 |
1.970 |
-0.894 |
| H12 |
0.213 |
-2.830 |
0.000 |
| H13 |
-1.078 |
-1.970 |
0.894 |
| H14 |
-1.078 |
-1.970 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5337 | 1.5309 | 2.5616 | 1.1135 | 1.1135 | 2.1700 | 2.1700 | 2.1986 | 2.1933 | 2.1933 | 3.5287 | 2.8376 | 2.8376 |
C2 | 1.5337 | | 2.5616 | 1.5309 | 2.1700 | 2.1700 | 1.1135 | 1.1135 | 3.5287 | 2.8376 | 2.8376 | 2.1986 | 2.1933 | 2.1933 | C3 | 1.5309 | 2.5616 | | 3.9317 | 2.1750 | 2.1750 | 2.7876 | 2.7876 | 1.1088 | 1.1105 | 1.1105 | 4.7549 | 4.2640 | 4.2640 | C4 | 2.5616 | 1.5309 | 3.9317 | | 2.7876 | 2.7876 | 2.1750 | 2.1750 | 4.7549 | 4.2640 | 4.2640 | 1.1088 | 1.1105 | 1.1105 | H5 | 1.1135 | 2.1700 | 2.1750 | 2.7876 | | 1.7745 | 2.5290 | 3.0895 | 2.5256 | 2.5487 | 3.1094 | 3.8082 | 2.6058 | 3.1564 | H6 | 1.1135 | 2.1700 | 2.1750 | 2.7876 | 1.7745 | | 3.0895 | 2.5290 | 2.5256 | 3.1094 | 2.5487 | 3.8082 | 3.1564 | 2.6058 | H7 | 2.1700 | 1.1135 | 2.7876 | 2.1750 | 2.5290 | 3.0895 | | 1.7745 | 3.8082 | 2.6058 | 3.1564 | 2.5256 | 2.5487 | 3.1094 | H8 | 2.1700 | 1.1135 | 2.7876 | 2.1750 | 3.0895 | 2.5290 | 1.7745 | | 3.8082 | 3.1564 | 2.6058 | 2.5256 | 3.1094 | 2.5487 | H9 | 2.1986 | 3.5287 | 1.1088 | 4.7549 | 2.5256 | 2.5256 | 3.8082 | 3.8082 | | 1.7901 | 1.7901 | 5.6761 | 4.9589 | 4.9589 | H10 | 2.1933 | 2.8376 | 1.1105 | 4.2640 | 2.5487 | 3.1094 | 2.6058 | 3.1564 | 1.7901 | | 1.7878 | 4.9589 | 4.4918 | 4.8345 | H11 | 2.1933 | 2.8376 | 1.1105 | 4.2640 | 3.1094 | 2.5487 | 3.1564 | 2.6058 | 1.7901 | 1.7878 | | 4.9589 | 4.8345 | 4.4918 | H12 | 3.5287 | 2.1986 | 4.7549 | 1.1088 | 3.8082 | 3.8082 | 2.5256 | 2.5256 | 5.6761 | 4.9589 | 4.9589 | | 1.7901 | 1.7901 | H13 | 2.8376 | 2.1933 | 4.2640 | 1.1105 | 2.6058 | 3.1564 | 2.5487 | 3.1094 | 4.9589 | 4.4918 | 4.8345 | 1.7901 | | 1.7878 | H14 | 2.8376 | 2.1933 | 4.2640 | 1.1105 | 3.1564 | 2.6058 | 3.1094 | 2.5487 | 4.9589 | 4.8345 | 4.4918 | 1.7901 | 1.7878 | |
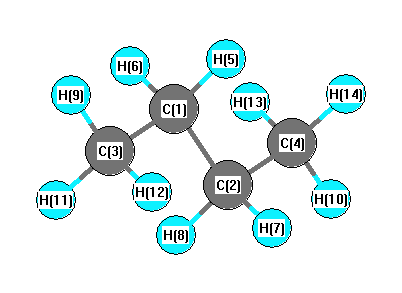 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.406 |
|
C1 |
C2 |
H7 |
109.079 |
| C1 |
C2 |
H8 |
109.079 |
|
C1 |
C3 |
H9 |
111.797 |
| C1 |
C3 |
H11 |
111.273 |
|
C1 |
C3 |
H12 |
30.833 |
| C2 |
C1 |
C3 |
113.406 |
|
C2 |
C1 |
H5 |
109.079 |
| C2 |
C1 |
H6 |
109.079 |
|
C2 |
C4 |
H10 |
17.169 |
| C2 |
C4 |
H13 |
111.273 |
|
C2 |
C4 |
H14 |
111.273 |
| C3 |
C1 |
H5 |
109.662 |
|
C3 |
C1 |
H6 |
109.662 |
| C4 |
C2 |
H7 |
109.662 |
|
C4 |
C2 |
H8 |
109.662 |
| H5 |
C1 |
H6 |
105.658 |
|
H7 |
C2 |
H8 |
105.658 |
| H9 |
C3 |
H11 |
107.529 |
|
H9 |
C3 |
H12 |
142.631 |
| H10 |
C4 |
H13 |
94.607 |
|
H10 |
C4 |
H14 |
114.698 |
| H11 |
C3 |
H12 |
94.067 |
|
H13 |
C4 |
H14 |
107.213 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.073 |
|
|
|
| 2 |
C |
-0.073 |
|
|
|
| 3 |
C |
-0.051 |
|
|
|
| 4 |
C |
-0.051 |
|
|
|
| 5 |
H |
0.019 |
|
|
|
| 6 |
H |
0.019 |
|
|
|
| 7 |
H |
0.019 |
|
|
|
| 8 |
H |
0.019 |
|
|
|
| 9 |
H |
0.026 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.026 |
|
|
|
| 13 |
H |
0.030 |
|
|
|
| 14 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.626 |
-0.323 |
0.000 |
| y |
-0.323 |
-28.899 |
0.000 |
| z |
0.000 |
0.000 |
-28.064 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.144 |
-0.323 |
0.000 |
| y |
-0.323 |
-0.555 |
0.000 |
| z |
0.000 |
0.000 |
0.699 |
|
| Polar |
|---|
| 3z2-r2 | 1.398 |
|---|
| x2-y2 | 0.274 |
|---|
| xy | -0.323 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.887 |
0.251 |
0.000 |
| y |
0.251 |
8.309 |
0.000 |
| z |
0.000 |
0.000 |
6.555 |
<r2> (average value of r
2) Å
2
| <r2> |
120.277 |
| (<r2>)1/2 |
10.967 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -158.213808 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 131.313313 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3053 |
3036 |
28.11 |
|
|
|
| 2 |
A |
3043 |
3025 |
4.88 |
|
|
|
| 3 |
A |
2989 |
2971 |
27.32 |
|
|
|
| 4 |
A |
2967 |
2950 |
27.10 |
|
|
|
| 5 |
A |
2948 |
2931 |
19.12 |
|
|
|
| 6 |
A |
1433 |
1425 |
0.19 |
|
|
|
| 7 |
A |
1427 |
1419 |
5.00 |
|
|
|
| 8 |
A |
1409 |
1401 |
0.06 |
|
|
|
| 9 |
A |
1346 |
1338 |
1.69 |
|
|
|
| 10 |
A |
1322 |
1315 |
0.91 |
|
|
|
| 11 |
A |
1261 |
1254 |
0.12 |
|
|
|
| 12 |
A |
1143 |
1136 |
0.01 |
|
|
|
| 13 |
A |
1081 |
1075 |
0.07 |
|
|
|
| 14 |
A |
957 |
952 |
0.38 |
|
|
|
| 15 |
A |
820 |
815 |
0.02 |
|
|
|
| 16 |
A |
774 |
769 |
2.03 |
|
|
|
| 17 |
A |
319 |
317 |
0.03 |
|
|
|
| 18 |
A |
254 |
252 |
0.03 |
|
|
|
| 19 |
A |
112 |
111 |
0.01 |
|
|
|
| 20 |
B |
3050 |
3032 |
43.93 |
|
|
|
| 21 |
B |
3043 |
3025 |
64.77 |
|
|
|
| 22 |
B |
2992 |
2975 |
30.76 |
|
|
|
| 23 |
B |
2966 |
2948 |
25.07 |
|
|
|
| 24 |
B |
2948 |
2931 |
32.51 |
|
|
|
| 25 |
B |
1428 |
1420 |
8.04 |
|
|
|
| 26 |
B |
1419 |
1410 |
8.03 |
|
|
|
| 27 |
B |
1403 |
1395 |
0.50 |
|
|
|
| 28 |
B |
1351 |
1343 |
7.95 |
|
|
|
| 29 |
B |
1307 |
1299 |
3.73 |
|
|
|
| 30 |
B |
1236 |
1229 |
0.96 |
|
|
|
| 31 |
B |
1110 |
1103 |
2.61 |
|
|
|
| 32 |
B |
959 |
954 |
2.46 |
|
|
|
| 33 |
B |
931 |
925 |
2.37 |
|
|
|
| 34 |
B |
727 |
722 |
4.74 |
|
|
|
| 35 |
B |
418 |
416 |
0.53 |
|
|
|
| 36 |
B |
215 |
214 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28079.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27916.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
