Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -158.215049 |
| Energy at 298.15K | -158.225574 |
| HF Energy | -158.215049 |
| Nuclear repulsion energy | 129.605297 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
122 |
122 |
0.00 |
|
|
|
|
Au |
223 |
222 |
0.00 |
|
|
|
|
Bu |
250 |
248 |
0.07 |
|
|
|
|
Bg |
264 |
262 |
0.00 |
|
|
|
|
Ag |
416 |
413 |
0.00 |
|
|
|
|
Au |
721 |
717 |
4.54 |
|
|
|
|
Bg |
783 |
779 |
0.00 |
|
|
|
|
Ag |
829 |
824 |
0.00 |
|
|
|
|
Au |
924 |
918 |
1.57 |
|
|
|
|
Bu |
944 |
939 |
6.05 |
|
|
|
|
Bu |
1014 |
1008 |
0.17 |
|
|
|
|
Ag |
1062 |
1056 |
0.00 |
|
|
|
|
Ag |
1130 |
1124 |
0.00 |
|
|
|
|
Bg |
1154 |
1147 |
0.00 |
|
|
|
|
Au |
1234 |
1227 |
0.02 |
|
|
|
|
Bu |
1261 |
1253 |
6.49 |
|
|
|
|
Bg |
1279 |
1271 |
0.00 |
|
|
|
|
Ag |
1337 |
1329 |
0.00 |
|
|
|
|
Ag |
1348 |
1340 |
0.00 |
|
|
|
|
Bu |
1348 |
1340 |
2.36 |
|
|
|
|
Ag |
1409 |
1401 |
0.00 |
|
|
|
|
Bu |
1413 |
1405 |
1.72 |
|
|
|
|
Bg |
1419 |
1411 |
0.00 |
|
|
|
|
Au |
1422 |
1413 |
12.80 |
|
|
|
|
Ag |
1430 |
1421 |
0.00 |
|
|
|
|
Bu |
1434 |
1426 |
5.69 |
|
|
|
|
Ag |
2943 |
2926 |
0.00 |
|
|
|
|
Bu |
2950 |
2933 |
56.72 |
|
|
|
|
Bu |
2962 |
2945 |
72.31 |
|
|
|
|
Ag |
2963 |
2946 |
0.00 |
|
|
|
|
Bg |
2971 |
2953 |
0.00 |
|
|
|
|
Au |
2994 |
2977 |
30.30 |
|
|
|
|
Bg |
3038 |
3020 |
0.00 |
|
|
|
|
Au |
3040 |
3023 |
88.46 |
|
|
|
|
Ag |
3048 |
3030 |
0.00 |
|
|
|
|
Bu |
3049 |
3031 |
63.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28064.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27901.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.420 |
0.641 |
0.000 |
| C2 |
0.420 |
-0.641 |
0.000 |
| C3 |
0.420 |
1.921 |
0.000 |
| C4 |
-0.420 |
-1.921 |
0.000 |
| H5 |
-1.093 |
0.635 |
0.887 |
| H6 |
-1.093 |
0.635 |
-0.887 |
| H7 |
1.093 |
-0.635 |
0.887 |
| H8 |
1.093 |
-0.635 |
-0.887 |
| H9 |
-0.214 |
2.830 |
0.000 |
| H10 |
1.077 |
1.971 |
0.894 |
| H11 |
1.077 |
1.971 |
-0.894 |
| H12 |
0.214 |
-2.830 |
0.000 |
| H13 |
-1.077 |
-1.971 |
0.894 |
| H14 |
-1.077 |
-1.971 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5336 | 1.5308 | 2.5618 | 1.1136 | 1.1136 | 2.1699 | 2.1699 | 2.1984 | 2.1935 | 2.1935 | 3.5288 | 2.8383 | 2.8383 |
C2 | 1.5336 | | 2.5618 | 1.5308 | 2.1699 | 2.1699 | 1.1136 | 1.1136 | 3.5288 | 2.8383 | 2.8383 | 2.1984 | 2.1935 | 2.1935 | C3 | 1.5308 | 2.5618 | | 3.9320 | 2.1748 | 2.1748 | 2.7880 | 2.7880 | 1.1088 | 1.1105 | 1.1105 | 4.7550 | 4.2647 | 4.2647 | C4 | 2.5618 | 1.5308 | 3.9320 | | 2.7880 | 2.7880 | 2.1748 | 2.1748 | 4.7550 | 4.2647 | 4.2647 | 1.1088 | 1.1105 | 1.1105 | H5 | 1.1136 | 2.1699 | 2.1748 | 2.7880 | | 1.7745 | 2.5291 | 3.0895 | 2.5252 | 2.5488 | 3.1095 | 3.8086 | 2.6067 | 3.1571 | H6 | 1.1136 | 2.1699 | 2.1748 | 2.7880 | 1.7745 | | 3.0895 | 2.5291 | 2.5252 | 3.1095 | 2.5488 | 3.8086 | 3.1571 | 2.6067 | H7 | 2.1699 | 1.1136 | 2.7880 | 2.1748 | 2.5291 | 3.0895 | | 1.7745 | 3.8086 | 2.6067 | 3.1571 | 2.5252 | 2.5488 | 3.1095 | H8 | 2.1699 | 1.1136 | 2.7880 | 2.1748 | 3.0895 | 2.5291 | 1.7745 | | 3.8086 | 3.1571 | 2.6067 | 2.5252 | 3.1095 | 2.5488 | H9 | 2.1984 | 3.5288 | 1.1088 | 4.7550 | 2.5252 | 2.5252 | 3.8086 | 3.8086 | | 1.7900 | 1.7900 | 5.6762 | 4.9595 | 4.9595 | H10 | 2.1935 | 2.8383 | 1.1105 | 4.2647 | 2.5488 | 3.1095 | 2.6067 | 3.1571 | 1.7900 | | 1.7878 | 4.9595 | 4.4929 | 4.8355 | H11 | 2.1935 | 2.8383 | 1.1105 | 4.2647 | 3.1095 | 2.5488 | 3.1571 | 2.6067 | 1.7900 | 1.7878 | | 4.9595 | 4.8355 | 4.4929 | H12 | 3.5288 | 2.1984 | 4.7550 | 1.1088 | 3.8086 | 3.8086 | 2.5252 | 2.5252 | 5.6762 | 4.9595 | 4.9595 | | 1.7900 | 1.7900 | H13 | 2.8383 | 2.1935 | 4.2647 | 1.1105 | 2.6067 | 3.1571 | 2.5488 | 3.1095 | 4.9595 | 4.4929 | 4.8355 | 1.7900 | | 1.7878 | H14 | 2.8383 | 2.1935 | 4.2647 | 1.1105 | 3.1571 | 2.6067 | 3.1095 | 2.5488 | 4.9595 | 4.8355 | 4.4929 | 1.7900 | 1.7878 | |
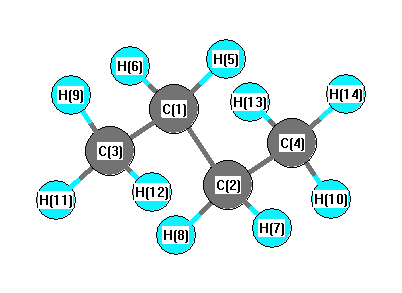 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.436 |
|
C1 |
C2 |
H7 |
109.081 |
| C1 |
C2 |
H8 |
109.081 |
|
C1 |
C3 |
H9 |
111.793 |
| C1 |
C3 |
H11 |
111.291 |
|
C1 |
C3 |
H12 |
30.827 |
| C2 |
C1 |
C3 |
113.436 |
|
C2 |
C1 |
H5 |
109.081 |
| C2 |
C1 |
H6 |
109.081 |
|
C2 |
C4 |
H10 |
17.164 |
| C2 |
C4 |
H13 |
111.291 |
|
C2 |
C4 |
H14 |
111.291 |
| C3 |
C1 |
H5 |
109.650 |
|
C3 |
C1 |
H6 |
109.650 |
| C4 |
C2 |
H7 |
109.650 |
|
C4 |
C2 |
H8 |
109.650 |
| H5 |
C1 |
H6 |
105.647 |
|
H7 |
C2 |
H8 |
105.647 |
| H9 |
C3 |
H11 |
107.516 |
|
H9 |
C3 |
H12 |
142.620 |
| H10 |
C4 |
H13 |
94.631 |
|
H10 |
C4 |
H14 |
114.718 |
| H11 |
C3 |
H12 |
94.089 |
|
H13 |
C4 |
H14 |
107.204 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.073 |
|
|
|
| 2 |
C |
-0.073 |
|
|
|
| 3 |
C |
-0.051 |
|
|
|
| 4 |
C |
-0.051 |
|
|
|
| 5 |
H |
0.019 |
|
|
|
| 6 |
H |
0.019 |
|
|
|
| 7 |
H |
0.019 |
|
|
|
| 8 |
H |
0.019 |
|
|
|
| 9 |
H |
0.026 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.026 |
|
|
|
| 13 |
H |
0.030 |
|
|
|
| 14 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.626 |
-0.323 |
0.000 |
| y |
-0.323 |
-28.897 |
0.000 |
| z |
0.000 |
0.000 |
-28.065 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.145 |
-0.323 |
0.000 |
| y |
-0.323 |
-0.551 |
0.000 |
| z |
0.000 |
0.000 |
0.697 |
|
| Polar |
|---|
| 3z2-r2 | 1.393 |
|---|
| x2-y2 | 0.270 |
|---|
| xy | -0.323 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.888 |
0.251 |
0.000 |
| y |
0.251 |
8.309 |
0.000 |
| z |
0.000 |
0.000 |
6.556 |
<r2> (average value of r
2) Å
2
| <r2> |
120.292 |
| (<r2>)1/2 |
10.968 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -158.213816 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3052 |
3034 |
25.78 |
|
|
|
| 2 |
A |
3042 |
3025 |
6.79 |
|
|
|
| 3 |
A |
2990 |
2973 |
27.58 |
|
|
|
| 4 |
A |
2967 |
2949 |
26.93 |
|
|
|
| 5 |
A |
2949 |
2932 |
19.42 |
|
|
|
| 6 |
A |
1433 |
1424 |
0.05 |
|
|
|
| 7 |
A |
1427 |
1419 |
5.06 |
|
|
|
| 8 |
A |
1409 |
1401 |
0.08 |
|
|
|
| 9 |
A |
1345 |
1338 |
1.80 |
|
|
|
| 10 |
A |
1324 |
1317 |
0.88 |
|
|
|
| 11 |
A |
1262 |
1255 |
0.12 |
|
|
|
| 12 |
A |
1144 |
1137 |
0.01 |
|
|
|
| 13 |
A |
1081 |
1075 |
0.07 |
|
|
|
| 14 |
A |
958 |
952 |
0.37 |
|
|
|
| 15 |
A |
820 |
815 |
0.01 |
|
|
|
| 16 |
A |
775 |
770 |
2.02 |
|
|
|
| 17 |
A |
319 |
317 |
0.03 |
|
|
|
| 18 |
A |
247 |
246 |
0.02 |
|
|
|
| 19 |
A |
115 |
114 |
0.01 |
|
|
|
| 20 |
B |
3048 |
3031 |
48.38 |
|
|
|
| 21 |
B |
3042 |
3024 |
63.45 |
|
|
|
| 22 |
B |
2993 |
2976 |
28.44 |
|
|
|
| 23 |
B |
2964 |
2947 |
24.48 |
|
|
|
| 24 |
B |
2949 |
2932 |
32.43 |
|
|
|
| 25 |
B |
1429 |
1420 |
8.51 |
|
|
|
| 26 |
B |
1419 |
1411 |
7.61 |
|
|
|
| 27 |
B |
1404 |
1396 |
0.46 |
|
|
|
| 28 |
B |
1351 |
1343 |
8.06 |
|
|
|
| 29 |
B |
1308 |
1300 |
3.68 |
|
|
|
| 30 |
B |
1237 |
1229 |
0.93 |
|
|
|
| 31 |
B |
1111 |
1105 |
2.59 |
|
|
|
| 32 |
B |
959 |
954 |
2.48 |
|
|
|
| 33 |
B |
932 |
927 |
2.38 |
|
|
|
| 34 |
B |
728 |
723 |
4.75 |
|
|
|
| 35 |
B |
421 |
418 |
0.53 |
|
|
|
| 36 |
B |
217 |
216 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28084.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27922.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
