return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-156.736909
Energy at 298.15K-156.744873
HF Energy-156.117755
Nuclear repulsion energy116.178297
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3256 3124 17.46      
2 A 3167 3038 4.55      
3 A 3153 3025 28.93      
4 A 3142 3015 33.95      
5 A 3136 3009 32.63      
6 A 3097 2972 9.81      
7 A 3055 2931 22.72      
8 A 3048 2924 36.53      
9 A 1726 1656 5.60      
10 A 1505 1444 3.80      
11 A 1496 1435 4.71      
12 A 1482 1422 2.56      
13 A 1458 1399 1.59      
14 A 1416 1359 0.70      
15 A 1355 1300 3.92      
16 A 1313 1260 1.95      
17 A 1294 1241 0.16      
18 A 1205 1156 0.39      
19 A 1102 1058 3.40      
20 A 1055 1012 1.44      
21 A 1019 977 12.23      
22 A 986 946 1.38      
23 A 929 892 30.67      
24 A 872 837 1.14      
25 A 794 761 1.81      
26 A 647 621 7.76      
27 A 431 414 0.58      
28 A 317 304 0.41      
29 A 241 231 0.03      
30 A 105 101 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 23900.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 22930.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.74041 0.13664 0.13359

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.729 -0.243 -0.303
C2 0.546 0.519 0.320
C3 -0.721 -0.307 0.341
C4 -1.868 0.026 -0.282
H5 -0.674 -1.252 0.905
H6 1.512 -0.513 -1.352
H7 2.650 0.368 -0.286
H8 1.932 -1.178 0.252
H9 0.809 0.810 1.357
H10 0.363 1.457 -0.238
H11 -1.956 0.958 -0.856
H12 -2.754 -0.619 -0.237

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.53872.53413.60732.87291.10541.10511.10622.17012.18123.91524.4991
C21.53871.51242.53632.22902.18952.19472.19231.10851.10652.79933.5344
C32.53411.51241.34751.10152.81003.49452.79392.14852.15012.13552.1365
C43.60732.53631.34752.11393.58604.53094.02223.23482.65121.09821.0964
H52.87292.22901.10152.11393.22803.88512.68802.57963.11823.10332.4556
H61.10542.18952.81003.58603.22801.79111.78693.09582.53773.79954.4102
H71.10512.19473.49454.53093.88511.79111.78742.50712.53294.67855.4932
H81.10622.19232.79394.02222.68801.78691.78742.53663.10554.57274.7445
H92.17011.10852.14853.23482.57963.09582.50712.53661.77803.54434.1557
H102.18121.10652.15012.65123.11822.53772.53293.10551.77802.45173.7450
H113.91522.79932.13551.09823.10333.79954.67854.57273.54432.45171.8724
H124.49913.53442.13651.09642.45564.41025.49324.74454.15573.74501.8724

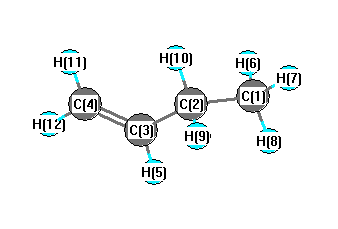
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.307 C1 C2 H9 109.043
C1 C2 H10 110.016 C2 C1 H6 110.736
C2 C1 H7 111.165 C2 C1 H8 110.902
C2 C3 C4 124.858 C2 C3 H5 116.138
C3 C2 H9 109.155 C3 C2 H10 109.388
C3 C4 H11 121.322 C3 C4 H12 121.558
C4 C3 H5 118.999 H6 C1 H7 108.252
H6 C1 H8 107.795 H7 C1 H8 107.860
H9 C2 H10 106.775 H11 C4 H12 117.120
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability