return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-156.732690
Energy at 298.15K-156.740665
HF Energy-156.117974
Nuclear repulsion energy116.252474
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3266 3125 16.56      
2 A 3176 3038 4.37      
3 A 3162 3025 29.64      
4 A 3155 3018 31.43      
5 A 3148 3012 29.32      
6 A 3109 2975 9.82      
7 A 3064 2932 21.99      
8 A 3058 2926 34.60      
9 A 1741 1665 4.28      
10 A 1508 1442 4.01      
11 A 1498 1433 4.91      
12 A 1485 1421 2.54      
13 A 1462 1399 1.67      
14 A 1420 1358 0.82      
15 A 1359 1300 3.14      
16 A 1318 1261 2.14      
17 A 1297 1240 0.14      
18 A 1207 1155 0.37      
19 A 1105 1058 3.44      
20 A 1058 1012 1.50      
21 A 1023 979 11.92      
22 A 988 946 1.33      
23 A 940 899 31.17      
24 A 875 837 1.21      
25 A 795 761 1.88      
26 A 650 622 8.05      
27 A 433 414 0.57      
28 A 318 304 0.41      
29 A 241 230 0.03      
30 A 106 101 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 23981.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22943.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.74075 0.13683 0.13379

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.728 -0.243 -0.303
C2 0.545 0.519 0.320
C3 -0.722 -0.306 0.342
C4 -1.866 0.026 -0.282
H5 -0.676 -1.250 0.907
H6 1.511 -0.514 -1.352
H7 2.649 0.367 -0.287
H8 1.931 -1.178 0.252
H9 0.808 0.811 1.356
H10 0.363 1.456 -0.239
H11 -1.953 0.956 -0.859
H12 -2.752 -0.618 -0.237

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.53852.53393.60402.87331.10491.10461.10572.16962.18043.91114.4959
C21.53851.51202.53352.22822.18892.19422.19171.10811.10612.79643.5316
C32.53391.51201.34501.10112.80973.49382.79382.14822.14932.13312.1339
C43.60402.53351.34502.11153.58254.52734.01893.23252.64891.09781.0960
H52.87332.22821.10112.11153.22863.88482.68902.57863.11703.10082.4530
H61.10492.18892.80973.58253.22861.79031.78603.09472.53673.79474.4068
H71.10462.19423.49384.52733.88481.79031.78652.50632.53194.67415.4896
H81.10572.19172.79384.01892.68901.78601.78652.53623.10424.56874.7415
H92.16961.10812.14823.23252.57863.09472.50632.53621.77703.54224.1533
H102.18041.10612.14932.64893.11702.53672.53193.10421.77702.44923.7423
H113.91112.79642.13311.09783.10083.79474.67414.56873.54222.44921.8714
H124.49593.53162.13391.09602.45304.40685.48964.74154.15333.74231.8714

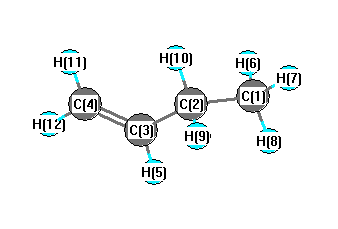
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.332 C1 C2 H9 109.042
C1 C2 H10 109.994 C2 C1 H6 110.733
C2 C1 H7 111.170 C2 C1 H8 110.906
C2 C3 C4 124.847 C2 C3 H5 116.131
C3 C2 H9 109.180 C3 C2 H10 109.382
C3 C4 H11 121.343 C3 C4 H12 121.567
C4 C3 H5 119.017 H6 C1 H7 108.252
H6 C1 H8 107.791 H7 C1 H8 107.858
H9 C2 H10 106.749 H11 C4 H12 117.089
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability