Jump to
S1C2
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Geometric Data calculated at QCISD(T)/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -155.534447 |
| Energy at 298.15K | -155.539699 |
| HF Energy | -154.926148 |
| Nuclear repulsion energy | 103.608921 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3251 |
3098 |
|
|
|
|
| 2 |
A |
3168 |
3020 |
|
|
|
|
| 3 |
A |
3150 |
3002 |
|
|
|
|
| 4 |
A |
1670 |
1591 |
|
|
|
|
| 5 |
A |
1457 |
1389 |
|
|
|
|
| 6 |
A |
1321 |
1259 |
|
|
|
|
| 7 |
A |
1052 |
1003 |
|
|
|
|
| 8 |
A |
983 |
937 |
|
|
|
|
| 9 |
A |
901 |
859 |
|
|
|
|
| 10 |
A |
887 |
845 |
|
|
|
|
| 11 |
A |
727 |
693 |
|
|
|
|
| 12 |
A |
269 |
256 |
|
|
|
|
| 13 |
A |
156 |
149 |
|
|
|
|
| 14 |
B |
3249 |
3097 |
|
|
|
|
| 15 |
B |
3156 |
3008 |
|
|
|
|
| 16 |
B |
3146 |
2998 |
|
|
|
|
| 17 |
B |
1672 |
1593 |
|
|
|
|
| 18 |
B |
1421 |
1355 |
|
|
|
|
| 19 |
B |
1288 |
1228 |
|
|
|
|
| 20 |
B |
1091 |
1040 |
|
|
|
|
| 21 |
B |
1001 |
954 |
|
|
|
|
| 22 |
B |
903 |
861 |
|
|
|
|
| 23 |
B |
601 |
572 |
|
|
|
|
| 24 |
B |
461 |
439 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18489.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 17622.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.107 |
0.735 |
0.566 |
| C2 |
-0.107 |
-0.735 |
0.566 |
| C3 |
-0.107 |
1.545 |
-0.502 |
| C4 |
0.107 |
-1.545 |
-0.502 |
| H5 |
0.443 |
1.182 |
1.514 |
| H6 |
-0.443 |
-1.182 |
1.514 |
| H7 |
0.088 |
2.623 |
-0.445 |
| H8 |
-0.493 |
1.149 |
-1.452 |
| H9 |
-0.088 |
-2.623 |
-0.445 |
| H10 |
0.493 |
-1.149 |
-1.452 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4852 | 1.3574 | 2.5174 | 1.1011 | 2.2082 | 2.1420 | 2.1453 | 3.5123 | 2.7871 |
C2 | 1.4852 | | 2.5174 | 1.3574 | 2.2082 | 1.1011 | 3.5123 | 2.7871 | 2.1420 | 2.1453 | C3 | 1.3574 | 2.5174 | | 3.0970 | 2.1216 | 3.4078 | 1.0975 | 1.0988 | 4.1685 | 2.9186 | C4 | 2.5174 | 1.3574 | 3.0970 | | 3.4078 | 2.1216 | 4.1685 | 2.9186 | 1.0975 | 1.0988 | H5 | 1.1011 | 2.2082 | 2.1216 | 3.4078 | | 2.5243 | 2.4582 | 3.1105 | 4.3126 | 3.7725 | H6 | 2.2082 | 1.1011 | 3.4078 | 2.1216 | 2.5243 | | 4.3126 | 3.7725 | 2.4582 | 3.1105 | H7 | 2.1420 | 3.5123 | 1.0975 | 4.1685 | 2.4582 | 4.3126 | | 1.8775 | 5.2494 | 3.9251 | H8 | 2.1453 | 2.7871 | 1.0988 | 2.9186 | 3.1105 | 3.7725 | 1.8775 | | 3.9251 | 2.5005 | H9 | 3.5123 | 2.1420 | 4.1685 | 1.0975 | 4.3126 | 2.4582 | 5.2494 | 3.9251 | | 1.8775 | H10 | 2.7871 | 2.1453 | 2.9186 | 1.0988 | 3.7725 | 3.1105 | 3.9251 | 2.5005 | 1.8775 | |
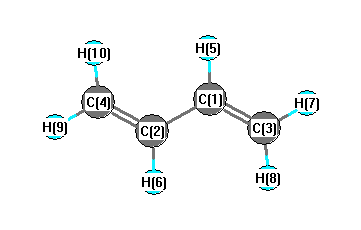 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
124.586 |
|
C1 |
C2 |
H6 |
116.467 |
| C1 |
C3 |
H7 |
121.144 |
|
C1 |
C3 |
H8 |
121.360 |
| C2 |
C1 |
C3 |
124.586 |
|
C2 |
C1 |
H5 |
116.467 |
| C2 |
C4 |
H9 |
121.144 |
|
C2 |
C4 |
H10 |
121.360 |
| C3 |
C1 |
H5 |
118.934 |
|
C4 |
C2 |
H6 |
118.934 |
| H7 |
C3 |
H8 |
117.492 |
|
H9 |
C4 |
H10 |
117.492 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
