Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -155.512725 |
| Energy at 298.15K | -155.518037 |
| HF Energy | -154.927249 |
| Nuclear repulsion energy | 103.821650 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3267 |
3135 |
5.59 |
|
|
|
| 2 |
A |
3187 |
3057 |
8.71 |
|
|
|
| 3 |
A |
3168 |
3039 |
15.72 |
|
|
|
| 4 |
A |
1709 |
1640 |
1.39 |
|
|
|
| 5 |
A |
1470 |
1411 |
7.44 |
|
|
|
| 6 |
A |
1336 |
1281 |
0.01 |
|
|
|
| 7 |
A |
1065 |
1022 |
0.00 |
|
|
|
| 8 |
A |
1002 |
962 |
2.93 |
|
|
|
| 9 |
A |
930 |
893 |
5.22 |
|
|
|
| 10 |
A |
894 |
858 |
0.45 |
|
|
|
| 11 |
A |
743 |
713 |
3.67 |
|
|
|
| 12 |
A |
274 |
263 |
0.00 |
|
|
|
| 13 |
A |
160 |
154 |
0.07 |
|
|
|
| 14 |
B |
3266 |
3133 |
19.54 |
|
|
|
| 15 |
B |
3175 |
3046 |
12.54 |
|
|
|
| 16 |
B |
3164 |
3036 |
3.43 |
|
|
|
| 17 |
B |
1703 |
1634 |
2.80 |
|
|
|
| 18 |
B |
1435 |
1377 |
0.75 |
|
|
|
| 19 |
B |
1304 |
1251 |
0.76 |
|
|
|
| 20 |
B |
1102 |
1057 |
3.22 |
|
|
|
| 21 |
B |
1021 |
979 |
29.44 |
|
|
|
| 22 |
B |
932 |
894 |
53.81 |
|
|
|
| 23 |
B |
613 |
588 |
6.18 |
|
|
|
| 24 |
B |
463 |
445 |
10.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18691.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 17932.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.113 |
0.734 |
0.560 |
| C2 |
-0.113 |
-0.734 |
0.560 |
| C3 |
-0.113 |
1.545 |
-0.497 |
| C4 |
0.113 |
-1.545 |
-0.497 |
| H5 |
0.472 |
1.173 |
1.502 |
| H6 |
-0.472 |
-1.173 |
1.502 |
| H7 |
0.091 |
2.621 |
-0.440 |
| H8 |
-0.519 |
1.157 |
-1.439 |
| H9 |
-0.091 |
-2.621 |
-0.440 |
| H10 |
0.519 |
-1.157 |
-1.439 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4857 | 1.3511 | 2.5125 | 1.0995 | 2.2064 | 2.1354 | 2.1389 | 3.5068 | 2.7818 |
C2 | 1.4857 | | 2.5125 | 1.3511 | 2.2064 | 1.0995 | 3.5068 | 2.7818 | 2.1354 | 2.1389 | C3 | 1.3511 | 2.5125 | | 3.0993 | 2.1153 | 3.3932 | 1.0960 | 1.0972 | 4.1668 | 2.9314 | C4 | 2.5125 | 1.3511 | 3.0993 | | 3.3932 | 2.1153 | 4.1668 | 2.9314 | 1.0960 | 1.0972 | H5 | 1.0995 | 2.2064 | 2.1153 | 3.3932 | | 2.5296 | 2.4518 | 3.1035 | 4.2992 | 3.7527 | H6 | 2.2064 | 1.0995 | 3.3932 | 2.1153 | 2.5296 | | 4.2992 | 3.7527 | 2.4518 | 3.1035 | H7 | 2.1354 | 3.5068 | 1.0960 | 4.1668 | 2.4518 | 4.2992 | | 1.8741 | 5.2449 | 3.9314 | H8 | 2.1389 | 2.7818 | 1.0972 | 2.9314 | 3.1035 | 3.7527 | 1.8741 | | 3.9314 | 2.5366 | H9 | 3.5068 | 2.1354 | 4.1668 | 1.0960 | 4.2992 | 2.4518 | 5.2449 | 3.9314 | | 1.8741 | H10 | 2.7818 | 2.1389 | 2.9314 | 1.0972 | 3.7527 | 3.1035 | 3.9314 | 2.5366 | 1.8741 | |
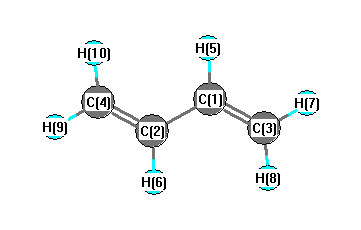 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
124.606 |
|
C1 |
C2 |
H6 |
116.391 |
| C1 |
C3 |
H7 |
121.172 |
|
C1 |
C3 |
H8 |
121.417 |
| C2 |
C1 |
C3 |
124.606 |
|
C2 |
C1 |
H5 |
116.391 |
| C2 |
C4 |
H9 |
121.172 |
|
C2 |
C4 |
H10 |
121.417 |
| C3 |
C1 |
H5 |
118.992 |
|
C4 |
C2 |
H6 |
118.992 |
| H7 |
C3 |
H8 |
117.408 |
|
H9 |
C4 |
H10 |
117.408 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
