Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -155.772047 |
| Energy at 298.15K | -155.777246 |
| HF Energy | -155.772047 |
| Nuclear repulsion energy | 103.872497 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3158 |
5.68 |
|
|
|
| 2 |
A |
3086 |
3068 |
4.76 |
|
|
|
| 3 |
A |
3074 |
3056 |
23.09 |
|
|
|
| 4 |
A |
1634 |
1625 |
1.78 |
|
|
|
| 5 |
A |
1399 |
1390 |
7.82 |
|
|
|
| 6 |
A |
1293 |
1285 |
0.42 |
|
|
|
| 7 |
A |
1032 |
1026 |
0.06 |
|
|
|
| 8 |
A |
983 |
978 |
1.23 |
|
|
|
| 9 |
A |
889 |
884 |
2.49 |
|
|
|
| 10 |
A |
870 |
865 |
0.92 |
|
|
|
| 11 |
A |
730 |
726 |
2.11 |
|
|
|
| 12 |
A |
266 |
265 |
0.00 |
|
|
|
| 13 |
A |
147 |
146 |
0.30 |
|
|
|
| 14 |
B |
3176 |
3157 |
13.59 |
|
|
|
| 15 |
B |
3078 |
3060 |
4.29 |
|
|
|
| 16 |
B |
3066 |
3048 |
8.50 |
|
|
|
| 17 |
B |
1650 |
1640 |
2.69 |
|
|
|
| 18 |
B |
1364 |
1356 |
2.34 |
|
|
|
| 19 |
B |
1259 |
1252 |
0.20 |
|
|
|
| 20 |
B |
1063 |
1056 |
5.89 |
|
|
|
| 21 |
B |
991 |
985 |
25.08 |
|
|
|
| 22 |
B |
890 |
885 |
56.81 |
|
|
|
| 23 |
B |
589 |
585 |
5.31 |
|
|
|
| 24 |
B |
462 |
459 |
10.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18082.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 17977.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.086 |
0.730 |
0.559 |
| C2 |
-0.086 |
-0.730 |
0.559 |
| C3 |
-0.086 |
1.551 |
-0.499 |
| C4 |
0.086 |
-1.551 |
-0.499 |
| H5 |
0.360 |
1.178 |
1.531 |
| H6 |
-0.360 |
-1.178 |
1.531 |
| H7 |
0.084 |
2.634 |
-0.413 |
| H8 |
-0.417 |
1.168 |
-1.478 |
| H9 |
-0.084 |
-2.634 |
-0.413 |
| H10 |
0.417 |
-1.168 |
-1.478 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4694 | 1.3507 | 2.5139 | 1.1044 | 2.1868 | 2.1379 | 2.1437 | 3.5050 | 2.8037 |
C2 | 1.4694 | | 2.5139 | 1.3507 | 2.1868 | 1.1044 | 3.5050 | 2.8037 | 2.1379 | 2.1437 | C3 | 1.3507 | 2.5139 | | 3.1061 | 2.1115 | 3.4118 | 1.0996 | 1.1014 | 4.1851 | 2.9332 | C4 | 2.5139 | 1.3507 | 3.1061 | | 3.4118 | 2.1115 | 4.1851 | 2.9332 | 1.0996 | 1.1014 | H5 | 1.1044 | 2.1868 | 2.1115 | 3.4118 | | 2.4639 | 2.4441 | 3.1072 | 4.3016 | 3.8156 | H6 | 2.1868 | 1.1044 | 3.4118 | 2.1115 | 2.4639 | | 4.3016 | 3.8156 | 2.4441 | 3.1072 | H7 | 2.1379 | 3.5050 | 1.0996 | 4.1851 | 2.4441 | 4.3016 | | 1.8791 | 5.2698 | 3.9621 | H8 | 2.1437 | 2.8037 | 1.1014 | 2.9332 | 3.1072 | 3.8156 | 1.8791 | | 3.9621 | 2.4812 | H9 | 3.5050 | 2.1379 | 4.1851 | 1.0996 | 4.3016 | 2.4441 | 5.2698 | 3.9621 | | 1.8791 | H10 | 2.8037 | 2.1437 | 2.9332 | 1.1014 | 3.8156 | 3.1072 | 3.9621 | 2.4812 | 1.8791 | |
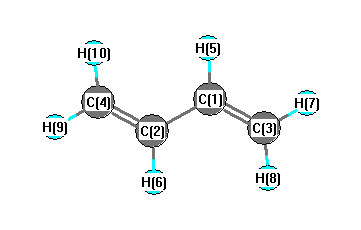 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
126.057 |
|
C1 |
C2 |
H6 |
115.622 |
| C1 |
C3 |
H7 |
121.169 |
|
C1 |
C3 |
H8 |
121.578 |
| C2 |
C1 |
C3 |
126.057 |
|
C2 |
C1 |
H5 |
115.622 |
| C2 |
C4 |
H9 |
121.169 |
|
C2 |
C4 |
H10 |
121.578 |
| C3 |
C1 |
H5 |
118.305 |
|
C4 |
C2 |
H6 |
118.305 |
| H7 |
C3 |
H8 |
117.247 |
|
H9 |
C4 |
H10 |
117.247 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.056 |
|
|
|
| 2 |
C |
-0.056 |
|
|
|
| 3 |
C |
0.003 |
|
|
|
| 4 |
C |
0.003 |
|
|
|
| 5 |
H |
-0.001 |
|
|
|
| 6 |
H |
-0.001 |
|
|
|
| 7 |
H |
0.030 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
| 9 |
H |
0.030 |
|
|
|
| 10 |
H |
0.024 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.116 |
0.116 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.813 |
0.558 |
0.000 |
| y |
0.558 |
-23.208 |
0.000 |
| z |
0.000 |
0.000 |
-23.195 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.612 |
0.558 |
0.000 |
| y |
0.558 |
2.296 |
0.000 |
| z |
0.000 |
0.000 |
2.316 |
|
| Polar |
|---|
| 3z2-r2 | 4.631 |
|---|
| x2-y2 | -4.605 |
|---|
| xy | 0.558 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.154 |
0.059 |
0.000 |
| y |
0.059 |
9.835 |
0.000 |
| z |
0.000 |
0.000 |
7.197 |
<r2> (average value of r
2) Å
2
| <r2> |
87.008 |
| (<r2>)1/2 |
9.328 |
