return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHCCH2CH3 (1-Butyne)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-155.516016
Energy at 298.15K-155.520877
HF Energy-154.907937
Nuclear repulsion energy101.550761
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3457 3295        
2 A' 3136 2989        
3 A' 3052 2909        
4 A' 3048 2905        
5 A' 2157 2055        
6 A' 1493 1423        
7 A' 1468 1399        
8 A' 1405 1339        
9 A' 1347 1284        
10 A' 1090 1039        
11 A' 1037 989        
12 A' 848 808        
13 A' 595 567        
14 A' 491 468        
15 A' 186 177        
16 A" 3144 2997        
17 A" 3094 2948        
18 A" 1484 1415        
19 A" 1284 1223        
20 A" 1103 1051        
21 A" 780 744        
22 A" 591 563        
23 A" 338 322        
24 A" 222 212        

Unscaled Zero Point Vibrational Energy (zpe) 18424.0 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 17559.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.89528 0.14749 0.13303

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.642 -1.900 0.000
C2 0.000 -0.850 0.000
C3 0.752 0.427 0.000
C4 -0.176 1.658 0.000
H5 -1.202 -2.821 0.000
H6 1.414 0.451 0.888
H7 1.414 0.451 -0.888
H8 0.421 2.589 0.000
H9 -0.825 1.657 -0.895
H10 -0.825 1.657 0.895

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.23052.71273.58781.07883.24673.24674.61253.67193.6719
C21.23051.48232.51392.30932.11672.11673.46432.78672.7867
C32.71271.48231.54193.79151.10751.10752.18692.19152.1915
C43.58782.51391.54194.59532.18472.18471.10611.10561.1056
H51.07882.30933.79154.59534.28294.28295.64854.58244.5824
H63.24672.11671.10752.18474.28291.77642.51863.10592.5427
H73.24672.11671.10752.18474.28291.77642.51862.54273.1059
H84.61253.46432.18691.10615.64852.51862.51861.79521.7952
H93.67192.78672.19151.10564.58243.10592.54271.79521.7906
H103.67192.78672.19151.10564.58242.54273.10591.79521.7906

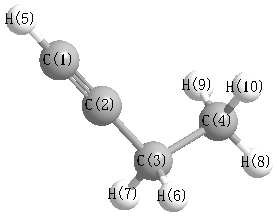
picture of 1-Butyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 179.064 C2 C1 H5 179.855
C2 C3 C4 112.443 C2 C3 H6 108.773
C2 C3 H7 108.773 C3 C4 H8 110.265
C3 C4 H9 110.656 C3 C4 H10 110.656
C4 C3 H6 110.016 C4 C3 H7 110.016
H6 C3 H7 106.639 H8 C4 H9 108.521
H8 C4 H10 108.521 H9 C4 H10 108.147
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability