Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.688357 |
| Energy at 298.15K | |
| HF Energy | -191.688357 |
| Nuclear repulsion energy | 102.135126 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3174 |
3156 |
4.36 |
74.03 |
0.55 |
0.71 |
| 2 |
A' |
3113 |
3095 |
2.52 |
122.19 |
0.30 |
0.47 |
| 3 |
A' |
3068 |
3050 |
4.96 |
80.92 |
0.10 |
0.19 |
| 4 |
A' |
2765 |
2749 |
122.19 |
116.22 |
0.29 |
0.44 |
| 5 |
A' |
1736 |
1726 |
192.09 |
49.67 |
0.45 |
0.62 |
| 6 |
A' |
1635 |
1625 |
2.58 |
26.21 |
0.20 |
0.33 |
| 7 |
A' |
1382 |
1374 |
11.29 |
4.93 |
0.65 |
0.79 |
| 8 |
A' |
1325 |
1317 |
6.04 |
20.97 |
0.47 |
0.64 |
| 9 |
A' |
1238 |
1231 |
6.11 |
5.67 |
0.32 |
0.48 |
| 10 |
A' |
1126 |
1120 |
28.20 |
9.28 |
0.58 |
0.74 |
| 11 |
A' |
889 |
884 |
17.15 |
2.18 |
0.15 |
0.26 |
| 12 |
A' |
555 |
552 |
5.50 |
6.13 |
0.34 |
0.51 |
| 13 |
A' |
303 |
301 |
7.54 |
1.12 |
0.35 |
0.52 |
| 14 |
A" |
1003 |
997 |
8.34 |
1.16 |
0.75 |
0.86 |
| 15 |
A" |
983 |
977 |
8.00 |
4.70 |
0.75 |
0.86 |
| 16 |
A" |
944 |
939 |
27.13 |
0.35 |
0.75 |
0.86 |
| 17 |
A" |
592 |
589 |
6.64 |
2.85 |
0.75 |
0.86 |
| 18 |
A" |
171 |
170 |
2.41 |
1.80 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13001.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12925.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.153 |
-0.748 |
0.000 |
| C2 |
0.000 |
0.723 |
0.000 |
| C3 |
1.223 |
1.292 |
0.000 |
| O4 |
-1.227 |
-1.332 |
0.000 |
| H5 |
0.832 |
-1.310 |
0.000 |
| H6 |
-0.929 |
1.315 |
0.000 |
| H7 |
1.363 |
2.383 |
0.000 |
| H8 |
2.131 |
0.666 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4781 | 2.4603 | 1.2234 | 1.1336 | 2.2040 | 3.4785 | 2.6859 |
C2 | 1.4781 | | 1.3489 | 2.3936 | 2.1962 | 1.1021 | 2.1485 | 2.1318 | C3 | 2.4603 | 1.3489 | | 3.5905 | 2.6316 | 2.1521 | 1.0998 | 1.1032 | O4 | 1.2234 | 2.3936 | 3.5905 | | 2.0589 | 2.6641 | 4.5296 | 3.9081 | H5 | 1.1336 | 2.1962 | 2.6316 | 2.0589 | | 3.1610 | 3.7313 | 2.3653 | H6 | 2.2040 | 1.1021 | 2.1521 | 2.6641 | 3.1610 | | 2.5294 | 3.1284 | H7 | 3.4785 | 2.1485 | 1.0998 | 4.5296 | 3.7313 | 2.5294 | | 1.8806 | H8 | 2.6859 | 2.1318 | 1.1032 | 3.9081 | 2.3653 | 3.1284 | 1.8806 | |
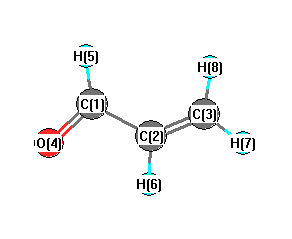 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.913 |
|
C1 |
C2 |
H6 |
116.584 |
| C2 |
C1 |
O4 |
124.487 |
|
C2 |
C1 |
H5 |
113.820 |
| C2 |
C3 |
H7 |
122.337 |
|
C2 |
C3 |
H8 |
120.440 |
| C3 |
C2 |
H6 |
122.503 |
|
O4 |
C1 |
H5 |
121.693 |
| H7 |
C3 |
H8 |
117.222 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.146 |
|
|
|
| 2 |
C |
-0.080 |
|
|
|
| 3 |
C |
0.030 |
|
|
|
| 4 |
O |
-0.176 |
|
|
|
| 5 |
H |
-0.021 |
|
|
|
| 6 |
H |
0.012 |
|
|
|
| 7 |
H |
0.050 |
|
|
|
| 8 |
H |
0.040 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.103 |
1.950 |
0.000 |
2.868 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.105 |
-2.083 |
0.000 |
| y |
-2.083 |
-23.751 |
0.000 |
| z |
0.000 |
0.000 |
-24.545 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.043 |
-2.083 |
0.000 |
| y |
-2.083 |
0.574 |
0.000 |
| z |
0.000 |
0.000 |
-0.617 |
|
| Polar |
|---|
| 3z2-r2 | -1.234 |
|---|
| x2-y2 | -0.355 |
|---|
| xy | -2.083 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.098 |
1.609 |
0.000 |
| y |
1.609 |
6.715 |
0.000 |
| z |
0.000 |
0.000 |
2.395 |
<r2> (average value of r
2) Å
2
| <r2> |
84.467 |
| (<r2>)1/2 |
9.191 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.685843 |
| Energy at 298.15K | |
| HF Energy | -191.685843 |
| Nuclear repulsion energy | 103.735714 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3187 |
3168 |
1.46 |
62.87 |
0.64 |
0.78 |
| 2 |
A' |
3101 |
3083 |
11.57 |
145.06 |
0.23 |
0.38 |
| 3 |
A' |
3077 |
3059 |
5.03 |
59.50 |
0.14 |
0.24 |
| 4 |
A' |
2802 |
2786 |
170.79 |
192.03 |
0.32 |
0.48 |
| 5 |
A' |
1737 |
1727 |
89.39 |
10.44 |
0.62 |
0.76 |
| 6 |
A' |
1623 |
1613 |
65.27 |
43.37 |
0.23 |
0.37 |
| 7 |
A' |
1372 |
1364 |
20.56 |
10.51 |
0.65 |
0.79 |
| 8 |
A' |
1352 |
1344 |
18.52 |
5.80 |
0.27 |
0.43 |
| 9 |
A' |
1259 |
1251 |
3.10 |
16.08 |
0.44 |
0.61 |
| 10 |
A' |
1025 |
1019 |
4.80 |
5.31 |
0.70 |
0.82 |
| 11 |
A' |
919 |
913 |
53.68 |
5.53 |
0.20 |
0.34 |
| 12 |
A' |
658 |
654 |
7.71 |
0.62 |
0.74 |
0.85 |
| 13 |
A' |
278 |
276 |
6.62 |
3.20 |
0.44 |
0.61 |
| 14 |
A" |
999 |
994 |
10.62 |
6.34 |
0.75 |
0.86 |
| 15 |
A" |
990 |
985 |
13.19 |
2.22 |
0.75 |
0.86 |
| 16 |
A" |
958 |
952 |
12.30 |
0.20 |
0.75 |
0.86 |
| 17 |
A" |
541 |
538 |
6.65 |
6.52 |
0.75 |
0.86 |
| 18 |
A" |
192 |
191 |
4.81 |
1.25 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13033.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12958.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.894 |
-0.286 |
0.000 |
| C2 |
0.000 |
0.905 |
0.000 |
| C3 |
1.342 |
0.768 |
0.000 |
| O4 |
-0.500 |
-1.445 |
0.000 |
| H5 |
-1.999 |
-0.049 |
0.000 |
| H6 |
-0.479 |
1.898 |
0.000 |
| H7 |
2.018 |
1.635 |
0.000 |
| H8 |
1.780 |
-0.243 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4889 | 2.4721 | 1.2247 | 1.1299 | 2.2225 | 3.4889 | 2.6749 |
C2 | 1.4889 | | 1.3486 | 2.4025 | 2.2149 | 1.1027 | 2.1461 | 2.1177 | C3 | 2.4721 | 1.3486 | | 2.8797 | 3.4394 | 2.1428 | 1.0997 | 1.1017 | O4 | 1.2247 | 2.4025 | 2.8797 | | 2.0486 | 3.3430 | 3.9789 | 2.5781 | H5 | 1.1299 | 2.2149 | 3.4394 | 2.0486 | | 2.4696 | 4.3558 | 3.7843 | H6 | 2.2225 | 1.1027 | 2.1428 | 3.3430 | 2.4696 | | 2.5108 | 3.1121 | H7 | 3.4889 | 2.1461 | 1.0997 | 3.9789 | 4.3558 | 2.5108 | | 1.8930 | H8 | 2.6749 | 2.1177 | 1.1017 | 2.5781 | 3.7843 | 3.1121 | 1.8930 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.120 |
|
C1 |
C2 |
H6 |
117.312 |
| C2 |
C1 |
O4 |
124.301 |
|
C2 |
C1 |
H5 |
114.820 |
| C2 |
C3 |
H7 |
122.134 |
|
C2 |
C3 |
H8 |
119.252 |
| C3 |
C2 |
H6 |
121.568 |
|
O4 |
C1 |
H5 |
120.878 |
| H7 |
C3 |
H8 |
118.614 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.154 |
|
|
|
| 2 |
C |
-0.125 |
|
|
|
| 3 |
C |
0.058 |
|
|
|
| 4 |
O |
-0.172 |
|
|
|
| 5 |
H |
-0.010 |
|
|
|
| 6 |
H |
0.003 |
|
|
|
| 7 |
H |
0.045 |
|
|
|
| 8 |
H |
0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.329 |
2.373 |
0.000 |
2.395 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.649 |
-0.514 |
0.000 |
| y |
-0.514 |
-25.514 |
0.000 |
| z |
0.000 |
0.000 |
-24.522 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.369 |
-0.514 |
0.000 |
| y |
-0.514 |
-2.929 |
0.000 |
| z |
0.000 |
0.000 |
-1.440 |
|
| Polar |
|---|
| 3z2-r2 | -2.881 |
|---|
| x2-y2 | 4.865 |
|---|
| xy | -0.514 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.489 |
0.529 |
0.000 |
| y |
0.529 |
5.939 |
0.000 |
| z |
0.000 |
0.000 |
2.393 |
<r2> (average value of r
2) Å
2
| <r2> |
75.775 |
| (<r2>)1/2 |
8.705 |
