Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3110.557000 |
| Energy at 298.15K | |
| HF Energy | -3109.961117 |
| Nuclear repulsion energy | 287.216335 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
199 |
191 |
6.06 |
|
|
|
| 2 |
A' |
254 |
243 |
0.71 |
|
|
|
| 3 |
A' |
666 |
639 |
45.23 |
|
|
|
| 4 |
A' |
773 |
741 |
22.41 |
|
|
|
| 5 |
A' |
1091 |
1047 |
1.16 |
|
|
|
| 6 |
A' |
1233 |
1183 |
47.27 |
|
|
|
| 7 |
A' |
1334 |
1280 |
1.15 |
|
|
|
| 8 |
A' |
1486 |
1426 |
4.28 |
|
|
|
| 9 |
A' |
1494 |
1433 |
0.57 |
|
|
|
| 10 |
A' |
3126 |
2999 |
5.05 |
|
|
|
| 11 |
A' |
3134 |
3007 |
9.74 |
|
|
|
| 12 |
A" |
116 |
111 |
5.06 |
|
|
|
| 13 |
A" |
769 |
737 |
2.41 |
|
|
|
| 14 |
A" |
986 |
946 |
0.33 |
|
|
|
| 15 |
A" |
1134 |
1088 |
2.37 |
|
|
|
| 16 |
A" |
1295 |
1243 |
0.02 |
|
|
|
| 17 |
A" |
3188 |
3059 |
0.84 |
|
|
|
| 18 |
A" |
3211 |
3081 |
3.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12744.2 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12226.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.643 |
0.000 |
| C2 |
1.243 |
-0.236 |
0.000 |
| Br3 |
-1.600 |
-0.487 |
0.000 |
| Cl4 |
2.711 |
0.811 |
0.000 |
| H5 |
-0.048 |
1.274 |
0.900 |
| H6 |
-0.048 |
1.274 |
-0.900 |
| H7 |
1.284 |
-0.871 |
0.898 |
| H8 |
1.284 |
-0.871 |
-0.898 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5223 | 1.9589 | 2.7159 | 1.1001 | 1.1001 | 2.1788 | 2.1788 |
C2 | 1.5223 | | 2.8540 | 1.8033 | 2.1812 | 2.1812 | 1.1007 | 1.1007 | Br3 | 1.9589 | 2.8540 | | 4.5021 | 2.5138 | 2.5138 | 3.0452 | 3.0452 | Cl4 | 2.7159 | 1.8033 | 4.5021 | | 2.9387 | 2.9387 | 2.3815 | 2.3815 | H5 | 1.1001 | 2.1812 | 2.5138 | 2.9387 | | 1.7998 | 2.5250 | 3.0999 | H6 | 1.1001 | 2.1812 | 2.5138 | 2.9387 | 1.7998 | | 3.0999 | 2.5250 | H7 | 2.1788 | 1.1007 | 3.0452 | 2.3815 | 2.5250 | 3.0999 | | 1.7968 | H8 | 2.1788 | 1.1007 | 3.0452 | 2.3815 | 3.0999 | 2.5250 | 1.7968 | |
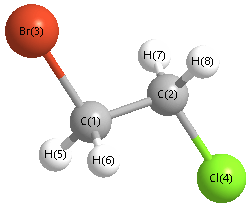 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.213 |
|
C1 |
C2 |
H7 |
111.320 |
| C1 |
C2 |
H8 |
111.320 |
|
C2 |
C1 |
Br3 |
109.495 |
| C2 |
C1 |
Cl4 |
38.829 |
|
C2 |
C1 |
H6 |
111.540 |
| Br3 |
C1 |
H5 |
107.154 |
|
Br3 |
C1 |
H6 |
107.154 |
| Cl4 |
C2 |
H7 |
107.720 |
|
Cl4 |
C2 |
H8 |
107.720 |
| H5 |
C1 |
H6 |
109.765 |
|
H7 |
C2 |
H8 |
109.412 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3110.554263 |
| Energy at 298.15K | |
| HF Energy | -3109.957758 |
| Nuclear repulsion energy | 303.356178 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3191 |
3061 |
3.59 |
|
|
|
| 2 |
A |
3173 |
3044 |
2.62 |
|
|
|
| 3 |
A |
3118 |
2991 |
18.04 |
|
|
|
| 4 |
A |
3102 |
2976 |
9.45 |
|
|
|
| 5 |
A |
1477 |
1417 |
1.41 |
|
|
|
| 6 |
A |
1466 |
1407 |
6.60 |
|
|
|
| 7 |
A |
1345 |
1290 |
23.71 |
|
|
|
| 8 |
A |
1305 |
1252 |
49.62 |
|
|
|
| 9 |
A |
1224 |
1174 |
2.43 |
|
|
|
| 10 |
A |
1150 |
1103 |
1.66 |
|
|
|
| 11 |
A |
1066 |
1022 |
1.44 |
|
|
|
| 12 |
A |
950 |
912 |
6.88 |
|
|
|
| 13 |
A |
882 |
846 |
15.61 |
|
|
|
| 14 |
A |
697 |
669 |
16.81 |
|
|
|
| 15 |
A |
591 |
567 |
10.04 |
|
|
|
| 16 |
A |
391 |
375 |
5.88 |
|
|
|
| 17 |
A |
246 |
236 |
0.88 |
|
|
|
| 18 |
A |
103 |
99 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12737.6 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12220.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.007 |
1.106 |
-0.386 |
| C2 |
1.291 |
0.903 |
0.405 |
| Br3 |
-1.359 |
-0.226 |
0.036 |
| Cl4 |
2.175 |
-0.580 |
-0.087 |
| H5 |
-0.429 |
2.087 |
-0.137 |
| H6 |
0.190 |
1.047 |
-1.470 |
| H7 |
1.966 |
1.757 |
0.220 |
| H8 |
1.088 |
0.829 |
1.485 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5223 | 1.9543 | 2.7630 | 1.1023 | 1.1006 | 2.1514 | 2.1786 |
C2 | 1.5223 | | 2.9048 | 1.7958 | 2.1577 | 2.1795 | 1.1035 | 1.1012 | Br3 | 1.9543 | 2.9048 | | 3.5543 | 2.4997 | 2.5077 | 3.8759 | 3.0333 | Cl4 | 2.7630 | 1.7958 | 3.5543 | | 3.7283 | 2.9157 | 2.3669 | 2.3749 | H5 | 1.1023 | 2.1577 | 2.4997 | 3.7283 | | 1.8004 | 2.4434 | 2.5525 | H6 | 1.1006 | 2.1795 | 2.5077 | 2.9157 | 1.8004 | | 2.5525 | 3.0959 | H7 | 2.1514 | 1.1035 | 3.8759 | 2.3669 | 2.4434 | 2.5525 | | 1.7974 | H8 | 2.1786 | 1.1012 | 3.0333 | 2.3749 | 2.5525 | 3.0959 | 1.7974 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.498 |
|
C1 |
C2 |
H7 |
108.994 |
| C1 |
C2 |
H8 |
111.270 |
|
C2 |
C1 |
Br3 |
112.755 |
| C2 |
C1 |
Cl4 |
36.904 |
|
C2 |
C1 |
H6 |
111.378 |
| Br3 |
C1 |
H5 |
106.368 |
|
Br3 |
C1 |
H6 |
107.000 |
| Cl4 |
C2 |
H7 |
107.014 |
|
Cl4 |
C2 |
H8 |
107.706 |
| H5 |
C1 |
H6 |
109.621 |
|
H7 |
C2 |
H8 |
109.228 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
