Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3110.553072 |
| Energy at 298.15K | |
| HF Energy | -3109.961138 |
| Nuclear repulsion energy | 287.609739 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
200 |
192 |
5.96 |
|
|
|
| 2 |
A' |
256 |
245 |
0.72 |
|
|
|
| 3 |
A' |
680 |
651 |
39.74 |
|
|
|
| 4 |
A' |
788 |
754 |
21.33 |
|
|
|
| 5 |
A' |
1094 |
1047 |
1.20 |
|
|
|
| 6 |
A' |
1242 |
1188 |
45.05 |
|
|
|
| 7 |
A' |
1343 |
1285 |
1.13 |
|
|
|
| 8 |
A' |
1490 |
1425 |
4.35 |
|
|
|
| 9 |
A' |
1497 |
1432 |
0.70 |
|
|
|
| 10 |
A' |
3133 |
2997 |
4.65 |
|
|
|
| 11 |
A' |
3141 |
3005 |
9.72 |
|
|
|
| 12 |
A" |
117 |
112 |
5.02 |
|
|
|
| 13 |
A" |
770 |
736 |
2.47 |
|
|
|
| 14 |
A" |
990 |
947 |
0.33 |
|
|
|
| 15 |
A" |
1141 |
1091 |
2.35 |
|
|
|
| 16 |
A" |
1300 |
1244 |
0.02 |
|
|
|
| 17 |
A" |
3195 |
3057 |
0.80 |
|
|
|
| 18 |
A" |
3218 |
3079 |
3.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12796.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12242.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.641 |
0.000 |
| C2 |
1.242 |
-0.239 |
0.000 |
| Br3 |
-1.600 |
-0.481 |
0.000 |
| Cl4 |
2.710 |
0.801 |
0.000 |
| H5 |
-0.045 |
1.273 |
0.899 |
| H6 |
-0.045 |
1.273 |
-0.899 |
| H7 |
1.281 |
-0.875 |
0.898 |
| H8 |
1.281 |
-0.875 |
-0.898 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5224 | 1.9537 | 2.7147 | 1.0999 | 1.0999 | 2.1778 | 2.1778 |
C2 | 1.5224 | | 2.8523 | 1.7991 | 2.1797 | 2.1797 | 1.1005 | 1.1005 | Br3 | 1.9537 | 2.8523 | | 4.4964 | 2.5104 | 2.5104 | 3.0428 | 3.0428 | Cl4 | 2.7147 | 1.7991 | 4.4964 | | 2.9357 | 2.9357 | 2.3785 | 2.3785 | H5 | 1.0999 | 2.1797 | 2.5104 | 2.9357 | | 1.7979 | 2.5238 | 3.0979 | H6 | 1.0999 | 2.1797 | 2.5104 | 2.9357 | 1.7979 | | 3.0979 | 2.5238 | H7 | 2.1778 | 1.1005 | 3.0428 | 2.3785 | 2.5238 | 3.0979 | | 1.7952 | H8 | 2.1778 | 1.1005 | 3.0428 | 2.3785 | 3.0979 | 2.5238 | 1.7952 | |
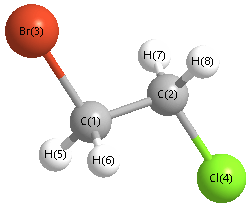 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.351 |
|
C1 |
C2 |
H7 |
111.246 |
| C1 |
C2 |
H8 |
111.246 |
|
C2 |
C1 |
Br3 |
109.658 |
| C2 |
C1 |
Cl4 |
38.704 |
|
C2 |
C1 |
H6 |
111.430 |
| Br3 |
C1 |
H5 |
107.261 |
|
Br3 |
C1 |
H6 |
107.261 |
| Cl4 |
C2 |
H7 |
107.786 |
|
Cl4 |
C2 |
H8 |
107.786 |
| H5 |
C1 |
H6 |
109.625 |
|
H7 |
C2 |
H8 |
109.303 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3110.550385 |
| Energy at 298.15K | |
| HF Energy | -3109.957785 |
| Nuclear repulsion energy | 303.808270 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3058 |
3.44 |
|
|
|
| 2 |
A |
3179 |
3042 |
2.59 |
|
|
|
| 3 |
A |
3124 |
2988 |
17.30 |
|
|
|
| 4 |
A |
3109 |
2974 |
9.07 |
|
|
|
| 5 |
A |
1480 |
1416 |
1.47 |
|
|
|
| 6 |
A |
1470 |
1406 |
6.79 |
|
|
|
| 7 |
A |
1353 |
1294 |
22.64 |
|
|
|
| 8 |
A |
1314 |
1257 |
46.73 |
|
|
|
| 9 |
A |
1230 |
1177 |
1.97 |
|
|
|
| 10 |
A |
1157 |
1107 |
1.64 |
|
|
|
| 11 |
A |
1071 |
1024 |
1.36 |
|
|
|
| 12 |
A |
955 |
914 |
6.74 |
|
|
|
| 13 |
A |
888 |
849 |
15.24 |
|
|
|
| 14 |
A |
708 |
678 |
14.74 |
|
|
|
| 15 |
A |
603 |
577 |
8.50 |
|
|
|
| 16 |
A |
394 |
377 |
5.49 |
|
|
|
| 17 |
A |
248 |
237 |
0.86 |
|
|
|
| 18 |
A |
103 |
99 |
0.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12790.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12236.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.004 |
1.103 |
-0.385 |
| C2 |
1.290 |
0.901 |
0.404 |
| Br3 |
-1.357 |
-0.226 |
0.036 |
| Cl4 |
2.173 |
-0.579 |
-0.087 |
| H5 |
-0.431 |
2.085 |
-0.137 |
| H6 |
0.187 |
1.045 |
-1.469 |
| H7 |
1.961 |
1.757 |
0.219 |
| H8 |
1.087 |
0.829 |
1.484 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5222 | 1.9487 | 2.7607 | 1.1022 | 1.1006 | 2.1502 | 2.1778 |
C2 | 1.5222 | | 2.9005 | 1.7922 | 2.1571 | 2.1787 | 1.1033 | 1.1011 | Br3 | 1.9487 | 2.9005 | | 3.5502 | 2.4957 | 2.5036 | 3.8701 | 3.0303 | Cl4 | 2.7607 | 1.7922 | 3.5502 | | 3.7250 | 2.9141 | 2.3651 | 2.3727 | H5 | 1.1022 | 2.1571 | 2.4957 | 3.7250 | | 1.7991 | 2.4403 | 2.5511 | H6 | 1.1006 | 2.1787 | 2.5036 | 2.9141 | 1.7991 | | 2.5505 | 3.0950 | H7 | 2.1502 | 1.1033 | 3.8701 | 2.3651 | 2.4403 | 2.5505 | | 1.7960 | H8 | 2.1778 | 1.1011 | 3.0303 | 2.3727 | 2.5511 | 3.0950 | 1.7960 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.547 |
|
C1 |
C2 |
H7 |
108.918 |
| C1 |
C2 |
H8 |
111.218 |
|
C2 |
C1 |
Br3 |
112.794 |
| C2 |
C1 |
Cl4 |
36.839 |
|
C2 |
C1 |
H6 |
111.322 |
| Br3 |
C1 |
H5 |
106.447 |
|
Br3 |
C1 |
H6 |
107.068 |
| Cl4 |
C2 |
H7 |
107.124 |
|
Cl4 |
C2 |
H8 |
107.781 |
| H5 |
C1 |
H6 |
109.522 |
|
H7 |
C2 |
H8 |
109.125 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
