Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3112.271605 |
| Energy at 298.15K | |
| HF Energy | -3112.271605 |
| Nuclear repulsion energy | 286.103971 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3039 |
3021 |
6.63 |
46.41 |
0.05 |
0.09 |
| 2 |
A' |
3026 |
3008 |
3.20 |
147.55 |
0.05 |
0.10 |
| 3 |
A' |
1415 |
1407 |
0.41 |
14.43 |
0.72 |
0.84 |
| 4 |
A' |
1408 |
1400 |
4.77 |
1.26 |
0.70 |
0.83 |
| 5 |
A' |
1240 |
1233 |
1.15 |
17.01 |
0.66 |
0.79 |
| 6 |
A' |
1155 |
1149 |
36.33 |
2.02 |
0.51 |
0.68 |
| 7 |
A' |
1054 |
1048 |
0.95 |
8.81 |
0.74 |
0.85 |
| 8 |
A' |
710 |
706 |
30.74 |
51.78 |
0.29 |
0.45 |
| 9 |
A' |
619 |
615 |
58.77 |
14.87 |
0.26 |
0.41 |
| 10 |
A' |
240 |
239 |
0.85 |
4.71 |
0.37 |
0.54 |
| 11 |
A' |
186 |
185 |
7.51 |
0.21 |
0.40 |
0.57 |
| 12 |
A" |
3118 |
3100 |
1.39 |
9.98 |
0.75 |
0.86 |
| 13 |
A" |
3090 |
3072 |
0.63 |
102.05 |
0.75 |
0.86 |
| 14 |
A" |
1230 |
1223 |
0.02 |
5.42 |
0.75 |
0.86 |
| 15 |
A" |
1062 |
1056 |
2.35 |
0.16 |
0.75 |
0.86 |
| 16 |
A" |
924 |
919 |
0.19 |
8.32 |
0.75 |
0.86 |
| 17 |
A" |
745 |
740 |
4.56 |
0.04 |
0.75 |
0.86 |
| 18 |
A" |
109 |
108 |
4.78 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12184.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12114.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.649 |
0.000 |
| C2 |
1.241 |
-0.222 |
0.000 |
| Br3 |
-1.600 |
-0.508 |
0.000 |
| Cl4 |
2.711 |
0.845 |
0.000 |
| H5 |
-0.068 |
1.279 |
0.905 |
| H6 |
-0.068 |
1.279 |
-0.905 |
| H7 |
1.298 |
-0.858 |
0.903 |
| H8 |
1.298 |
-0.858 |
-0.903 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5158 | 1.9739 | 2.7184 | 1.1049 | 1.1049 | 2.1841 | 2.1841 |
C2 | 1.5158 | | 2.8549 | 1.8172 | 2.1871 | 2.1871 | 1.1057 | 1.1057 | Br3 | 1.9739 | 2.8549 | | 4.5185 | 2.5215 | 2.5215 | 3.0560 | 3.0560 | Cl4 | 2.7184 | 1.8172 | 4.5185 | | 2.9545 | 2.9545 | 2.3898 | 2.3898 | H5 | 1.1049 | 2.1871 | 2.5215 | 2.9545 | | 1.8101 | 2.5357 | 3.1144 | H6 | 1.1049 | 2.1871 | 2.5215 | 2.9545 | 1.8101 | | 3.1144 | 2.5357 | H7 | 2.1841 | 1.1057 | 3.0560 | 2.3898 | 2.5357 | 3.1144 | | 1.8064 | H8 | 2.1841 | 1.1057 | 3.0560 | 2.3898 | 3.1144 | 2.5357 | 1.8064 | |
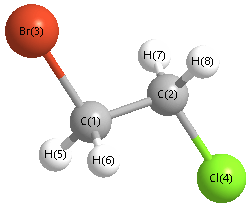 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.961 |
|
C1 |
C2 |
H7 |
111.891 |
| C1 |
C2 |
H8 |
111.891 |
|
C2 |
C1 |
Br3 |
109.083 |
| C2 |
C1 |
Cl4 |
39.213 |
|
C2 |
C1 |
H6 |
112.192 |
| Br3 |
C1 |
H5 |
106.527 |
|
Br3 |
C1 |
H6 |
106.527 |
| Cl4 |
C2 |
H7 |
107.170 |
|
Cl4 |
C2 |
H8 |
107.170 |
| H5 |
C1 |
H6 |
109.997 |
|
H7 |
C2 |
H8 |
109.533 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.104 |
|
|
|
| 2 |
C |
-0.072 |
|
|
|
| 3 |
Br |
-0.090 |
|
|
|
| 4 |
Cl |
-0.115 |
|
|
|
| 5 |
H |
0.096 |
|
|
|
| 6 |
H |
0.096 |
|
|
|
| 7 |
H |
0.095 |
|
|
|
| 8 |
H |
0.095 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.108 |
-0.047 |
0.000 |
0.118 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.133 |
-1.989 |
0.000 |
| y |
-1.989 |
-43.002 |
0.000 |
| z |
0.000 |
0.000 |
-43.235 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.015 |
-1.989 |
0.000 |
| y |
-1.989 |
3.182 |
0.000 |
| z |
0.000 |
0.000 |
2.833 |
|
| Polar |
|---|
| 3z2-r2 | 5.666 |
|---|
| x2-y2 | -6.131 |
|---|
| xy | -1.989 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.577 |
2.117 |
0.000 |
| y |
2.117 |
6.436 |
0.000 |
| z |
0.000 |
0.000 |
4.958 |
<r2> (average value of r
2) Å
2
| <r2> |
287.357 |
| (<r2>)1/2 |
16.952 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3112.268983 |
| Energy at 298.15K | |
| HF Energy | -3112.268983 |
| Nuclear repulsion energy | 301.862489 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3094 |
3076 |
1.45 |
50.28 |
0.75 |
0.86 |
| 2 |
A |
3070 |
3052 |
1.41 |
99.35 |
0.75 |
0.86 |
| 3 |
A |
3018 |
3001 |
10.72 |
188.33 |
0.03 |
0.07 |
| 4 |
A |
2997 |
2980 |
8.96 |
134.12 |
0.19 |
0.32 |
| 5 |
A |
1388 |
1380 |
2.35 |
5.73 |
0.71 |
0.83 |
| 6 |
A |
1380 |
1372 |
9.77 |
14.46 |
0.75 |
0.86 |
| 7 |
A |
1265 |
1258 |
20.61 |
4.45 |
0.75 |
0.86 |
| 8 |
A |
1222 |
1215 |
49.71 |
1.88 |
0.59 |
0.74 |
| 9 |
A |
1155 |
1149 |
3.75 |
15.72 |
0.75 |
0.85 |
| 10 |
A |
1089 |
1082 |
1.20 |
6.12 |
0.75 |
0.85 |
| 11 |
A |
1016 |
1011 |
2.74 |
3.46 |
0.63 |
0.77 |
| 12 |
A |
895 |
889 |
9.11 |
7.39 |
0.49 |
0.66 |
| 13 |
A |
830 |
825 |
22.81 |
1.35 |
0.75 |
0.86 |
| 14 |
A |
656 |
653 |
20.78 |
8.14 |
0.39 |
0.56 |
| 15 |
A |
558 |
555 |
11.86 |
13.05 |
0.19 |
0.32 |
| 16 |
A |
375 |
373 |
7.92 |
3.00 |
0.73 |
0.84 |
| 17 |
A |
237 |
236 |
1.22 |
1.69 |
0.46 |
0.63 |
| 18 |
A |
95 |
95 |
0.42 |
1.79 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 12170.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12099.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.012 |
1.111 |
-0.381 |
| C2 |
1.294 |
0.908 |
0.399 |
| Br3 |
-1.368 |
-0.227 |
0.036 |
| Cl4 |
2.189 |
-0.583 |
-0.086 |
| H5 |
-0.431 |
2.093 |
-0.121 |
| H6 |
0.178 |
1.056 |
-1.472 |
| H7 |
1.974 |
1.763 |
0.204 |
| H8 |
1.105 |
0.836 |
1.487 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5142 | 1.9671 | 2.7747 | 1.1075 | 1.1054 | 2.1489 | 2.1815 |
C2 | 1.5142 | | 2.9166 | 1.8060 | 2.1562 | 2.1834 | 1.1095 | 1.1064 | Br3 | 1.9671 | 2.9166 | | 3.5770 | 2.5066 | 2.5120 | 3.8935 | 3.0582 | Cl4 | 2.7747 | 1.8060 | 3.5770 | | 3.7454 | 2.9412 | 2.3733 | 2.3797 | H5 | 1.1075 | 2.1562 | 2.5066 | 3.7454 | | 1.8091 | 2.4501 | 2.5543 | H6 | 1.1054 | 2.1834 | 2.5120 | 2.9412 | 1.8091 | | 2.5566 | 3.1085 | H7 | 2.1489 | 1.1095 | 3.8935 | 2.3733 | 2.4501 | 2.5566 | | 1.8053 | H8 | 2.1815 | 1.1064 | 3.0582 | 2.3797 | 2.5543 | 3.1085 | 1.8053 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
113.093 |
|
C1 |
C2 |
H7 |
109.008 |
| C1 |
C2 |
H8 |
111.760 |
|
C2 |
C1 |
Br3 |
113.178 |
| C2 |
C1 |
Cl4 |
36.777 |
|
C2 |
C1 |
H6 |
111.976 |
| Br3 |
C1 |
H5 |
105.807 |
|
Br3 |
C1 |
H6 |
106.275 |
| Cl4 |
C2 |
H7 |
106.529 |
|
Cl4 |
C2 |
H8 |
107.137 |
| H5 |
C1 |
H6 |
109.675 |
|
H7 |
C2 |
H8 |
109.122 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.107 |
|
|
|
| 2 |
C |
-0.073 |
|
|
|
| 3 |
Br |
-0.082 |
|
|
|
| 4 |
Cl |
-0.106 |
|
|
|
| 5 |
H |
0.089 |
|
|
|
| 6 |
H |
0.096 |
|
|
|
| 7 |
H |
0.089 |
|
|
|
| 8 |
H |
0.094 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.316 |
2.531 |
0.106 |
2.553 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.833 |
2.629 |
0.353 |
| y |
2.629 |
-40.257 |
-0.089 |
| z |
0.353 |
-0.089 |
-42.875 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.267 |
2.629 |
0.353 |
| y |
2.629 |
4.596 |
-0.089 |
| z |
0.353 |
-0.089 |
0.671 |
|
| Polar |
|---|
| 3z2-r2 | 1.341 |
|---|
| x2-y2 | -6.575 |
|---|
| xy | 2.629 |
|---|
| xz | 0.353 |
|---|
| yz | -0.089 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.375 |
0.314 |
-0.231 |
| y |
0.314 |
7.308 |
0.048 |
| z |
-0.231 |
0.048 |
5.279 |
<r2> (average value of r
2) Å
2
| <r2> |
225.092 |
| (<r2>)1/2 |
15.003 |
