Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -997.681489 |
| Energy at 298.15K | -997.686354 |
| HF Energy | -997.076041 |
| Nuclear repulsion energy | 201.079655 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
3043 |
1.80 |
|
|
|
| 2 |
A |
3112 |
2986 |
26.91 |
|
|
|
| 3 |
A |
1478 |
1418 |
0.22 |
|
|
|
| 4 |
A |
1356 |
1301 |
21.30 |
|
|
|
| 5 |
A |
1240 |
1190 |
0.97 |
|
|
|
| 6 |
A |
1074 |
1030 |
1.21 |
|
|
|
| 7 |
A |
973 |
934 |
8.85 |
|
|
|
| 8 |
A |
686 |
658 |
17.12 |
|
|
|
| 9 |
A |
265 |
255 |
0.65 |
|
|
|
| 10 |
A |
120 |
115 |
0.84 |
|
|
|
| 11 |
B |
3182 |
3053 |
7.90 |
|
|
|
| 12 |
B |
3102 |
2976 |
4.75 |
|
|
|
| 13 |
B |
1472 |
1413 |
7.54 |
|
|
|
| 14 |
B |
1328 |
1274 |
39.17 |
|
|
|
| 15 |
B |
1168 |
1121 |
0.91 |
|
|
|
| 16 |
B |
909 |
872 |
14.19 |
|
|
|
| 17 |
B |
712 |
683 |
21.34 |
|
|
|
| 18 |
B |
416 |
399 |
6.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12881.4 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12358.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.297 |
0.701 |
0.898 |
| C2 |
-0.297 |
-0.701 |
0.898 |
| Cl3 |
-0.297 |
1.696 |
-0.473 |
| Cl4 |
0.297 |
-1.696 |
-0.473 |
| H5 |
0.004 |
1.218 |
1.827 |
| H6 |
1.396 |
0.664 |
0.833 |
| H7 |
-0.004 |
-1.218 |
1.827 |
| H8 |
-1.396 |
-0.664 |
0.833 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5224 | 1.7961 | 2.7614 | 1.1031 | 1.1013 | 2.1530 | 2.1759 |
C2 | 1.5224 | | 2.7614 | 1.7961 | 2.1530 | 2.1759 | 1.1031 | 1.1013 | Cl3 | 1.7961 | 2.7614 | | 3.4439 | 2.3691 | 2.3750 | 3.7243 | 2.9124 | Cl4 | 2.7614 | 1.7961 | 3.4439 | | 3.7243 | 2.9124 | 2.3691 | 2.3750 | H5 | 1.1031 | 2.1530 | 2.3691 | 3.7243 | | 1.7982 | 2.4360 | 2.5477 | H6 | 1.1013 | 2.1759 | 2.3750 | 2.9124 | 1.7982 | | 2.5477 | 3.0919 | H7 | 2.1530 | 1.1031 | 3.7243 | 2.3691 | 2.4360 | 2.5477 | | 1.7982 | H8 | 2.1759 | 1.1013 | 2.9124 | 2.3750 | 2.5477 | 3.0919 | 1.7982 | |
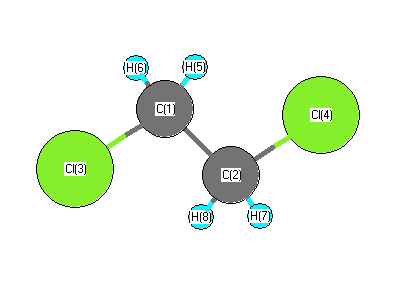 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.375 |
|
C1 |
C2 |
H7 |
109.135 |
| C1 |
C2 |
H8 |
111.040 |
|
C2 |
C1 |
Cl3 |
112.375 |
| C2 |
C1 |
H5 |
109.135 |
|
C2 |
C1 |
H6 |
111.040 |
| Cl3 |
C1 |
H5 |
107.169 |
|
Cl3 |
C1 |
H6 |
107.685 |
| Cl4 |
C2 |
H7 |
107.169 |
|
Cl4 |
C2 |
H8 |
107.685 |
| H5 |
C1 |
H6 |
109.324 |
|
H7 |
C2 |
H8 |
109.324 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
