Jump to
S1C2
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -997.677932 |
| Energy at 298.15K | -997.682816 |
| HF Energy | -997.076082 |
| Nuclear repulsion energy | 201.321418 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3180 |
3042 |
1.78 |
|
|
|
| 2 |
A |
3120 |
2985 |
25.79 |
|
|
|
| 3 |
A |
1481 |
1417 |
0.23 |
|
|
|
| 4 |
A |
1363 |
1304 |
20.37 |
|
|
|
| 5 |
A |
1245 |
1191 |
0.84 |
|
|
|
| 6 |
A |
1078 |
1032 |
1.19 |
|
|
|
| 7 |
A |
978 |
936 |
8.62 |
|
|
|
| 8 |
A |
696 |
666 |
15.56 |
|
|
|
| 9 |
A |
266 |
255 |
0.64 |
|
|
|
| 10 |
A |
120 |
115 |
0.85 |
|
|
|
| 11 |
B |
3190 |
3052 |
7.56 |
|
|
|
| 12 |
B |
3110 |
2975 |
4.63 |
|
|
|
| 13 |
B |
1476 |
1412 |
7.79 |
|
|
|
| 14 |
B |
1336 |
1278 |
37.49 |
|
|
|
| 15 |
B |
1174 |
1123 |
0.90 |
|
|
|
| 16 |
B |
914 |
874 |
14.15 |
|
|
|
| 17 |
B |
723 |
692 |
18.85 |
|
|
|
| 18 |
B |
418 |
400 |
6.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12933.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12373.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.297 |
0.701 |
0.896 |
| C2 |
-0.297 |
-0.701 |
0.896 |
| Cl3 |
-0.297 |
1.694 |
-0.473 |
| Cl4 |
0.297 |
-1.694 |
-0.473 |
| H5 |
0.004 |
1.217 |
1.826 |
| H6 |
1.395 |
0.664 |
0.833 |
| H7 |
-0.004 |
-1.217 |
1.826 |
| H8 |
-1.395 |
-0.664 |
0.833 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5223 | 1.7925 | 2.7589 | 1.1027 | 1.1010 | 2.1526 | 2.1745 |
C2 | 1.5223 | | 2.7589 | 1.7925 | 2.1526 | 2.1745 | 1.1027 | 1.1010 | Cl3 | 1.7925 | 2.7589 | | 3.4397 | 2.3667 | 2.3725 | 3.7210 | 2.9108 | Cl4 | 2.7589 | 1.7925 | 3.4397 | | 3.7210 | 2.9108 | 2.3667 | 2.3725 | H5 | 1.1027 | 2.1526 | 2.3667 | 3.7210 | | 1.7965 | 2.4346 | 2.5457 | H6 | 1.1010 | 2.1745 | 2.3725 | 2.9108 | 1.7965 | | 2.5457 | 3.0900 | H7 | 2.1526 | 1.1027 | 3.7210 | 2.3667 | 2.4346 | 2.5457 | | 1.7965 | H8 | 2.1745 | 1.1010 | 2.9108 | 2.3725 | 2.5457 | 3.0900 | 1.7965 | |
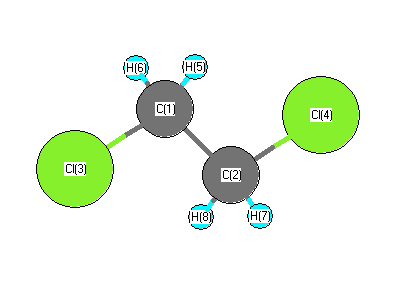 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.417 |
|
C1 |
C2 |
H7 |
109.130 |
| C1 |
C2 |
H8 |
110.958 |
|
C2 |
C1 |
Cl3 |
112.417 |
| C2 |
C1 |
H5 |
109.130 |
|
C2 |
C1 |
H6 |
110.958 |
| Cl3 |
C1 |
H5 |
107.248 |
|
Cl3 |
C1 |
H6 |
107.755 |
| Cl4 |
C2 |
H7 |
107.248 |
|
Cl4 |
C2 |
H8 |
107.755 |
| H5 |
C1 |
H6 |
109.221 |
|
H7 |
C2 |
H8 |
109.221 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
