Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -998.544802 |
| Energy at 298.15K | -998.549378 |
| Nuclear repulsion energy | 192.624510 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3025 |
3007 |
0.00 |
|
|
|
| 2 |
Ag |
1418 |
1409 |
0.00 |
|
|
|
| 3 |
Ag |
1260 |
1253 |
0.00 |
|
|
|
| 4 |
Ag |
1054 |
1048 |
0.00 |
|
|
|
| 5 |
Ag |
734 |
730 |
0.00 |
|
|
|
| 6 |
Ag |
291 |
290 |
0.00 |
|
|
|
| 7 |
Au |
3109 |
3091 |
3.93 |
|
|
|
| 8 |
Au |
1078 |
1072 |
1.81 |
|
|
|
| 9 |
Au |
758 |
753 |
3.60 |
|
|
|
| 10 |
Au |
117 |
116 |
6.05 |
|
|
|
| 11 |
Bg |
3086 |
3068 |
0.00 |
|
|
|
| 12 |
Bg |
1235 |
1228 |
0.00 |
|
|
|
| 13 |
Bg |
958 |
952 |
0.00 |
|
|
|
| 14 |
Bu |
3031 |
3014 |
14.72 |
|
|
|
| 15 |
Bu |
1414 |
1406 |
4.80 |
|
|
|
| 16 |
Bu |
1182 |
1175 |
29.11 |
|
|
|
| 17 |
Bu |
700 |
696 |
99.90 |
|
|
|
| 18 |
Bu |
207 |
206 |
9.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12328.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12257.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.478 |
0.589 |
0.000 |
| C2 |
-0.478 |
-0.589 |
0.000 |
| Cl3 |
-0.478 |
2.132 |
0.000 |
| Cl4 |
0.478 |
-2.132 |
0.000 |
| H5 |
1.116 |
0.598 |
0.904 |
| H6 |
1.116 |
0.598 |
-0.904 |
| H7 |
-1.116 |
-0.598 |
0.904 |
| H8 |
-1.116 |
-0.598 |
-0.904 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5183 | 1.8149 | 2.7210 | 1.1059 | 1.1059 | 2.1836 | 2.1836 |
C2 | 1.5183 | | 2.7210 | 1.8149 | 2.1836 | 2.1836 | 1.1059 | 1.1059 | Cl3 | 1.8149 | 2.7210 | | 4.3693 | 2.3896 | 2.3896 | 2.9451 | 2.9451 | Cl4 | 2.7210 | 1.8149 | 4.3693 | | 2.9451 | 2.9451 | 2.3896 | 2.3896 | H5 | 1.1059 | 2.1836 | 2.3896 | 2.9451 | | 1.8072 | 2.5320 | 3.1108 | H6 | 1.1059 | 2.1836 | 2.3896 | 2.9451 | 1.8072 | | 3.1108 | 2.5320 | H7 | 2.1836 | 1.1059 | 2.9451 | 2.3896 | 2.5320 | 3.1108 | | 1.8072 | H8 | 2.1836 | 1.1059 | 2.9451 | 2.3896 | 3.1108 | 2.5320 | 1.8072 | |
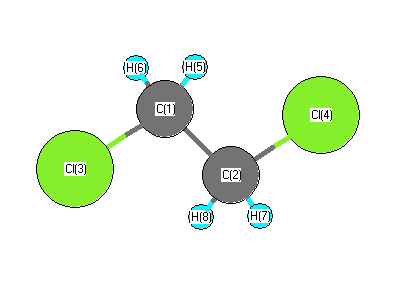 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.119 |
|
C1 |
C2 |
H7 |
111.674 |
| C1 |
C2 |
H8 |
111.674 |
|
C2 |
C1 |
Cl3 |
109.119 |
| C2 |
C1 |
H5 |
111.674 |
|
C2 |
C1 |
H6 |
111.674 |
| Cl3 |
C1 |
H5 |
107.294 |
|
Cl3 |
C1 |
H6 |
107.294 |
| Cl4 |
C2 |
H7 |
107.294 |
|
Cl4 |
C2 |
H8 |
107.294 |
| H5 |
C1 |
H6 |
109.591 |
|
H7 |
C2 |
H8 |
109.591 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.073 |
|
|
|
| 2 |
C |
-0.073 |
|
|
|
| 3 |
Cl |
-0.116 |
|
|
|
| 4 |
Cl |
-0.116 |
|
|
|
| 5 |
H |
0.094 |
|
|
|
| 6 |
H |
0.094 |
|
|
|
| 7 |
H |
0.094 |
|
|
|
| 8 |
H |
0.094 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.022 |
1.788 |
0.000 |
| y |
1.788 |
-44.132 |
0.000 |
| z |
0.000 |
0.000 |
-37.359 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.723 |
1.788 |
0.000 |
| y |
1.788 |
-6.941 |
0.000 |
| z |
0.000 |
0.000 |
3.218 |
|
| Polar |
|---|
| 3z2-r2 | 6.436 |
|---|
| x2-y2 | 7.110 |
|---|
| xy | 1.788 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.554 |
-1.460 |
0.000 |
| y |
-1.460 |
9.243 |
0.000 |
| z |
0.000 |
0.000 |
4.481 |
<r2> (average value of r
2) Å
2
| <r2> |
203.537 |
| (<r2>)1/2 |
14.267 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -998.542684 |
| Energy at 298.15K | -998.547401 |
| HF Energy | -998.542684 |
| Nuclear repulsion energy | 200.104689 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3068 |
3050 |
0.58 |
|
|
|
| 2 |
A |
3007 |
2990 |
21.24 |
|
|
|
| 3 |
A |
1389 |
1381 |
1.36 |
|
|
|
| 4 |
A |
1275 |
1268 |
19.41 |
|
|
|
| 5 |
A |
1173 |
1166 |
1.46 |
|
|
|
| 6 |
A |
1022 |
1016 |
2.50 |
|
|
|
| 7 |
A |
916 |
911 |
11.72 |
|
|
|
| 8 |
A |
649 |
645 |
18.29 |
|
|
|
| 9 |
A |
254 |
253 |
0.93 |
|
|
|
| 10 |
A |
111 |
111 |
0.72 |
|
|
|
| 11 |
B |
3081 |
3064 |
4.64 |
|
|
|
| 12 |
B |
2999 |
2981 |
3.38 |
|
|
|
| 13 |
B |
1384 |
1376 |
10.54 |
|
|
|
| 14 |
B |
1243 |
1236 |
38.77 |
|
|
|
| 15 |
B |
1106 |
1099 |
0.73 |
|
|
|
| 16 |
B |
855 |
850 |
20.93 |
|
|
|
| 17 |
B |
668 |
664 |
26.82 |
|
|
|
| 18 |
B |
399 |
396 |
8.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12299.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12228.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.294 |
0.698 |
0.901 |
| C2 |
-0.294 |
-0.698 |
0.901 |
| Cl3 |
-0.294 |
1.710 |
-0.475 |
| Cl4 |
0.294 |
-1.710 |
-0.475 |
| H5 |
-0.006 |
1.220 |
1.832 |
| H6 |
1.399 |
0.673 |
0.837 |
| H7 |
0.006 |
-1.220 |
1.832 |
| H8 |
-1.399 |
-0.673 |
0.837 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5159 | 1.8062 | 2.7738 | 1.1088 | 1.1065 | 2.1520 | 2.1795 |
C2 | 1.5159 | | 2.7738 | 1.8062 | 2.1520 | 2.1795 | 1.1088 | 1.1065 | Cl3 | 1.8062 | 2.7738 | | 3.4707 | 2.3761 | 2.3799 | 3.7414 | 2.9357 | Cl4 | 2.7738 | 1.8062 | 3.4707 | | 3.7414 | 2.9357 | 2.3761 | 2.3799 | H5 | 1.1088 | 2.1520 | 2.3761 | 3.7414 | | 1.8062 | 2.4398 | 2.5518 | H6 | 1.1065 | 2.1795 | 2.3799 | 2.9357 | 1.8062 | | 2.5518 | 3.1039 | H7 | 2.1520 | 1.1088 | 3.7414 | 2.3761 | 2.4398 | 2.5518 | | 1.8062 | H8 | 2.1795 | 1.1065 | 2.9357 | 2.3799 | 2.5518 | 3.1039 | 1.8062 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.933 |
|
C1 |
C2 |
H7 |
109.163 |
| C1 |
C2 |
H8 |
111.469 |
|
C2 |
C1 |
Cl3 |
112.933 |
| C2 |
C1 |
H5 |
109.163 |
|
C2 |
C1 |
H6 |
111.469 |
| Cl3 |
C1 |
H5 |
106.744 |
|
Cl3 |
C1 |
H6 |
107.127 |
| Cl4 |
C2 |
H7 |
106.744 |
|
Cl4 |
C2 |
H8 |
107.127 |
| H5 |
C1 |
H6 |
109.241 |
|
H7 |
C2 |
H8 |
109.241 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.073 |
|
|
|
| 2 |
C |
-0.073 |
|
|
|
| 3 |
Cl |
-0.107 |
|
|
|
| 4 |
Cl |
-0.107 |
|
|
|
| 5 |
H |
0.087 |
|
|
|
| 6 |
H |
0.094 |
|
|
|
| 7 |
H |
0.087 |
|
|
|
| 8 |
H |
0.094 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.636 |
2.636 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.226 |
1.047 |
0.000 |
| y |
1.047 |
-41.660 |
0.000 |
| z |
0.000 |
0.000 |
-35.118 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.163 |
1.047 |
0.000 |
| y |
1.047 |
-5.488 |
0.000 |
| z |
0.000 |
0.000 |
4.325 |
|
| Polar |
|---|
| 3z2-r2 | 8.650 |
|---|
| x2-y2 | 4.434 |
|---|
| xy | 1.047 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.902 |
-0.527 |
0.000 |
| y |
-0.527 |
7.112 |
0.000 |
| z |
0.000 |
0.000 |
6.627 |
<r2> (average value of r
2) Å
2
| <r2> |
166.330 |
| (<r2>)1/2 |
12.897 |
