return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-173.988048
Energy at 298.15K-173.998518
HF Energy-173.297774
Nuclear repulsion energy130.207830
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3445 3283        
2 A' 3125 2978        
3 A' 3037 2894        
4 A' 3029 2887        
5 A' 3018 2876        
6 A' 1657 1580        
7 A' 1497 1427        
8 A' 1485 1415        
9 A' 1473 1404        
10 A' 1409 1343        
11 A' 1399 1334        
12 A' 1316 1255        
13 A' 1159 1104        
14 A' 1109 1057        
15 A' 1057 1007        
16 A' 964 919        
17 A' 872 831        
18 A' 447 426        
19 A' 263 251        
20 A" 3527 3361        
21 A" 3117 2971        
22 A" 3082 2938        
23 A" 3055 2912        
24 A" 1488 1418        
25 A" 1395 1329        
26 A" 1315 1254        
27 A" 1240 1182        
28 A" 1035 986        
29 A" 857 817        
30 A" 743 708        
31 A" 316 301        
32 A" 235 224        
33 A" 134 127        

Unscaled Zero Point Vibrational Energy (zpe) 26648.9 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 25399.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.81199 0.12184 0.11476

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.435 1.296 0.000
C2 0.000 0.748 0.000
C3 -0.053 -0.790 0.000
N4 -1.399 -1.387 0.000
H5 1.447 2.402 0.000
H6 1.990 0.951 0.894
H7 1.990 0.951 -0.894
H8 -0.548 1.126 0.889
H9 -0.548 1.126 -0.889
H10 0.491 -1.175 -0.887
H11 0.491 -1.175 0.887
H12 -1.907 -1.007 -0.809
H13 -1.907 -1.007 0.809

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.53642.56273.90331.10601.10771.10772.17982.17982.79042.79044.13844.1384
C21.53641.53882.55272.19792.19132.19131.11091.11092.17382.17382.71432.7143
C32.56271.53881.47303.52712.82922.82922.16962.16961.10961.10962.03452.0345
N43.90332.55271.47304.73934.21374.21372.79852.79852.09912.09911.02791.0279
H51.10602.19793.52714.73931.78931.78932.52952.52953.80763.80764.84984.8498
H61.10772.19132.82924.21371.78931.78822.54393.10693.15282.60134.68174.3620
H71.10772.19132.82924.21371.78931.78823.10692.54392.60133.15284.36204.6817
H82.17981.11092.16962.79852.52952.54393.10691.77893.08702.52443.04592.5299
H92.17981.11092.16962.79852.52953.10692.54391.77892.52443.08702.52993.0459
H102.79042.17381.10962.09913.80763.15282.60133.08702.52441.77462.40532.9420
H112.79042.17381.10962.09913.80762.60133.15282.52443.08701.77462.94202.4053
H124.13842.71432.03451.02794.84984.68174.36203.04592.52992.40532.94201.6172
H134.13842.71432.03451.02794.84984.36204.68172.52993.04592.94202.40531.6172

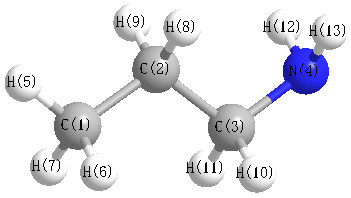
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.888 C1 C2 H8 109.815
C1 C2 H9 109.815 C2 C1 H5 111.526
C2 C1 H6 110.898 C2 C1 H7 110.898
C2 C3 N4 115.871 C2 C3 H10 109.256
C2 C3 H11 109.256 C3 C2 H8 108.857
C3 C2 H9 108.857 C3 N4 H12 107.540
C3 N4 H13 107.540 N4 C3 H10 107.910
N4 C3 H11 107.910 H5 C1 H6 107.859
H5 C1 H7 107.859 H6 C1 H7 107.642
H8 C2 H9 106.392 H10 C3 H11 106.204
H12 N4 H13 103.748
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability