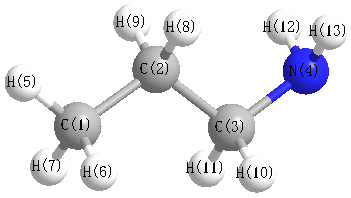
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3470 |
3348 |
1.55 |
|
|
|
| 2 |
A' |
3123 |
3014 |
29.92 |
|
|
|
| 3 |
A' |
3036 |
2930 |
36.11 |
|
|
|
| 4 |
A' |
3024 |
2918 |
62.01 |
|
|
|
| 5 |
A' |
3014 |
2909 |
13.32 |
|
|
|
| 6 |
A' |
1649 |
1592 |
16.74 |
|
|
|
| 7 |
A' |
1480 |
1428 |
4.73 |
|
|
|
| 8 |
A' |
1466 |
1415 |
1.14 |
|
|
|
| 9 |
A' |
1457 |
1406 |
0.13 |
|
|
|
| 10 |
A' |
1393 |
1344 |
0.96 |
|
|
|
| 11 |
A' |
1387 |
1339 |
7.70 |
|
|
|
| 12 |
A' |
1309 |
1263 |
11.70 |
|
|
|
| 13 |
A' |
1147 |
1107 |
2.71 |
|
|
|
| 14 |
A' |
1109 |
1070 |
18.17 |
|
|
|
| 15 |
A' |
1055 |
1018 |
0.09 |
|
|
|
| 16 |
A' |
927 |
894 |
121.73 |
|
|
|
| 17 |
A' |
859 |
828 |
60.09 |
|
|
|
| 18 |
A' |
447 |
431 |
2.72 |
|
|
|
| 19 |
A' |
262 |
253 |
3.27 |
|
|
|
| 20 |
A" |
3548 |
3424 |
0.09 |
|
|
|
| 21 |
A" |
3115 |
3006 |
46.66 |
|
|
|
| 22 |
A" |
3074 |
2967 |
48.14 |
|
|
|
| 23 |
A" |
3047 |
2940 |
1.29 |
|
|
|
| 24 |
A" |
1468 |
1416 |
7.23 |
|
|
|
| 25 |
A" |
1383 |
1334 |
0.45 |
|
|
|
| 26 |
A" |
1311 |
1265 |
0.69 |
|
|
|
| 27 |
A" |
1229 |
1186 |
0.06 |
|
|
|
| 28 |
A" |
1032 |
996 |
0.00 |
|
|
|
| 29 |
A" |
856 |
826 |
1.45 |
|
|
|
| 30 |
A" |
748 |
722 |
3.37 |
|
|
|
| 31 |
A" |
314 |
303 |
40.27 |
|
|
|
| 32 |
A" |
230 |
222 |
3.90 |
|
|
|
| 33 |
A" |
134 |
129 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26549.8 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 25620.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.089 |
|
|
|
| 2 |
C |
-0.118 |
|
|
|
| 3 |
C |
-0.000 |
|
|
|
| 4 |
N |
-0.213 |
|
|
|
| 5 |
H |
0.039 |
|
|
|
| 6 |
H |
0.042 |
|
|
|
| 7 |
H |
0.042 |
|
|
|
| 8 |
H |
0.027 |
|
|
|
| 9 |
H |
0.027 |
|
|
|
| 10 |
H |
0.038 |
|
|
|
| 11 |
H |
0.038 |
|
|
|
| 12 |
H |
0.083 |
|
|
|
| 13 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.246 |
1.272 |
0.000 |
1.295 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.724 |
-1.735 |
0.000 |
| y |
-1.735 |
-31.933 |
0.000 |
| z |
0.000 |
0.000 |
-25.573 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.029 |
-1.735 |
0.000 |
| y |
-1.735 |
-6.284 |
0.000 |
| z |
0.000 |
0.000 |
3.255 |
|
| Polar |
|---|
| 3z2-r2 | 6.510 |
|---|
| x2-y2 | 6.209 |
|---|
| xy | -1.735 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.834 |
0.562 |
0.000 |
| y |
0.562 |
6.303 |
0.000 |
| z |
0.000 |
0.000 |
5.906 |
<r2> (average value of r
2) Å
2
| <r2> |
112.775 |
| (<r2>)1/2 |
10.620 |