return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-173.966070
Energy at 298.15K-173.976587
HF Energy-173.298467
Nuclear repulsion energy130.447595
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3493 3342 1.74      
2 A' 3151 3014 31.88      
3 A' 3062 2930 36.44      
4 A' 3057 2925 41.50      
5 A' 3047 2915 24.34      
6 A' 1676 1603 16.63      
7 A' 1511 1445 3.39      
8 A' 1498 1433 1.15      
9 A' 1488 1424 0.06      
10 A' 1429 1367 1.33      
11 A' 1418 1357 6.76      
12 A' 1333 1276 14.58      
13 A' 1170 1119 7.58      
14 A' 1128 1079 16.29      
15 A' 1070 1024 0.64      
16 A' 970 928 126.60      
17 A' 883 845 27.87      
18 A' 453 434 2.45      
19 A' 268 256 3.46      
20 A" 3575 3420 0.53      
21 A" 3143 3007 51.62      
22 A" 3110 2975 45.55      
23 A" 3084 2951 2.12      
24 A" 1500 1435 5.92      
25 A" 1408 1347 0.68      
26 A" 1329 1272 0.69      
27 A" 1253 1199 0.00      
28 A" 1046 1000 0.23      
29 A" 867 829 1.46      
30 A" 751 718 1.31      
31 A" 316 302 42.06      
32 A" 238 228 3.69      
33 A" 137 131 0.30      

Unscaled Zero Point Vibrational Energy (zpe) 26929.6 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25763.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.81537 0.12224 0.11515

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.433 1.294 0.000
C2 0.000 0.747 0.000
C3 -0.054 -0.789 0.000
N4 -1.396 -1.383 0.000
H5 1.444 2.398 0.000
H6 1.988 0.950 0.893
H7 1.988 0.950 -0.893
H8 -0.546 1.125 0.888
H9 -0.546 1.125 -0.888
H10 0.491 -1.173 -0.885
H11 0.491 -1.173 0.885
H12 -1.907 -1.015 -0.808
H13 -1.907 -1.015 0.808

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.53392.55923.89491.10421.10591.10592.17572.17572.78552.78554.14034.1403
C21.53391.53672.54662.19412.18852.18851.10881.10882.17022.17022.71942.7194
C32.55921.53671.46793.52182.82642.82642.16662.16661.10761.10762.03462.0346
N43.89492.54661.46794.72944.20604.20602.79312.79312.09422.09421.02471.0247
H51.10422.19413.52184.72941.78591.78592.52422.52423.80113.80114.85164.8516
H61.10592.18852.82644.20601.78591.78502.54023.10183.14792.59814.68254.3635
H71.10592.18852.82644.20601.78591.78503.10182.54022.59813.14794.36354.6825
H82.17571.10882.16662.79312.52422.54023.10181.77503.08202.52133.05112.5376
H92.17571.10882.16662.79312.52423.10182.54021.77502.52133.08202.53763.0511
H102.78552.17021.10762.09423.80113.14792.59813.08202.52131.76992.40392.9394
H112.78552.17021.10762.09423.80112.59813.14792.52133.08201.76992.93942.4039
H124.14032.71942.03461.02474.85164.68254.36353.05112.53762.40392.93941.6167
H134.14032.71942.03461.02474.85164.36354.68252.53763.05112.93942.40391.6167

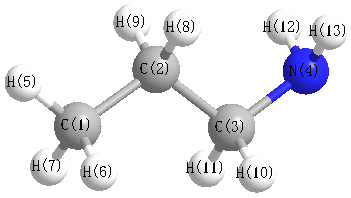
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.912 C1 C2 H8 109.784
C1 C2 H9 109.784 C2 C1 H5 111.507
C2 C1 H6 110.960 C2 C1 H7 110.960
C2 C3 N4 115.881 C2 C3 H10 109.235
C2 C3 H11 109.235 C3 C2 H8 108.898
C3 C2 H9 108.898 C3 N4 H12 108.106
C3 N4 H13 108.106 N4 C3 H10 107.989
N4 C3 H11 107.989 H5 C1 H6 107.813
H5 C1 H7 107.813 H6 C1 H7 107.622
H8 C2 H9 106.347 H10 C3 H11 106.063
H12 N4 H13 104.155
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability