Jump to
S1C2
Vibrational Frequencies calculated at BLYP/cc-pVDZ
Geometric Data calculated at BLYP/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.910151 |
| Energy at 298.15K | -207.913423 |
| HF Energy | -207.910151 |
| Nuclear repulsion energy | 100.913230 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3604 |
3609 |
23.04 |
75.19 |
0.28 |
0.43 |
| 2 |
A |
3003 |
3008 |
6.81 |
99.32 |
0.35 |
0.52 |
| 3 |
A |
2899 |
2904 |
35.77 |
132.56 |
0.18 |
0.31 |
| 4 |
A |
2249 |
2253 |
0.10 |
61.81 |
0.24 |
0.38 |
| 5 |
A |
1412 |
1414 |
2.55 |
16.10 |
0.64 |
0.78 |
| 6 |
A |
1369 |
1372 |
41.40 |
7.46 |
0.74 |
0.85 |
| 7 |
A |
1316 |
1318 |
1.68 |
6.73 |
0.72 |
0.84 |
| 8 |
A |
1179 |
1181 |
15.40 |
5.42 |
0.58 |
0.73 |
| 9 |
A |
1013 |
1014 |
86.65 |
5.63 |
0.34 |
0.50 |
| 10 |
A |
950 |
951 |
22.28 |
0.79 |
0.45 |
0.62 |
| 11 |
A |
865 |
866 |
13.14 |
2.09 |
0.17 |
0.29 |
| 12 |
A |
558 |
558 |
2.60 |
1.98 |
0.25 |
0.40 |
| 13 |
A |
383 |
384 |
58.63 |
1.31 |
0.72 |
0.84 |
| 14 |
A |
297 |
298 |
67.24 |
4.54 |
0.75 |
0.86 |
| 15 |
A |
204 |
204 |
6.06 |
3.72 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 10650.1 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 10667.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.577 |
0.596 |
0.039 |
| C2 |
0.832 |
0.127 |
-0.006 |
| O3 |
-1.524 |
-0.463 |
-0.110 |
| H4 |
-0.716 |
1.182 |
0.978 |
| H5 |
-0.749 |
1.290 |
-0.809 |
| H6 |
-1.381 |
-1.087 |
0.633 |
| N7 |
1.930 |
-0.289 |
-0.016 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4859 | 1.4286 | 1.1163 | 1.1087 | 1.9575 | 2.6594 |
C2 | 1.4859 | | 2.4309 | 2.1165 | 2.1205 | 2.6034 | 1.1742 | O3 | 1.4286 | 2.4309 | | 2.1322 | 2.0400 | 0.9806 | 3.4599 | H4 | 1.1163 | 2.1165 | 2.1322 | | 1.7912 | 2.3899 | 3.1864 | H5 | 1.1087 | 2.1205 | 2.0400 | 1.7912 | | 2.8507 | 3.2090 | H6 | 1.9575 | 2.6034 | 0.9806 | 2.3899 | 2.8507 | | 3.4673 | N7 | 2.6594 | 1.1742 | 3.4599 | 3.1864 | 3.2090 | 3.4673 | |
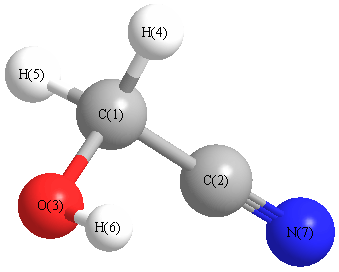 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
177.398 |
|
C1 |
O3 |
H6 |
107.217 |
| C2 |
C1 |
O3 |
113.021 |
|
C2 |
C1 |
H4 |
108.004 |
| C2 |
C1 |
H5 |
108.755 |
|
O3 |
C1 |
H4 |
113.252 |
| O3 |
C1 |
H5 |
106.341 |
|
H4 |
C1 |
H5 |
107.229 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.195 |
|
|
|
| 2 |
C |
-0.152 |
|
|
|
| 3 |
O |
-0.218 |
|
|
|
| 4 |
H |
0.043 |
|
|
|
| 5 |
H |
0.062 |
|
|
|
| 6 |
H |
0.146 |
|
|
|
| 7 |
N |
-0.076 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.309 |
0.977 |
1.197 |
2.778 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.800 |
1.159 |
-1.696 |
| y |
1.159 |
-20.565 |
-1.647 |
| z |
-1.696 |
-1.647 |
-21.711 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.662 |
1.159 |
-1.696 |
| y |
1.159 |
5.691 |
-1.647 |
| z |
-1.696 |
-1.647 |
3.971 |
|
| Polar |
|---|
| 3z2-r2 | 7.942 |
|---|
| x2-y2 | -10.235 |
|---|
| xy | 1.159 |
|---|
| xz | -1.696 |
|---|
| yz | -1.647 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.639 |
-0.443 |
-0.047 |
| y |
-0.443 |
3.807 |
-0.147 |
| z |
-0.047 |
-0.147 |
3.195 |
<r2> (average value of r
2) Å
2
| <r2> |
79.896 |
| (<r2>)1/2 |
8.938 |
