Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
Geometric Data calculated at B3PW91/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.887131 |
| Energy at 298.15K | -207.890507 |
| HF Energy | -207.887131 |
| Nuclear repulsion energy | 102.051197 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3803 |
3670 |
46.41 |
|
|
|
| 2 |
A |
3111 |
3002 |
4.03 |
|
|
|
| 3 |
A |
3013 |
2908 |
27.01 |
|
|
|
| 4 |
A |
2365 |
2282 |
0.62 |
|
|
|
| 5 |
A |
1455 |
1404 |
4.16 |
|
|
|
| 6 |
A |
1408 |
1359 |
45.85 |
|
|
|
| 7 |
A |
1362 |
1314 |
2.03 |
|
|
|
| 8 |
A |
1217 |
1175 |
13.39 |
|
|
|
| 9 |
A |
1110 |
1071 |
105.50 |
|
|
|
| 10 |
A |
979 |
944 |
16.31 |
|
|
|
| 11 |
A |
908 |
876 |
14.60 |
|
|
|
| 12 |
A |
586 |
566 |
2.23 |
|
|
|
| 13 |
A |
397 |
383 |
57.54 |
|
|
|
| 14 |
A |
311 |
301 |
82.55 |
|
|
|
| 15 |
A |
214 |
206 |
7.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11119.8 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 10730.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.574 |
0.584 |
0.038 |
| C2 |
0.823 |
0.117 |
-0.006 |
| O3 |
-1.509 |
-0.452 |
-0.111 |
| H4 |
-0.707 |
1.163 |
0.973 |
| H5 |
-0.732 |
1.282 |
-0.798 |
| H6 |
-1.388 |
-1.070 |
0.624 |
| N7 |
1.915 |
-0.280 |
-0.015 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4743 | 1.4035 | 1.1074 | 1.1007 | 1.9347 | 2.6355 |
C2 | 1.4743 | | 2.4028 | 2.0961 | 2.0986 | 2.5877 | 1.1616 | O3 | 1.4035 | 2.4028 | | 2.1040 | 2.0207 | 0.9677 | 3.4297 | H4 | 1.1074 | 2.0961 | 2.1040 | | 1.7756 | 2.3608 | 3.1515 | H5 | 1.1007 | 2.0986 | 2.0207 | 1.7756 | | 2.8260 | 3.1715 | H6 | 1.9347 | 2.5877 | 0.9677 | 2.3608 | 2.8260 | | 3.4563 | N7 | 2.6355 | 1.1616 | 3.4297 | 3.1515 | 3.1715 | 3.4563 | |
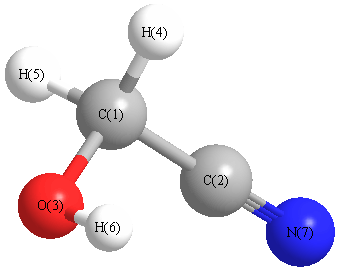 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.082 |
|
C1 |
O3 |
H6 |
107.941 |
| C2 |
C1 |
O3 |
113.200 |
|
C2 |
C1 |
H4 |
107.722 |
| C2 |
C1 |
H5 |
108.301 |
|
O3 |
C1 |
H4 |
113.317 |
| O3 |
C1 |
H5 |
106.977 |
|
H4 |
C1 |
H5 |
107.052 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.157 |
|
|
|
| 2 |
C |
-0.107 |
|
|
|
| 3 |
O |
-0.255 |
|
|
|
| 4 |
H |
0.078 |
|
|
|
| 5 |
H |
0.098 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
N |
-0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.413 |
1.040 |
1.268 |
2.917 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.913 |
1.154 |
-1.819 |
| y |
1.154 |
-20.321 |
-1.694 |
| z |
-1.819 |
-1.694 |
-21.507 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.999 |
1.154 |
-1.819 |
| y |
1.154 |
5.889 |
-1.694 |
| z |
-1.819 |
-1.694 |
4.110 |
|
| Polar |
|---|
| 3z2-r2 | 8.221 |
|---|
| x2-y2 | -10.592 |
|---|
| xy | 1.154 |
|---|
| xz | -1.819 |
|---|
| yz | -1.694 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.360 |
-0.447 |
-0.040 |
| y |
-0.447 |
3.625 |
-0.128 |
| z |
-0.040 |
-0.128 |
3.086 |
<r2> (average value of r
2) Å
2
| <r2> |
78.576 |
| (<r2>)1/2 |
8.864 |
