Jump to
S1C2
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.407586 |
| Energy at 298.15K | -207.410977 |
| HF Energy | -206.784780 |
| Nuclear repulsion energy | 101.526757 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3876 |
3708 |
42.78 |
|
|
|
| 2 |
A |
3160 |
3023 |
7.27 |
|
|
|
| 3 |
A |
3073 |
2940 |
27.99 |
|
|
|
| 4 |
A |
2358 |
2256 |
2.77 |
|
|
|
| 5 |
A |
1499 |
1435 |
1.39 |
|
|
|
| 6 |
A |
1448 |
1385 |
53.26 |
|
|
|
| 7 |
A |
1401 |
1340 |
4.35 |
|
|
|
| 8 |
A |
1244 |
1190 |
14.18 |
|
|
|
| 9 |
A |
1138 |
1089 |
94.60 |
|
|
|
| 10 |
A |
1001 |
958 |
17.09 |
|
|
|
| 11 |
A |
912 |
872 |
19.72 |
|
|
|
| 12 |
A |
582 |
557 |
2.73 |
|
|
|
| 13 |
A |
396 |
379 |
75.12 |
|
|
|
| 14 |
A |
312 |
299 |
67.97 |
|
|
|
| 15 |
A |
214 |
205 |
7.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11307.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 10817.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.585 |
0.588 |
0.038 |
| C2 |
0.829 |
0.113 |
-0.005 |
| O3 |
-1.514 |
-0.458 |
-0.113 |
| H4 |
-0.722 |
1.159 |
0.976 |
| H5 |
-0.741 |
1.283 |
-0.803 |
| H6 |
-1.400 |
-1.055 |
0.637 |
| N7 |
1.930 |
-0.276 |
-0.015 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4922 | 1.4070 | 1.1071 | 1.1015 | 1.9295 | 2.6599 |
C2 | 1.4922 | | 2.4143 | 2.1123 | 2.1143 | 2.5973 | 1.1679 | O3 | 1.4070 | 2.4143 | | 2.1037 | 2.0258 | 0.9654 | 3.4506 | H4 | 1.1071 | 2.1123 | 2.1037 | | 1.7834 | 2.3394 | 3.1743 | H5 | 1.1015 | 2.1143 | 2.0258 | 1.7834 | | 2.8236 | 3.1917 | H6 | 1.9295 | 2.5973 | 0.9654 | 2.3394 | 2.8236 | | 3.4818 | N7 | 2.6599 | 1.1679 | 3.4506 | 3.1743 | 3.1917 | 3.4818 | |
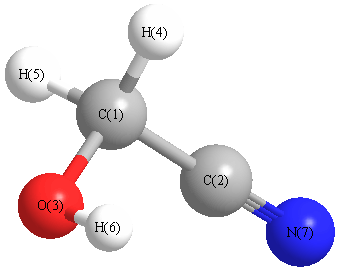 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.549 |
|
C1 |
O3 |
H6 |
107.373 |
| C2 |
C1 |
O3 |
112.731 |
|
C2 |
C1 |
H4 |
107.790 |
| C2 |
C1 |
H5 |
108.266 |
|
O3 |
C1 |
H4 |
113.058 |
| O3 |
C1 |
H5 |
107.091 |
|
H4 |
C1 |
H5 |
107.698 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
