return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-192.572323
Energy at 298.15K-192.578578
HF Energy-191.934940
Nuclear repulsion energy115.985054
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3885 3717 24.04      
2 A 3270 3128 14.58      
3 A 3197 3058 6.00      
4 A 3173 3035 9.61      
5 A 3055 2923 57.32      
6 A 3011 2881 58.56      
7 A 1747 1671 0.50      
8 A 1521 1455 1.19      
9 A 1498 1433 13.83      
10 A 1439 1376 5.04      
11 A 1311 1255 4.05      
12 A 1293 1237 44.54      
13 A 1263 1208 46.63      
14 A 1173 1122 17.83      
15 A 1124 1075 63.80      
16 A 1022 978 15.51      
17 A 979 936 4.32      
18 A 949 908 30.46      
19 A 929 889 1.74      
20 A 652 624 4.32      
21 A 448 428 3.78      
22 A 332 318 9.36      
23 A 270 258 114.89      
24 A 112 107 1.25      

Unscaled Zero Point Vibrational Energy (zpe) 18824.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 18009.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.92112 0.14239 0.13651

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.597 0.452 0.284
C2 -0.667 -0.368 0.249
C3 -1.846 0.082 -0.214
O4 1.630 -0.304 -0.333
H5 0.427 1.420 -0.232
H6 0.852 0.679 1.342
H7 -0.579 -1.389 0.645
H8 -2.745 -0.544 -0.196
H9 -1.951 1.094 -0.625
H10 2.453 0.178 -0.194

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.50752.52061.42131.11001.11162.21403.52032.78021.9358
C21.50751.34382.37102.15072.14431.09892.13202.13273.1986
C32.52061.34383.49962.63753.17082.12341.09581.09764.2999
O41.42132.37103.49962.10492.09252.64804.38403.85520.9639
H51.11002.15072.63752.10491.79103.10983.73102.43172.3762
H61.11162.14433.17082.09251.79102.60904.09843.44892.2747
H72.21401.09892.12342.64803.10982.60902.47293.10853.5143
H83.52032.13201.09584.38403.73104.09842.47291.87085.2479
H92.78022.13271.09763.85522.43173.44893.10851.87084.5182
H101.93583.19864.29990.96392.37622.27473.51435.24794.5182

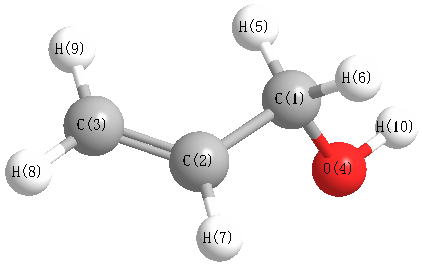
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.163 C1 C2 H7 115.412
C1 O4 H10 106.936 C2 C1 O4 108.068
C2 C1 H5 109.568 C2 C1 H6 108.981
C2 C3 H8 121.505 C2 C3 H9 121.430
C3 C2 H7 120.423 O4 C1 H5 111.926
O4 C1 H6 110.816 H5 C1 H6 107.446
H8 C3 H9 117.065
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability