Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -612.465707 |
| Energy at 298.15K | -612.469011 |
| HF Energy | -611.843694 |
| Nuclear repulsion energy | 142.147513 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3189 |
3059 |
1.57 |
|
|
|
| 2 |
A |
3105 |
2979 |
9.25 |
|
|
|
| 3 |
A |
2983 |
2862 |
71.91 |
|
|
|
| 4 |
A |
1827 |
1753 |
128.47 |
|
|
|
| 5 |
A |
1457 |
1398 |
8.06 |
|
|
|
| 6 |
A |
1414 |
1356 |
6.63 |
|
|
|
| 7 |
A |
1276 |
1224 |
20.60 |
|
|
|
| 8 |
A |
1191 |
1142 |
6.33 |
|
|
|
| 9 |
A |
1069 |
1026 |
26.20 |
|
|
|
| 10 |
A |
1033 |
991 |
1.63 |
|
|
|
| 11 |
A |
835 |
801 |
20.17 |
|
|
|
| 12 |
A |
717 |
688 |
12.84 |
|
|
|
| 13 |
A |
462 |
443 |
10.01 |
|
|
|
| 14 |
A |
289 |
278 |
9.68 |
|
|
|
| 15 |
A |
66 |
64 |
12.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10456.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 10031.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.015 |
0.685 |
0.142 |
| C2 |
1.168 |
-0.312 |
0.225 |
| Cl3 |
-1.545 |
-0.174 |
-0.084 |
| O4 |
2.263 |
-0.081 |
-0.240 |
| H5 |
-0.048 |
1.254 |
1.085 |
| H6 |
0.179 |
1.377 |
-0.698 |
| H7 |
0.931 |
-1.263 |
0.765 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5264 | 1.7952 | 2.4048 | 1.1039 | 1.1003 | 2.2406 |
C2 | 1.5264 | | 2.7344 | 1.2114 | 2.1610 | 2.1639 | 1.1187 | Cl3 | 1.7952 | 2.7344 | | 3.8119 | 2.3769 | 2.3991 | 2.8353 | O4 | 2.4048 | 1.2114 | 3.8119 | | 2.9793 | 2.5836 | 2.0442 | H5 | 1.1039 | 2.1610 | 2.3769 | 2.9793 | | 1.8015 | 2.7193 | H6 | 1.1003 | 2.1639 | 2.3991 | 2.5836 | 1.8015 | | 3.1101 | H7 | 2.2406 | 1.1187 | 2.8353 | 2.0442 | 2.7193 | 3.1101 | |
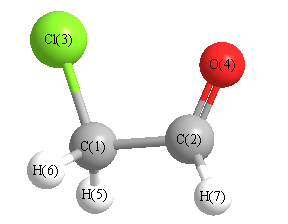 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
122.476 |
|
C1 |
C2 |
H7 |
114.922 |
| C2 |
C1 |
Cl3 |
110.556 |
|
C2 |
C1 |
H5 |
109.436 |
| C2 |
C1 |
H6 |
109.874 |
|
Cl3 |
C1 |
H5 |
107.752 |
| Cl3 |
C1 |
H6 |
109.553 |
|
O4 |
C2 |
H7 |
122.593 |
| H5 |
C1 |
H6 |
109.636 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -612.464355 |
| Energy at 298.15K | -612.467821 |
| HF Energy | -611.841468 |
| Nuclear repulsion energy | 146.007034 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3090 |
2964 |
18.40 |
|
|
|
| 2 |
A' |
2937 |
2818 |
110.39 |
|
|
|
| 3 |
A' |
1847 |
1772 |
121.24 |
|
|
|
| 4 |
A' |
1448 |
1389 |
18.12 |
|
|
|
| 5 |
A' |
1413 |
1356 |
14.67 |
|
|
|
| 6 |
A' |
1322 |
1269 |
17.06 |
|
|
|
| 7 |
A' |
967 |
928 |
5.10 |
|
|
|
| 8 |
A' |
784 |
753 |
7.22 |
|
|
|
| 9 |
A' |
641 |
615 |
41.09 |
|
|
|
| 10 |
A' |
216 |
207 |
2.64 |
|
|
|
| 11 |
A" |
3147 |
3020 |
2.99 |
|
|
|
| 12 |
A" |
1210 |
1161 |
1.92 |
|
|
|
| 13 |
A" |
1042 |
1000 |
0.89 |
|
|
|
| 14 |
A" |
704 |
675 |
0.78 |
|
|
|
| 15 |
A" |
166 |
160 |
0.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10466.7 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 10041.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.930 |
0.000 |
| C2 |
1.372 |
0.264 |
0.000 |
| Cl3 |
-1.350 |
-0.232 |
0.000 |
| O4 |
1.585 |
-0.925 |
0.000 |
| H5 |
-0.080 |
1.576 |
0.892 |
| H6 |
-0.080 |
1.576 |
-0.892 |
| H7 |
2.204 |
1.018 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5249 | 1.7810 | 2.4394 | 1.1046 | 1.1046 | 2.2060 |
C2 | 1.5249 | | 2.7674 | 1.2080 | 2.1502 | 2.1502 | 1.1224 | Cl3 | 1.7810 | 2.7674 | | 3.0163 | 2.3829 | 2.3829 | 3.7679 | O4 | 2.4394 | 1.2080 | 3.0163 | | 3.1337 | 3.1337 | 2.0387 | H5 | 1.1046 | 2.1502 | 2.3829 | 3.1337 | | 1.7844 | 2.5146 | H6 | 1.1046 | 2.1502 | 2.3829 | 3.1337 | 1.7844 | | 2.5146 | H7 | 2.2060 | 1.1224 | 3.7679 | 2.0387 | 2.5146 | 2.5146 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.017 |
|
C1 |
C2 |
H7 |
111.983 |
| C2 |
C1 |
Cl3 |
113.448 |
|
C2 |
C1 |
H5 |
108.665 |
| C2 |
C1 |
H6 |
108.665 |
|
Cl3 |
C1 |
H5 |
109.079 |
| Cl3 |
C1 |
H6 |
109.079 |
|
O4 |
C2 |
H7 |
122.001 |
| H5 |
C1 |
H6 |
107.753 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
