Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -612.458432 |
| Energy at 298.15K | -612.461769 |
| HF Energy | -611.844278 |
| Nuclear repulsion energy | 142.503461 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3058 |
1.55 |
|
|
|
| 2 |
A |
3113 |
2978 |
9.17 |
|
|
|
| 3 |
A |
3000 |
2870 |
65.66 |
|
|
|
| 4 |
A |
1872 |
1791 |
119.68 |
|
|
|
| 5 |
A |
1462 |
1399 |
8.30 |
|
|
|
| 6 |
A |
1426 |
1364 |
7.25 |
|
|
|
| 7 |
A |
1286 |
1230 |
18.93 |
|
|
|
| 8 |
A |
1197 |
1145 |
6.38 |
|
|
|
| 9 |
A |
1078 |
1031 |
24.93 |
|
|
|
| 10 |
A |
1038 |
993 |
1.87 |
|
|
|
| 11 |
A |
850 |
813 |
17.95 |
|
|
|
| 12 |
A |
727 |
696 |
12.06 |
|
|
|
| 13 |
A |
468 |
448 |
10.60 |
|
|
|
| 14 |
A |
294 |
281 |
9.26 |
|
|
|
| 15 |
A |
72 |
69 |
11.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10539.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 10082.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.013 |
0.691 |
0.143 |
| C2 |
1.169 |
-0.299 |
0.244 |
| Cl3 |
-1.537 |
-0.179 |
-0.088 |
| O4 |
2.246 |
-0.091 |
-0.256 |
| H5 |
-0.062 |
1.268 |
1.081 |
| H6 |
0.181 |
1.376 |
-0.701 |
| H7 |
0.948 |
-1.224 |
0.831 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5249 | 1.7917 | 2.4003 | 1.1035 | 1.0999 | 2.2391 |
C2 | 1.5249 | | 2.7283 | 1.2062 | 2.1606 | 2.1617 | 1.1177 | Cl3 | 1.7917 | 2.7283 | | 3.7879 | 2.3736 | 2.3963 | 2.8475 | O4 | 2.4003 | 1.2062 | 3.7879 | | 2.9941 | 2.5727 | 2.0373 | H5 | 1.1035 | 2.1606 | 2.3736 | 2.9941 | | 1.8012 | 2.7004 | H6 | 1.0999 | 2.1617 | 2.3963 | 2.5727 | 1.8012 | | 3.1134 | H7 | 2.2391 | 1.1177 | 2.8475 | 2.0373 | 2.7004 | 3.1134 | |
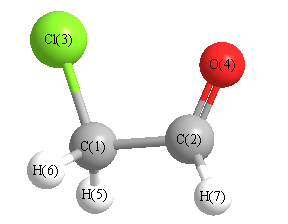 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
122.583 |
|
C1 |
C2 |
H7 |
114.968 |
| C2 |
C1 |
Cl3 |
110.435 |
|
C2 |
C1 |
H5 |
109.533 |
| C2 |
C1 |
H6 |
109.827 |
|
Cl3 |
C1 |
H5 |
107.758 |
| Cl3 |
C1 |
H6 |
109.602 |
|
O4 |
C2 |
H7 |
122.442 |
| H5 |
C1 |
H6 |
109.654 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -612.457154 |
| Energy at 298.15K | -612.460638 |
| HF Energy | -611.842064 |
| Nuclear repulsion energy | 146.280627 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3098 |
2964 |
17.76 |
|
|
|
| 2 |
A' |
2957 |
2829 |
99.19 |
|
|
|
| 3 |
A' |
1889 |
1807 |
112.03 |
|
|
|
| 4 |
A' |
1453 |
1390 |
18.93 |
|
|
|
| 5 |
A' |
1424 |
1362 |
15.52 |
|
|
|
| 6 |
A' |
1332 |
1274 |
15.92 |
|
|
|
| 7 |
A' |
976 |
934 |
5.09 |
|
|
|
| 8 |
A' |
794 |
760 |
6.36 |
|
|
|
| 9 |
A' |
648 |
620 |
40.56 |
|
|
|
| 10 |
A' |
218 |
209 |
2.65 |
|
|
|
| 11 |
A" |
3156 |
3019 |
2.86 |
|
|
|
| 12 |
A" |
1217 |
1164 |
2.12 |
|
|
|
| 13 |
A" |
1048 |
1002 |
0.66 |
|
|
|
| 14 |
A" |
708 |
678 |
0.94 |
|
|
|
| 15 |
A" |
167 |
160 |
1.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10542.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 10085.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.927 |
0.000 |
| C2 |
1.371 |
0.263 |
0.000 |
| Cl3 |
-1.349 |
-0.231 |
0.000 |
| O4 |
1.582 |
-0.921 |
0.000 |
| H5 |
-0.079 |
1.574 |
0.891 |
| H6 |
-0.079 |
1.574 |
-0.891 |
| H7 |
2.205 |
1.013 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5235 | 1.7780 | 2.4329 | 1.1042 | 1.1042 | 2.2064 |
C2 | 1.5235 | | 2.7644 | 1.2031 | 2.1488 | 2.1488 | 1.1212 | Cl3 | 1.7780 | 2.7644 | | 3.0104 | 2.3803 | 2.3803 | 3.7649 | O4 | 2.4329 | 1.2031 | 3.0104 | | 3.1274 | 3.1274 | 2.0322 | H5 | 1.1042 | 2.1488 | 2.3803 | 3.1274 | | 1.7828 | 2.5152 | H6 | 1.1042 | 2.1488 | 2.3803 | 3.1274 | 1.7828 | | 2.5152 | H7 | 2.2064 | 1.1212 | 3.7649 | 2.0322 | 2.5152 | 2.5152 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
125.919 |
|
C1 |
C2 |
H7 |
112.184 |
| C2 |
C1 |
Cl3 |
113.487 |
|
C2 |
C1 |
H5 |
108.664 |
| C2 |
C1 |
H6 |
108.664 |
|
Cl3 |
C1 |
H5 |
109.101 |
| Cl3 |
C1 |
H6 |
109.101 |
|
O4 |
C2 |
H7 |
121.897 |
| H5 |
C1 |
H6 |
107.663 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
