Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -613.066566 |
| Energy at 298.15K | -613.069721 |
| HF Energy | -613.066566 |
| Nuclear repulsion energy | 141.569784 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3079 |
3061 |
0.65 |
|
|
|
| 2 |
A |
2990 |
2973 |
6.84 |
|
|
|
| 3 |
A |
2815 |
2799 |
85.52 |
|
|
|
| 4 |
A |
1774 |
1763 |
139.47 |
|
|
|
| 5 |
A |
1359 |
1351 |
12.82 |
|
|
|
| 6 |
A |
1336 |
1328 |
3.74 |
|
|
|
| 7 |
A |
1190 |
1183 |
16.49 |
|
|
|
| 8 |
A |
1125 |
1118 |
5.20 |
|
|
|
| 9 |
A |
1015 |
1009 |
37.39 |
|
|
|
| 10 |
A |
977 |
972 |
2.28 |
|
|
|
| 11 |
A |
776 |
771 |
27.68 |
|
|
|
| 12 |
A |
663 |
659 |
11.96 |
|
|
|
| 13 |
A |
442 |
440 |
7.62 |
|
|
|
| 14 |
A |
265 |
263 |
9.95 |
|
|
|
| 15 |
A |
58 |
58 |
10.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9931.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 9874.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.024 |
0.668 |
0.135 |
| C2 |
1.173 |
-0.330 |
0.193 |
| Cl3 |
-1.561 |
-0.165 |
-0.078 |
| O4 |
2.287 |
-0.064 |
-0.213 |
| H5 |
-0.021 |
1.225 |
1.096 |
| H6 |
0.179 |
1.386 |
-0.692 |
| H7 |
0.907 |
-1.325 |
0.657 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5229 | 1.8032 | 2.4035 | 1.1109 | 1.1062 | 2.2420 |
C2 | 1.5229 | | 2.7519 | 1.2149 | 2.1579 | 2.1712 | 1.1299 | Cl3 | 1.8032 | 2.7519 | | 3.8512 | 2.3836 | 2.4106 | 2.8247 | O4 | 2.4035 | 1.2149 | 3.8512 | | 2.9489 | 2.6025 | 2.0615 | H5 | 1.1109 | 2.1579 | 2.3836 | 2.9489 | | 1.8058 | 2.7489 | H6 | 1.1062 | 2.1712 | 2.4106 | 2.6025 | 1.8058 | | 3.1147 | H7 | 2.2420 | 1.1299 | 2.8247 | 2.0615 | 2.7489 | 3.1147 | |
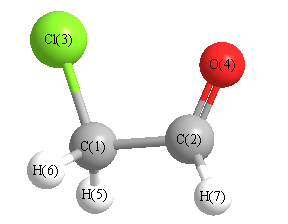 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
122.378 |
|
C1 |
C2 |
H7 |
114.565 |
| C2 |
C1 |
Cl3 |
111.383 |
|
C2 |
C1 |
H5 |
109.029 |
| C2 |
C1 |
H6 |
110.346 |
|
Cl3 |
C1 |
H5 |
107.367 |
| Cl3 |
C1 |
H6 |
109.566 |
|
O4 |
C2 |
H7 |
123.049 |
| H5 |
C1 |
H6 |
109.075 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.105 |
|
|
|
| 2 |
C |
0.141 |
|
|
|
| 3 |
Cl |
-0.108 |
|
|
|
| 4 |
O |
-0.149 |
|
|
|
| 5 |
H |
0.110 |
|
|
|
| 6 |
H |
0.097 |
|
|
|
| 7 |
H |
0.014 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.518 |
0.667 |
0.872 |
1.214 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.445 |
-0.066 |
1.138 |
| y |
-0.066 |
-28.373 |
0.124 |
| z |
1.138 |
0.124 |
-29.109 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.704 |
-0.066 |
1.138 |
| y |
-0.066 |
4.904 |
0.124 |
| z |
1.138 |
0.124 |
3.800 |
|
| Polar |
|---|
| 3z2-r2 | 7.600 |
|---|
| x2-y2 | -9.072 |
|---|
| xy | -0.066 |
|---|
| xz | 1.138 |
|---|
| yz | 0.124 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.992 |
0.439 |
-0.142 |
| y |
0.439 |
4.693 |
-0.297 |
| z |
-0.142 |
-0.297 |
3.554 |
<r2> (average value of r
2) Å
2
| <r2> |
124.037 |
| (<r2>)1/2 |
11.137 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -613.065673 |
| Energy at 298.15K | -613.069044 |
| HF Energy | -613.065673 |
| Nuclear repulsion energy | 145.480458 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2976 |
2959 |
15.92 |
|
|
|
| 2 |
A' |
2757 |
2741 |
125.59 |
|
|
|
| 3 |
A' |
1797 |
1787 |
141.37 |
|
|
|
| 4 |
A' |
1348 |
1340 |
28.85 |
|
|
|
| 5 |
A' |
1339 |
1331 |
9.16 |
|
|
|
| 6 |
A' |
1232 |
1225 |
19.82 |
|
|
|
| 7 |
A' |
910 |
905 |
4.90 |
|
|
|
| 8 |
A' |
751 |
747 |
9.46 |
|
|
|
| 9 |
A' |
619 |
615 |
41.56 |
|
|
|
| 10 |
A' |
197 |
196 |
2.04 |
|
|
|
| 11 |
A" |
3029 |
3012 |
0.47 |
|
|
|
| 12 |
A" |
1142 |
1135 |
1.72 |
|
|
|
| 13 |
A" |
981 |
975 |
1.46 |
|
|
|
| 14 |
A" |
646 |
643 |
2.25 |
|
|
|
| 15 |
A" |
186 |
185 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9954.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 9896.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.922 |
0.000 |
| C2 |
1.373 |
0.272 |
0.000 |
| Cl3 |
-1.360 |
-0.237 |
0.000 |
| O4 |
1.606 |
-0.917 |
0.000 |
| H5 |
-0.083 |
1.578 |
0.893 |
| H6 |
-0.083 |
1.578 |
-0.893 |
| H7 |
2.197 |
1.052 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5188 | 1.7867 | 2.4413 | 1.1115 | 1.1115 | 2.2004 |
C2 | 1.5188 | | 2.7794 | 1.2111 | 2.1502 | 2.1502 | 1.1350 | Cl3 | 1.7867 | 2.7794 | | 3.0429 | 2.3922 | 2.3922 | 3.7829 | O4 | 2.4413 | 1.2111 | 3.0429 | | 3.1424 | 3.1424 | 2.0557 | H5 | 1.1115 | 2.1502 | 2.3922 | 3.1424 | | 1.7867 | 2.5041 | H6 | 1.1115 | 2.1502 | 2.3922 | 3.1424 | 1.7867 | | 2.5041 | H7 | 2.2004 | 1.1350 | 3.7829 | 2.0557 | 2.5041 | 2.5041 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.463 |
|
C1 |
C2 |
H7 |
111.198 |
| C2 |
C1 |
Cl3 |
114.220 |
|
C2 |
C1 |
H5 |
108.674 |
| C2 |
C1 |
H6 |
108.674 |
|
Cl3 |
C1 |
H5 |
109.027 |
| Cl3 |
C1 |
H6 |
109.027 |
|
O4 |
C2 |
H7 |
122.339 |
| H5 |
C1 |
H6 |
106.974 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.125 |
|
|
|
| 2 |
C |
0.137 |
|
|
|
| 3 |
Cl |
-0.084 |
|
|
|
| 4 |
O |
-0.132 |
|
|
|
| 5 |
H |
0.101 |
|
|
|
| 6 |
H |
0.101 |
|
|
|
| 7 |
H |
0.002 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.484 |
3.031 |
0.000 |
3.070 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.179 |
3.151 |
0.000 |
| y |
3.151 |
-29.207 |
0.000 |
| z |
0.000 |
0.000 |
-29.141 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.005 |
3.151 |
0.000 |
| y |
3.151 |
1.453 |
0.000 |
| z |
0.000 |
0.000 |
1.552 |
|
| Polar |
|---|
| 3z2-r2 | 3.104 |
|---|
| x2-y2 | -2.972 |
|---|
| xy | 3.151 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.328 |
0.691 |
0.000 |
| y |
0.691 |
5.661 |
0.000 |
| z |
0.000 |
0.000 |
3.263 |
<r2> (average value of r
2) Å
2
| <r2> |
107.968 |
| (<r2>)1/2 |
10.391 |
