Jump to
S1C2
Vibrational Frequencies calculated at CCSD(T)/cc-pVDZ
Geometric Data calculated at CCSD(T)/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.593683 |
| Energy at 298.15K | -208.599561 |
| HF Energy | -207.900858 |
| Nuclear repulsion energy | 118.835811 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3834 |
3753 |
|
|
|
|
| 2 |
A' |
3195 |
3128 |
|
|
|
|
| 3 |
A' |
3173 |
3105 |
|
|
|
|
| 4 |
A' |
3054 |
2989 |
|
|
|
|
| 5 |
A' |
1717 |
1680 |
|
|
|
|
| 6 |
A' |
1472 |
1440 |
|
|
|
|
| 7 |
A' |
1401 |
1372 |
|
|
|
|
| 8 |
A' |
1380 |
1351 |
|
|
|
|
| 9 |
A' |
1328 |
1300 |
|
|
|
|
| 10 |
A' |
1144 |
1120 |
|
|
|
|
| 11 |
A' |
928 |
909 |
|
|
|
|
| 12 |
A' |
907 |
888 |
|
|
|
|
| 13 |
A' |
662 |
648 |
|
|
|
|
| 14 |
A' |
317 |
311 |
|
|
|
|
| 15 |
A" |
3122 |
3056 |
|
|
|
|
| 16 |
A" |
1482 |
1451 |
|
|
|
|
| 17 |
A" |
1053 |
1030 |
|
|
|
|
| 18 |
A" |
851 |
833 |
|
|
|
|
| 19 |
A" |
484 |
473 |
|
|
|
|
| 20 |
A" |
374 |
366 |
|
|
|
|
| 21 |
A" |
19 |
18 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15947.9 cm
-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 15609.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.458 |
0.481 |
0.000 |
| C2 |
0.000 |
0.870 |
0.000 |
| N3 |
1.034 |
0.091 |
0.000 |
| O4 |
0.652 |
-1.271 |
0.000 |
| H5 |
1.519 |
-1.701 |
0.000 |
| H6 |
-1.576 |
-0.613 |
0.000 |
| H7 |
-1.960 |
0.903 |
0.892 |
| H8 |
-1.960 |
0.903 |
-0.892 |
| H9 |
0.266 |
1.938 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5090 | 2.5227 | 2.7432 | 3.6912 | 1.1002 | 1.1068 | 1.1068 | 2.2572 |
C2 | 1.5090 | | 1.2950 | 2.2384 | 2.9861 | 2.1638 | 2.1533 | 2.1533 | 1.1004 | N3 | 2.5227 | 1.2950 | | 1.4145 | 1.8560 | 2.7033 | 3.2277 | 3.2277 | 2.0002 | O4 | 2.7432 | 2.2384 | 1.4145 | | 0.9673 | 2.3236 | 3.5134 | 3.5134 | 3.2321 | H5 | 3.6912 | 2.9861 | 1.8560 | 0.9673 | | 3.2807 | 4.4357 | 4.4357 | 3.8481 | H6 | 1.1002 | 2.1638 | 2.7033 | 2.3236 | 3.2807 | | 1.7999 | 1.7999 | 3.1462 | H7 | 1.1068 | 2.1533 | 3.2277 | 3.5134 | 4.4357 | 1.7999 | | 1.7842 | 2.6118 | H8 | 1.1068 | 2.1533 | 3.2277 | 3.5134 | 4.4357 | 1.7999 | 1.7842 | | 2.6118 | H9 | 2.2572 | 1.1004 | 2.0002 | 3.2321 | 3.8481 | 3.1462 | 2.6118 | 2.6118 | |
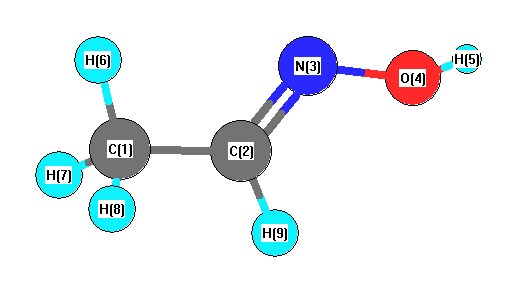 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.073 |
|
C1 |
C2 |
H9 |
118.940 |
| C2 |
C1 |
H6 |
111.082 |
|
C2 |
C1 |
H7 |
109.855 |
| C2 |
C1 |
H8 |
109.855 |
|
C2 |
N3 |
O4 |
111.334 |
| N3 |
C2 |
H9 |
112.987 |
|
N3 |
O4 |
H5 |
100.701 |
| H6 |
C1 |
H7 |
109.277 |
|
H6 |
C1 |
H8 |
109.277 |
| H7 |
C1 |
H8 |
107.417 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
