Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.573997 |
| Energy at 298.15K | -208.580016 |
| HF Energy | -207.903472 |
| Nuclear repulsion energy | 116.674034 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3858 |
3702 |
82.34 |
|
|
|
| 2 |
A |
3174 |
3045 |
13.57 |
|
|
|
| 3 |
A |
3143 |
3015 |
10.80 |
|
|
|
| 4 |
A |
3062 |
2938 |
21.42 |
|
|
|
| 5 |
A |
1750 |
1679 |
2.38 |
|
|
|
| 6 |
A |
1485 |
1424 |
9.64 |
|
|
|
| 7 |
A |
1463 |
1404 |
12.73 |
|
|
|
| 8 |
A |
1405 |
1348 |
17.81 |
|
|
|
| 9 |
A |
1294 |
1241 |
50.76 |
|
|
|
| 10 |
A |
1161 |
1114 |
2.90 |
|
|
|
| 11 |
A |
1034 |
992 |
102.75 |
|
|
|
| 12 |
A |
916 |
879 |
10.72 |
|
|
|
| 13 |
A |
568 |
545 |
13.63 |
|
|
|
| 14 |
A |
324 |
311 |
1.90 |
|
|
|
| 15 |
A |
3135 |
3007 |
17.47 |
|
|
|
| 16 |
A |
1480 |
1420 |
6.74 |
|
|
|
| 17 |
A |
1079 |
1035 |
1.20 |
|
|
|
| 18 |
A |
909 |
872 |
8.42 |
|
|
|
| 19 |
A |
392 |
376 |
127.30 |
|
|
|
| 20 |
A |
285 |
274 |
3.04 |
|
|
|
| 21 |
A |
205 |
197 |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16060.7 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15408.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.310 |
1.304 |
0.000 |
| C2 |
0.000 |
0.565 |
0.000 |
| N3 |
0.005 |
-0.721 |
0.000 |
| O4 |
1.326 |
-1.202 |
0.000 |
| H5 |
1.187 |
-2.158 |
0.000 |
| H6 |
-2.146 |
0.586 |
0.000 |
| H7 |
-1.390 |
1.953 |
0.892 |
| H8 |
-1.390 |
1.953 |
-0.892 |
| H9 |
0.951 |
1.123 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5038 | 2.4146 | 3.6370 | 4.2679 | 1.1022 | 1.1055 | 1.1055 | 2.2680 |
C2 | 1.5038 | | 1.2858 | 2.2092 | 2.9699 | 2.1464 | 2.1574 | 2.1574 | 1.1030 | N3 | 2.4146 | 1.2858 | | 1.4055 | 1.8600 | 2.5178 | 3.1452 | 3.1452 | 2.0725 | O4 | 3.6370 | 2.2092 | 1.4055 | | 0.9656 | 3.9059 | 4.2576 | 4.2576 | 2.3553 | H5 | 4.2679 | 2.9699 | 1.8600 | 0.9656 | | 4.3173 | 4.9327 | 4.9327 | 3.2893 | H6 | 1.1022 | 2.1464 | 2.5178 | 3.9059 | 4.3173 | | 1.7980 | 1.7980 | 3.1436 | H7 | 1.1055 | 2.1574 | 3.1452 | 4.2576 | 4.9327 | 1.7980 | | 1.7830 | 2.6394 | H8 | 1.1055 | 2.1574 | 3.1452 | 4.2576 | 4.9327 | 1.7980 | 1.7830 | | 2.6394 | H9 | 2.2680 | 1.1030 | 2.0725 | 2.3553 | 3.2893 | 3.1436 | 2.6394 | 2.6394 | |
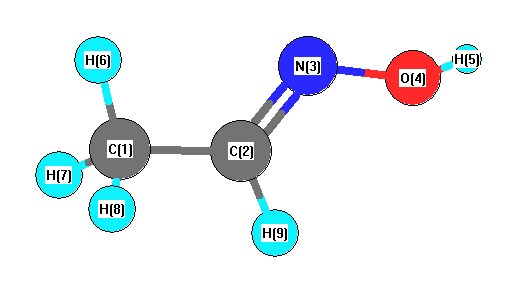 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.687 |
|
C1 |
C2 |
H9 |
120.145 |
| C2 |
C1 |
H6 |
109.938 |
|
C2 |
C1 |
H7 |
110.616 |
| C2 |
C1 |
H8 |
110.616 |
|
C2 |
N3 |
O4 |
110.262 |
| N3 |
C2 |
H9 |
120.168 |
|
N3 |
O4 |
H5 |
101.723 |
| H6 |
C1 |
H7 |
109.056 |
|
H6 |
C1 |
H8 |
109.056 |
| H7 |
C1 |
H8 |
107.501 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.573316 |
| Energy at 298.15K | |
| HF Energy | -207.902144 |
| Nuclear repulsion energy | 119.156951 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3863 |
3706 |
85.45 |
|
|
|
| 2 |
A' |
3209 |
3079 |
7.56 |
|
|
|
| 3 |
A' |
3192 |
3062 |
7.59 |
|
|
|
| 4 |
A' |
3069 |
2944 |
16.32 |
|
|
|
| 5 |
A' |
1758 |
1687 |
6.80 |
|
|
|
| 6 |
A' |
1481 |
1421 |
16.02 |
|
|
|
| 7 |
A' |
1416 |
1359 |
21.55 |
|
|
|
| 8 |
A' |
1394 |
1337 |
31.59 |
|
|
|
| 9 |
A' |
1346 |
1292 |
24.96 |
|
|
|
| 10 |
A' |
1155 |
1108 |
7.93 |
|
|
|
| 11 |
A' |
951 |
913 |
104.16 |
|
|
|
| 12 |
A' |
926 |
888 |
3.85 |
|
|
|
| 13 |
A' |
671 |
644 |
12.54 |
|
|
|
| 14 |
A' |
320 |
307 |
1.31 |
|
|
|
| 15 |
A" |
3136 |
3009 |
14.98 |
|
|
|
| 16 |
A" |
1491 |
1430 |
7.91 |
|
|
|
| 17 |
A" |
1066 |
1023 |
0.91 |
|
|
|
| 18 |
A" |
867 |
832 |
10.35 |
|
|
|
| 19 |
A" |
491 |
471 |
31.26 |
|
|
|
| 20 |
A" |
375 |
360 |
89.15 |
|
|
|
| 21 |
A" |
18i |
17i |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16079.7 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15426.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.457 |
0.480 |
0.000 |
| C2 |
0.000 |
0.866 |
0.000 |
| N3 |
1.029 |
0.090 |
0.000 |
| O4 |
0.656 |
-1.267 |
0.000 |
| H5 |
1.520 |
-1.698 |
0.000 |
| H6 |
-1.579 |
-0.612 |
0.000 |
| H7 |
-1.958 |
0.902 |
0.891 |
| H8 |
-1.958 |
0.902 |
-0.891 |
| H9 |
0.266 |
1.932 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5076 | 2.5166 | 2.7419 | 3.6888 | 1.0990 | 1.1054 | 1.1054 | 2.2534 |
C2 | 1.5076 | | 1.2888 | 2.2313 | 2.9807 | 2.1632 | 2.1512 | 2.1512 | 1.0987 | N3 | 2.5166 | 1.2888 | | 1.4067 | 1.8542 | 2.7011 | 3.2208 | 3.2208 | 1.9939 | O4 | 2.7419 | 2.2313 | 1.4067 | | 0.9654 | 2.3294 | 3.5114 | 3.5114 | 3.2224 | H5 | 3.6888 | 2.9807 | 1.8542 | 0.9654 | | 3.2839 | 4.4327 | 4.4327 | 3.8406 | H6 | 1.0990 | 2.1632 | 2.7011 | 2.3294 | 3.2839 | | 1.7972 | 1.7972 | 3.1429 | H7 | 1.1054 | 2.1512 | 3.2208 | 3.5114 | 4.4327 | 1.7972 | | 1.7813 | 2.6074 | H8 | 1.1054 | 2.1512 | 3.2208 | 3.5114 | 4.4327 | 1.7972 | 1.7813 | | 2.6074 | H9 | 2.2534 | 1.0987 | 1.9939 | 3.2224 | 3.8406 | 3.1429 | 2.6074 | 2.6074 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.134 |
|
C1 |
C2 |
H9 |
118.839 |
| C2 |
C1 |
H6 |
111.211 |
|
C2 |
C1 |
H7 |
109.867 |
| C2 |
C1 |
H8 |
109.867 |
|
C2 |
N3 |
O4 |
111.670 |
| N3 |
C2 |
H9 |
113.028 |
|
N3 |
O4 |
H5 |
101.192 |
| H6 |
C1 |
H7 |
109.226 |
|
H6 |
C1 |
H8 |
109.226 |
| H7 |
C1 |
H8 |
107.358 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
